Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.567022 |
| Energy at 298.15K | -208.573062 |
| HF Energy | -207.904206 |
| Nuclear repulsion energy | 116.909548 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3888 |
3720 |
83.16 |
|
|
|
| 2 |
A |
3184 |
3046 |
12.69 |
|
|
|
| 3 |
A |
3153 |
3016 |
9.86 |
|
|
|
| 4 |
A |
3071 |
2938 |
19.94 |
|
|
|
| 5 |
A |
1776 |
1699 |
1.02 |
|
|
|
| 6 |
A |
1490 |
1425 |
11.23 |
|
|
|
| 7 |
A |
1472 |
1409 |
11.85 |
|
|
|
| 8 |
A |
1411 |
1350 |
19.28 |
|
|
|
| 9 |
A |
1306 |
1250 |
51.22 |
|
|
|
| 10 |
A |
1165 |
1114 |
2.92 |
|
|
|
| 11 |
A |
1053 |
1008 |
103.35 |
|
|
|
| 12 |
A |
926 |
886 |
5.24 |
|
|
|
| 13 |
A |
574 |
549 |
13.55 |
|
|
|
| 14 |
A |
327 |
313 |
1.70 |
|
|
|
| 15 |
A |
3145 |
3009 |
16.65 |
|
|
|
| 16 |
A |
1482 |
1418 |
7.01 |
|
|
|
| 17 |
A |
1084 |
1037 |
1.47 |
|
|
|
| 18 |
A |
915 |
876 |
9.70 |
|
|
|
| 19 |
A |
389 |
372 |
129.35 |
|
|
|
| 20 |
A |
289 |
277 |
3.19 |
|
|
|
| 21 |
A |
207 |
198 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16153.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15453.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.310 |
1.301 |
0.000 |
| C2 |
0.000 |
0.563 |
0.000 |
| N3 |
0.009 |
-0.719 |
0.000 |
| O4 |
1.323 |
-1.199 |
0.000 |
| H5 |
1.189 |
-2.154 |
0.000 |
| H6 |
-2.146 |
0.583 |
0.000 |
| H7 |
-1.392 |
1.949 |
0.891 |
| H8 |
-1.392 |
1.949 |
-0.891 |
| H9 |
0.950 |
1.122 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5037 | 2.4127 | 3.6308 | 4.2639 | 1.1018 | 1.1051 | 1.1051 | 2.2670 |
C2 | 1.5037 | | 1.2820 | 2.2032 | 2.9653 | 2.1465 | 2.1574 | 2.1574 | 1.1024 | N3 | 2.4127 | 1.2820 | | 1.3989 | 1.8575 | 2.5186 | 3.1430 | 3.1430 | 2.0678 | O4 | 3.6308 | 2.2032 | 1.3989 | | 0.9641 | 3.9007 | 4.2518 | 4.2518 | 2.3508 | H5 | 4.2639 | 2.9653 | 1.8575 | 0.9641 | | 4.3150 | 4.9287 | 4.9287 | 3.2845 | H6 | 1.1018 | 2.1465 | 2.5186 | 3.9007 | 4.3150 | | 1.7968 | 1.7968 | 3.1428 | H7 | 1.1051 | 2.1574 | 3.1430 | 4.2518 | 4.9287 | 1.7968 | | 1.7821 | 2.6387 | H8 | 1.1051 | 2.1574 | 3.1430 | 4.2518 | 4.9287 | 1.7968 | 1.7821 | | 2.6387 | H9 | 2.2670 | 1.1024 | 2.0678 | 2.3508 | 3.2845 | 3.1428 | 2.6387 | 2.6387 | |
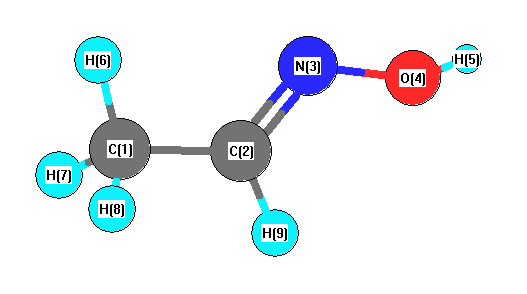 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.809 |
|
C1 |
C2 |
H9 |
120.107 |
| C2 |
C1 |
H6 |
109.982 |
|
C2 |
C1 |
H7 |
110.650 |
| C2 |
C1 |
H8 |
110.650 |
|
C2 |
N3 |
O4 |
110.462 |
| N3 |
C2 |
H9 |
120.084 |
|
N3 |
O4 |
H5 |
102.071 |
| H6 |
C1 |
H7 |
109.013 |
|
H6 |
C1 |
H8 |
109.013 |
| H7 |
C1 |
H8 |
107.471 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.566217 |
| Energy at 298.15K | -208.572161 |
| HF Energy | -207.902911 |
| Nuclear repulsion energy | 119.403041 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3892 |
3724 |
87.03 |
|
|
|
| 2 |
A' |
3218 |
3079 |
6.66 |
|
|
|
| 3 |
A' |
3199 |
3061 |
8.79 |
|
|
|
| 4 |
A' |
3078 |
2945 |
15.34 |
|
|
|
| 5 |
A' |
1784 |
1707 |
3.44 |
|
|
|
| 6 |
A' |
1485 |
1421 |
17.76 |
|
|
|
| 7 |
A' |
1424 |
1363 |
29.00 |
|
|
|
| 8 |
A' |
1405 |
1344 |
27.57 |
|
|
|
| 9 |
A' |
1356 |
1298 |
23.77 |
|
|
|
| 10 |
A' |
1159 |
1109 |
7.85 |
|
|
|
| 11 |
A' |
973 |
931 |
101.93 |
|
|
|
| 12 |
A' |
934 |
894 |
0.61 |
|
|
|
| 13 |
A' |
677 |
648 |
12.14 |
|
|
|
| 14 |
A' |
322 |
308 |
1.47 |
|
|
|
| 15 |
A" |
3147 |
3011 |
14.24 |
|
|
|
| 16 |
A" |
1493 |
1428 |
8.24 |
|
|
|
| 17 |
A" |
1072 |
1026 |
1.02 |
|
|
|
| 18 |
A" |
874 |
837 |
10.06 |
|
|
|
| 19 |
A" |
493 |
472 |
26.86 |
|
|
|
| 20 |
A" |
377 |
361 |
96.27 |
|
|
|
| 21 |
A" |
24 |
23 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16194.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15492.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.457 |
0.478 |
0.000 |
| C2 |
0.000 |
0.864 |
0.000 |
| N3 |
1.025 |
0.089 |
0.000 |
| O4 |
0.659 |
-1.262 |
0.000 |
| H5 |
1.520 |
-1.695 |
0.000 |
| H6 |
-1.581 |
-0.614 |
0.000 |
| H7 |
-1.957 |
0.900 |
0.890 |
| H8 |
-1.957 |
0.900 |
-0.890 |
| H9 |
0.264 |
1.930 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5071 | 2.5125 | 2.7394 | 3.6858 | 1.0986 | 1.1051 | 1.1051 | 2.2518 |
C2 | 1.5071 | | 1.2849 | 2.2251 | 2.9763 | 2.1639 | 2.1506 | 2.1506 | 1.0983 | N3 | 2.5125 | 1.2849 | | 1.3995 | 1.8518 | 2.6996 | 3.2166 | 3.2166 | 1.9915 | O4 | 2.7394 | 2.2251 | 1.3995 | | 0.9639 | 2.3324 | 3.5090 | 3.5090 | 3.2155 | H5 | 3.6858 | 2.9763 | 1.8518 | 0.9639 | | 3.2846 | 4.4296 | 4.4296 | 3.8362 | H6 | 1.0986 | 2.1639 | 2.6996 | 2.3324 | 3.2846 | | 1.7960 | 1.7960 | 3.1424 | H7 | 1.1051 | 2.1506 | 3.2166 | 3.5090 | 4.4296 | 1.7960 | | 1.7803 | 2.6055 | H8 | 1.1051 | 2.1506 | 3.2166 | 3.5090 | 4.4296 | 1.7960 | 1.7803 | | 2.6055 | H9 | 2.2518 | 1.0983 | 1.9915 | 3.2155 | 3.8362 | 3.1424 | 2.6055 | 2.6055 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.110 |
|
C1 |
C2 |
H9 |
118.757 |
| C2 |
C1 |
H6 |
111.323 |
|
C2 |
C1 |
H7 |
109.876 |
| C2 |
C1 |
H8 |
109.876 |
|
C2 |
N3 |
O4 |
111.902 |
| N3 |
C2 |
H9 |
113.132 |
|
N3 |
O4 |
H5 |
101.575 |
| H6 |
C1 |
H7 |
109.175 |
|
H6 |
C1 |
H8 |
109.175 |
| H7 |
C1 |
H8 |
107.323 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
