Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.900020 |
| Energy at 298.15K | -208.906006 |
| Nuclear repulsion energy | 116.469838 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3659 |
3638 |
77.56 |
|
|
|
| 2 |
A |
3079 |
3061 |
9.40 |
|
|
|
| 3 |
A |
3013 |
2995 |
20.24 |
|
|
|
| 4 |
A |
2961 |
2944 |
23.27 |
|
|
|
| 5 |
A |
1668 |
1659 |
0.07 |
|
|
|
| 6 |
A |
1407 |
1399 |
14.65 |
|
|
|
| 7 |
A |
1385 |
1377 |
8.30 |
|
|
|
| 8 |
A |
1328 |
1320 |
19.62 |
|
|
|
| 9 |
A |
1225 |
1218 |
47.29 |
|
|
|
| 10 |
A |
1117 |
1111 |
2.86 |
|
|
|
| 11 |
A |
992 |
986 |
152.33 |
|
|
|
| 12 |
A |
883 |
878 |
12.72 |
|
|
|
| 13 |
A |
550 |
547 |
13.21 |
|
|
|
| 14 |
A |
312 |
310 |
3.28 |
|
|
|
| 15 |
A |
3023 |
3005 |
12.75 |
|
|
|
| 16 |
A |
1397 |
1389 |
7.94 |
|
|
|
| 17 |
A |
1018 |
1012 |
0.18 |
|
|
|
| 18 |
A |
872 |
867 |
7.76 |
|
|
|
| 19 |
A |
440 |
437 |
116.78 |
|
|
|
| 20 |
A |
278 |
277 |
0.42 |
|
|
|
| 21 |
A |
217 |
216 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15412.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15322.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.302 |
1.305 |
0.000 |
| C2 |
0.000 |
0.572 |
0.000 |
| N3 |
0.006 |
-0.718 |
0.000 |
| O4 |
1.322 |
-1.214 |
0.000 |
| H5 |
1.164 |
-2.179 |
0.000 |
| H6 |
-2.146 |
0.590 |
0.000 |
| H7 |
-1.389 |
1.963 |
0.891 |
| H8 |
-1.389 |
1.963 |
-0.891 |
| H9 |
0.954 |
1.139 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4943 | 2.4090 | 3.6377 | 4.2683 | 1.1066 | 1.1112 | 1.1112 | 2.2622 |
C2 | 1.4943 | | 1.2896 | 2.2220 | 2.9867 | 2.1460 | 2.1588 | 2.1588 | 1.1099 | N3 | 2.4090 | 1.2896 | | 1.4068 | 1.8642 | 2.5178 | 3.1510 | 3.1510 | 2.0848 | O4 | 3.6377 | 2.2220 | 1.4068 | | 0.9778 | 3.9091 | 4.2710 | 4.2710 | 2.3814 | H5 | 4.2683 | 2.9867 | 1.8642 | 0.9778 | | 4.3148 | 4.9466 | 4.9466 | 3.3243 | H6 | 1.1066 | 2.1460 | 2.5178 | 3.9091 | 4.3148 | | 1.8038 | 1.8038 | 3.1484 | H7 | 1.1112 | 2.1588 | 3.1510 | 4.2710 | 4.9466 | 1.8038 | | 1.7822 | 2.6392 | H8 | 1.1112 | 2.1588 | 3.1510 | 4.2710 | 4.9466 | 1.8038 | 1.7822 | | 2.6392 | H9 | 2.2622 | 1.1099 | 2.0848 | 2.3814 | 3.3243 | 3.1484 | 2.6392 | 2.6392 | |
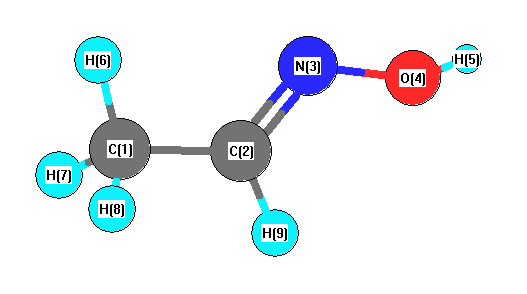 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.653 |
|
C1 |
C2 |
H9 |
119.885 |
| C2 |
C1 |
H6 |
110.308 |
|
C2 |
C1 |
H7 |
111.047 |
| C2 |
C1 |
H8 |
111.047 |
|
C2 |
N3 |
O4 |
110.911 |
| N3 |
C2 |
H9 |
120.462 |
|
N3 |
O4 |
H5 |
101.317 |
| H6 |
C1 |
H7 |
108.848 |
|
H6 |
C1 |
H8 |
108.848 |
| H7 |
C1 |
H8 |
106.635 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.036 |
|
|
|
| 2 |
C |
0.003 |
|
|
|
| 3 |
N |
-0.109 |
|
|
|
| 4 |
O |
-0.161 |
|
|
|
| 5 |
H |
0.158 |
|
|
|
| 6 |
H |
0.043 |
|
|
|
| 7 |
H |
0.053 |
|
|
|
| 8 |
H |
0.053 |
|
|
|
| 9 |
H |
-0.005 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.256 |
0.246 |
0.000 |
0.355 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.950 |
-2.250 |
0.000 |
| y |
-2.250 |
-18.598 |
0.000 |
| z |
0.000 |
0.000 |
-24.862 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.219 |
-2.250 |
0.000 |
| y |
-2.250 |
6.308 |
0.000 |
| z |
0.000 |
0.000 |
-3.088 |
|
| Polar |
|---|
| 3z2-r2 | -6.177 |
|---|
| x2-y2 | -6.351 |
|---|
| xy | -2.250 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.672 |
-1.537 |
0.000 |
| y |
-1.537 |
7.194 |
0.000 |
| z |
0.000 |
0.000 |
3.212 |
<r2> (average value of r
2) Å
2
| <r2> |
92.400 |
| (<r2>)1/2 |
9.612 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.899714 |
| Energy at 298.15K | -208.905621 |
| HF Energy | -208.899714 |
| Nuclear repulsion energy | 118.917128 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3667 |
3645 |
78.88 |
|
|
|
| 2 |
A' |
3108 |
3090 |
5.28 |
|
|
|
| 3 |
A' |
3084 |
3066 |
7.27 |
|
|
|
| 4 |
A' |
2971 |
2953 |
13.62 |
|
|
|
| 5 |
A' |
1672 |
1662 |
7.33 |
|
|
|
| 6 |
A' |
1401 |
1393 |
20.54 |
|
|
|
| 7 |
A' |
1335 |
1327 |
22.15 |
|
|
|
| 8 |
A' |
1317 |
1309 |
27.57 |
|
|
|
| 9 |
A' |
1279 |
1272 |
29.21 |
|
|
|
| 10 |
A' |
1106 |
1100 |
7.30 |
|
|
|
| 11 |
A' |
908 |
903 |
151.78 |
|
|
|
| 12 |
A' |
901 |
896 |
0.20 |
|
|
|
| 13 |
A' |
656 |
652 |
11.87 |
|
|
|
| 14 |
A' |
301 |
299 |
1.13 |
|
|
|
| 15 |
A" |
3031 |
3013 |
9.44 |
|
|
|
| 16 |
A" |
1407 |
1398 |
8.85 |
|
|
|
| 17 |
A" |
1002 |
997 |
0.11 |
|
|
|
| 18 |
A" |
819 |
814 |
15.12 |
|
|
|
| 19 |
A" |
507 |
504 |
39.96 |
|
|
|
| 20 |
A" |
385 |
383 |
65.12 |
|
|
|
| 21 |
A" |
54 |
53 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15454.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15364.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.451 |
0.497 |
0.000 |
| C2 |
0.000 |
0.867 |
0.000 |
| N3 |
1.027 |
0.081 |
0.000 |
| O4 |
0.651 |
-1.278 |
0.000 |
| H5 |
1.529 |
-1.708 |
0.000 |
| H6 |
-1.580 |
-0.600 |
0.000 |
| H7 |
-1.962 |
0.922 |
0.890 |
| H8 |
-1.962 |
0.922 |
-0.890 |
| H9 |
0.285 |
1.934 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4979 | 2.5126 | 2.7514 | 3.7072 | 1.1041 | 1.1107 | 1.1107 | 2.2544 |
C2 | 1.4979 | | 1.2928 | 2.2415 | 2.9948 | 2.1559 | 2.1552 | 2.1552 | 1.1044 | N3 | 2.5126 | 1.2928 | | 1.4097 | 1.8585 | 2.6939 | 3.2295 | 3.2295 | 1.9954 | O4 | 2.7514 | 2.2415 | 1.4097 | | 0.9773 | 2.3319 | 3.5297 | 3.5297 | 3.2323 | H5 | 3.7072 | 2.9948 | 1.8585 | 0.9773 | | 3.3003 | 4.4602 | 4.4602 | 3.8484 | H6 | 1.1041 | 2.1559 | 2.6939 | 2.3319 | 3.3003 | | 1.8037 | 1.8037 | 3.1463 | H7 | 1.1107 | 2.1552 | 3.2295 | 3.5297 | 4.4602 | 1.8037 | | 1.7805 | 2.6207 | H8 | 1.1107 | 2.1552 | 3.2295 | 3.5297 | 4.4602 | 1.8037 | 1.7805 | | 2.6207 | H9 | 2.2544 | 1.1044 | 1.9954 | 3.2323 | 3.8484 | 3.1463 | 2.6207 | 2.6207 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.253 |
|
C1 |
C2 |
H9 |
119.294 |
| C2 |
C1 |
H6 |
110.992 |
|
C2 |
C1 |
H7 |
110.536 |
| C2 |
C1 |
H8 |
110.536 |
|
C2 |
N3 |
O4 |
112.001 |
| N3 |
C2 |
H9 |
112.454 |
|
N3 |
O4 |
H5 |
100.697 |
| H6 |
C1 |
H7 |
109.053 |
|
H6 |
C1 |
H8 |
109.053 |
| H7 |
C1 |
H8 |
106.551 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.051 |
|
|
|
| 2 |
C |
-0.024 |
|
|
|
| 3 |
N |
-0.112 |
|
|
|
| 4 |
O |
-0.165 |
|
|
|
| 5 |
H |
0.163 |
|
|
|
| 6 |
H |
0.055 |
|
|
|
| 7 |
H |
0.055 |
|
|
|
| 8 |
H |
0.055 |
|
|
|
| 9 |
H |
0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.500 |
-0.034 |
0.000 |
0.501 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.658 |
-3.803 |
0.000 |
| y |
-3.803 |
-21.037 |
0.000 |
| z |
0.000 |
0.000 |
-24.867 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.294 |
-3.803 |
0.000 |
| y |
-3.803 |
2.726 |
0.000 |
| z |
0.000 |
0.000 |
-3.020 |
|
| Polar |
|---|
| 3z2-r2 | -6.039 |
|---|
| x2-y2 | -1.621 |
|---|
| xy | -3.803 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.026 |
-1.279 |
0.000 |
| y |
-1.279 |
6.184 |
0.000 |
| z |
0.000 |
0.000 |
3.202 |
<r2> (average value of r
2) Å
2
| <r2> |
79.697 |
| (<r2>)1/2 |
8.927 |
