Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.899707 |
| Energy at 298.15K | -208.905618 |
| HF Energy | -208.899707 |
| Nuclear repulsion energy | 118.912455 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3664 |
3643 |
78.91 |
|
|
|
| 2 |
A' |
3107 |
3089 |
5.17 |
|
|
|
| 3 |
A' |
3083 |
3065 |
7.38 |
|
|
|
| 4 |
A' |
2971 |
2954 |
13.62 |
|
|
|
| 5 |
A' |
1672 |
1662 |
7.32 |
|
|
|
| 6 |
A' |
1401 |
1393 |
20.63 |
|
|
|
| 7 |
A' |
1335 |
1327 |
22.85 |
|
|
|
| 8 |
A' |
1317 |
1309 |
27.47 |
|
|
|
| 9 |
A' |
1279 |
1272 |
28.70 |
|
|
|
| 10 |
A' |
1106 |
1099 |
7.30 |
|
|
|
| 11 |
A' |
908 |
903 |
151.66 |
|
|
|
| 12 |
A' |
901 |
896 |
0.21 |
|
|
|
| 13 |
A' |
655 |
651 |
11.83 |
|
|
|
| 14 |
A' |
300 |
298 |
1.11 |
|
|
|
| 15 |
A" |
3031 |
3013 |
9.45 |
|
|
|
| 16 |
A" |
1407 |
1399 |
8.88 |
|
|
|
| 17 |
A" |
1002 |
996 |
0.10 |
|
|
|
| 18 |
A" |
818 |
813 |
15.17 |
|
|
|
| 19 |
A" |
507 |
504 |
40.60 |
|
|
|
| 20 |
A" |
386 |
384 |
64.46 |
|
|
|
| 21 |
A" |
56 |
56 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15452.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15362.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.452 |
0.497 |
0.000 |
| C2 |
0.000 |
0.867 |
0.000 |
| N3 |
1.027 |
0.081 |
0.000 |
| O4 |
0.652 |
-1.278 |
0.000 |
| H5 |
1.529 |
-1.708 |
0.000 |
| H6 |
-1.581 |
-0.599 |
0.000 |
| H7 |
-1.962 |
0.922 |
0.890 |
| H8 |
-1.962 |
0.922 |
-0.890 |
| H9 |
0.285 |
1.934 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4979 | 2.5128 | 2.7519 | 3.7077 | 1.1041 | 1.1107 | 1.1107 | 2.2541 |
C2 | 1.4979 | | 1.2928 | 2.2415 | 2.9949 | 2.1564 | 2.1551 | 2.1551 | 1.1043 | N3 | 2.5128 | 1.2928 | | 1.4097 | 1.8586 | 2.6951 | 3.2296 | 3.2296 | 1.9953 | O4 | 2.7519 | 2.2415 | 1.4097 | | 0.9773 | 2.3335 | 3.5303 | 3.5303 | 3.2323 | H5 | 3.7077 | 2.9949 | 1.8586 | 0.9773 | | 3.3019 | 4.4607 | 4.4607 | 3.8484 | H6 | 1.1041 | 2.1564 | 2.6951 | 2.3335 | 3.3019 | | 1.8036 | 1.8036 | 3.1466 | H7 | 1.1107 | 2.1551 | 3.2296 | 3.5303 | 4.4607 | 1.8036 | | 1.7805 | 2.6203 | H8 | 1.1107 | 2.1551 | 3.2296 | 3.5303 | 4.4607 | 1.8036 | 1.7805 | | 2.6203 | H9 | 2.2541 | 1.1043 | 1.9953 | 3.2323 | 3.8484 | 3.1466 | 2.6203 | 2.6203 | |
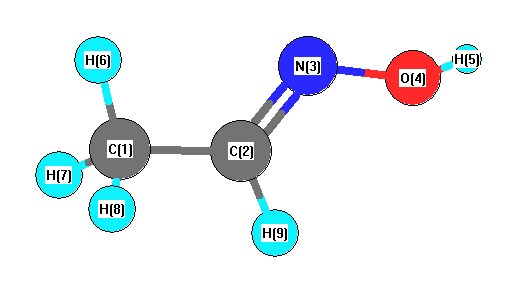 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.274 |
|
C1 |
C2 |
H9 |
119.275 |
| C2 |
C1 |
H6 |
111.037 |
|
C2 |
C1 |
H7 |
110.528 |
| C2 |
C1 |
H8 |
110.528 |
|
C2 |
N3 |
O4 |
112.005 |
| N3 |
C2 |
H9 |
112.451 |
|
N3 |
O4 |
H5 |
100.703 |
| H6 |
C1 |
H7 |
109.042 |
|
H6 |
C1 |
H8 |
109.042 |
| H7 |
C1 |
H8 |
106.543 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.051 |
|
|
|
| 2 |
C |
-0.023 |
|
|
|
| 3 |
N |
-0.112 |
|
|
|
| 4 |
O |
-0.165 |
|
|
|
| 5 |
H |
0.163 |
|
|
|
| 6 |
H |
0.055 |
|
|
|
| 7 |
H |
0.055 |
|
|
|
| 8 |
H |
0.055 |
|
|
|
| 9 |
H |
0.023 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.500 |
-0.034 |
0.000 |
0.502 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.656 |
-3.803 |
0.000 |
| y |
-3.803 |
-21.037 |
0.000 |
| z |
0.000 |
0.000 |
-24.867 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.296 |
-3.803 |
0.000 |
| y |
-3.803 |
2.725 |
0.000 |
| z |
0.000 |
0.000 |
-3.021 |
|
| Polar |
|---|
| 3z2-r2 | -6.041 |
|---|
| x2-y2 | -1.620 |
|---|
| xy | -3.803 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.026 |
-1.280 |
0.000 |
| y |
-1.280 |
6.185 |
0.000 |
| z |
0.000 |
0.000 |
3.202 |
<r2> (average value of r
2) Å
2
| <r2> |
79.708 |
| (<r2>)1/2 |
8.928 |
