Jump to
S1C2
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Geometric Data calculated at MP3=FULL/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.571166 |
| Energy at 298.15K | -208.577138 |
| HF Energy | -207.903439 |
| Nuclear repulsion energy | 119.625440 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3927 |
3692 |
92.38 |
|
|
|
| 2 |
A' |
3240 |
3047 |
6.89 |
|
|
|
| 3 |
A' |
3223 |
3031 |
7.78 |
|
|
|
| 4 |
A' |
3100 |
2915 |
14.01 |
|
|
|
| 5 |
A' |
1795 |
1688 |
3.96 |
|
|
|
| 6 |
A' |
1488 |
1399 |
20.12 |
|
|
|
| 7 |
A' |
1428 |
1343 |
33.94 |
|
|
|
| 8 |
A' |
1409 |
1325 |
25.35 |
|
|
|
| 9 |
A' |
1361 |
1280 |
22.60 |
|
|
|
| 10 |
A' |
1163 |
1093 |
8.07 |
|
|
|
| 11 |
A' |
982 |
923 |
107.64 |
|
|
|
| 12 |
A' |
939 |
883 |
2.12 |
|
|
|
| 13 |
A' |
681 |
640 |
12.20 |
|
|
|
| 14 |
A' |
323 |
304 |
1.47 |
|
|
|
| 15 |
A" |
3170 |
2980 |
12.96 |
|
|
|
| 16 |
A" |
1496 |
1407 |
8.72 |
|
|
|
| 17 |
A" |
1073 |
1009 |
0.98 |
|
|
|
| 18 |
A" |
878 |
826 |
10.54 |
|
|
|
| 19 |
A" |
497 |
467 |
27.18 |
|
|
|
| 20 |
A" |
380 |
358 |
96.72 |
|
|
|
| 21 |
A" |
31 |
29 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16292.0 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 15319.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.454 |
0.476 |
0.000 |
| C2 |
0.000 |
0.863 |
0.000 |
| N3 |
1.023 |
0.089 |
0.000 |
| O4 |
0.659 |
-1.259 |
0.000 |
| H5 |
1.517 |
-1.693 |
0.000 |
| H6 |
-1.576 |
-0.613 |
0.000 |
| H7 |
-1.955 |
0.897 |
0.888 |
| H8 |
-1.955 |
0.897 |
-0.888 |
| H9 |
0.264 |
1.927 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5049 | 2.5075 | 2.7343 | 3.6787 | 1.0965 | 1.1027 | 1.1027 | 2.2488 |
C2 | 1.5049 | | 1.2830 | 2.2221 | 2.9721 | 2.1596 | 2.1472 | 2.1472 | 1.0958 | N3 | 2.5075 | 1.2830 | | 1.3962 | 1.8486 | 2.6921 | 3.2106 | 3.2106 | 1.9884 | O4 | 2.7343 | 2.2221 | 1.3962 | | 0.9615 | 2.3260 | 3.5024 | 3.5024 | 3.2101 | H5 | 3.6787 | 2.9721 | 1.8486 | 0.9615 | | 3.2757 | 4.4211 | 4.4211 | 3.8300 | H6 | 1.0965 | 2.1596 | 2.6921 | 2.3260 | 3.2757 | | 1.7927 | 1.7927 | 3.1366 | H7 | 1.1027 | 2.1472 | 3.2106 | 3.5024 | 4.4211 | 1.7927 | | 1.7762 | 2.6022 | H8 | 1.1027 | 2.1472 | 3.2106 | 3.5024 | 4.4211 | 1.7927 | 1.7762 | | 2.6022 | H9 | 2.2488 | 1.0958 | 1.9884 | 3.2101 | 3.8300 | 3.1366 | 2.6022 | 2.6022 | |
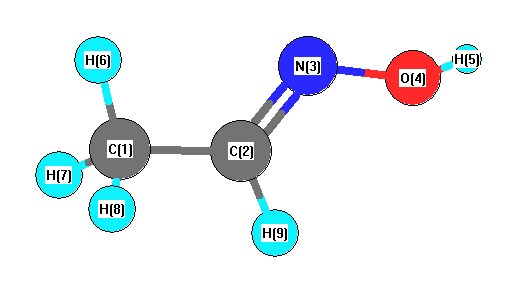 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.978 |
|
C1 |
C2 |
H9 |
118.844 |
| C2 |
C1 |
H6 |
111.259 |
|
C2 |
C1 |
H7 |
109.893 |
| C2 |
C1 |
H8 |
109.893 |
|
C2 |
N3 |
O4 |
112.002 |
| N3 |
C2 |
H9 |
113.177 |
|
N3 |
O4 |
H5 |
101.684 |
| H6 |
C1 |
H7 |
109.207 |
|
H6 |
C1 |
H8 |
109.207 |
| H7 |
C1 |
H8 |
107.293 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
