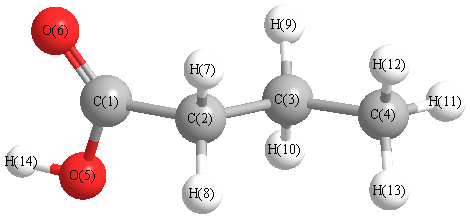return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Butanoic acid)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-306.886934
Energy at 298.15K-306.896385
HF Energy-305.905023
Nuclear repulsion energy237.885054
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3790 3636 60.68      
2 A 3151 3023 34.82      
3 A 3142 3014 29.65      
4 A 3135 3008 24.67      
5 A 3112 2986 0.47      
6 A 3077 2952 20.40      
7 A 3063 2939 10.81      
8 A 3053 2929 29.30      
9 A 1868 1793 284.69      
10 A 1508 1446 4.66      
11 A 1497 1436 6.61      
12 A 1494 1433 0.86      
13 A 1478 1418 8.99      
14 A 1429 1371 24.25      
15 A 1417 1359 29.95      
16 A 1392 1336 13.18      
17 A 1329 1275 1.76      
18 A 1296 1244 0.93      
19 A 1272 1220 57.37      
20 A 1232 1182 118.23      
21 A 1131 1085 3.50      
22 A 1100 1055 46.93      
23 A 1078 1035 14.30      
24 A 940 902 2.14      
25 A 897 861 1.28      
26 A 881 845 7.86      
27 A 759 729 11.09      
28 A 727 698 39.09      
29 A 623 598 71.90      
30 A 579 555 47.20      
31 A 433 415 2.15      
32 A 335 322 1.22      
33 A 254 244 0.02      
34 A 190 183 0.03      
35 A 96 92 0.19      
36 A 42 40 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 26401.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 25329.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.27552 0.06162 0.05461

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.194 -0.148 0.070
C2 -0.234 -0.429 0.493
C3 -1.267 0.263 -0.415
C4 -2.707 -0.057 0.006
O5 1.475 1.182 0.129
O6 2.008 -0.970 -0.288
H7 -0.370 -1.524 0.482
H8 -0.360 -0.074 1.533
H9 -1.103 -0.060 -1.461
H10 -1.096 1.355 -0.388
H11 -3.436 0.445 -0.655
H12 -2.900 -1.145 -0.038
H13 -2.900 0.279 1.042
H14 2.396 1.258 -0.171

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13 H14
C11.51572.54213.90231.36021.21122.12342.13582.76262.77734.72354.21534.22871.8657
C21.51571.54012.54752.37682.43481.10281.10672.17072.16873.51172.81112.81203.1951
C32.54211.54011.53362.94243.50202.19182.17531.10761.10522.18922.18922.18793.8042
C43.90232.54751.53364.36294.81122.79982.79982.17372.17771.10461.10591.10615.2729
O51.36022.37682.94244.36292.25583.29362.63013.27352.62815.02694.95814.55880.9721
O61.21122.43483.50204.81122.25582.55993.11903.44753.87975.63644.91755.23542.2651
H72.12341.10282.19182.79983.29362.55991.79092.54153.09383.81672.61083.15623.9773
H82.13581.10672.17532.79982.63013.11901.79093.08572.50483.81003.17292.61013.5040
H92.76262.17071.10762.17373.27353.44752.54153.08571.77612.51912.53663.09983.9561
H102.77732.16871.10522.17772.62813.87973.09382.50481.77612.52443.10272.54033.5008
H114.72353.51172.18921.10465.02695.63643.81673.81002.51912.52441.78751.78755.9085
H124.21532.81112.18921.10594.95814.91752.61083.17292.53663.10271.78751.78715.8179
H134.22872.81202.18791.10614.55885.23543.15622.61013.09982.54031.78751.78715.5205
H141.86573.19513.80425.27290.97212.26513.97733.50403.95613.50085.90855.81795.5205

picture of Butanoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.590 C1 C2 H7 107.320
C1 C2 H8 108.049 C1 O5 H14 105.030
C2 C1 O5 111.357 C2 C1 O6 126.111
C2 C3 C4 111.956 C2 C3 H9 109.051
C2 C3 H10 109.031 C3 C2 H7 110.981
C3 C2 H8 109.457 C3 C4 H11 111.120
C3 C4 H12 111.036 C3 C4 H13 110.921
C4 C3 H9 109.720 C4 C3 H10 110.172
O5 C1 O6 122.529 H7 C2 H8 108.303
H9 C3 H10 106.766 H11 C4 H12 107.925
H11 C4 H13 107.911 H12 C4 H13 107.782
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability