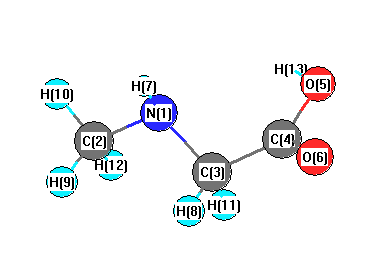
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3364 |
3370 |
0.75 |
|
|
|
| 2 |
A |
3183 |
3188 |
342.07 |
|
|
|
| 3 |
A |
3033 |
3037 |
22.75 |
|
|
|
| 4 |
A |
2992 |
2996 |
17.82 |
|
|
|
| 5 |
A |
2988 |
2993 |
20.74 |
|
|
|
| 6 |
A |
2938 |
2942 |
29.17 |
|
|
|
| 7 |
A |
2895 |
2900 |
88.45 |
|
|
|
| 8 |
A |
1791 |
1794 |
311.31 |
|
|
|
| 9 |
A |
1454 |
1456 |
9.72 |
|
|
|
| 10 |
A |
1433 |
1436 |
33.59 |
|
|
|
| 11 |
A |
1424 |
1426 |
21.05 |
|
|
|
| 12 |
A |
1411 |
1413 |
34.12 |
|
|
|
| 13 |
A |
1397 |
1399 |
60.84 |
|
|
|
| 14 |
A |
1387 |
1389 |
208.50 |
|
|
|
| 15 |
A |
1284 |
1286 |
5.46 |
|
|
|
| 16 |
A |
1234 |
1236 |
4.26 |
|
|
|
| 17 |
A |
1165 |
1167 |
28.52 |
|
|
|
| 18 |
A |
1120 |
1122 |
12.96 |
|
|
|
| 19 |
A |
1097 |
1098 |
41.87 |
|
|
|
| 20 |
A |
1083 |
1085 |
15.06 |
|
|
|
| 21 |
A |
970 |
972 |
19.34 |
|
|
|
| 22 |
A |
925 |
927 |
14.33 |
|
|
|
| 23 |
A |
894 |
895 |
43.26 |
|
|
|
| 24 |
A |
832 |
834 |
16.33 |
|
|
|
| 25 |
A |
765 |
766 |
54.58 |
|
|
|
| 26 |
A |
619 |
620 |
2.79 |
|
|
|
| 27 |
A |
557 |
558 |
5.03 |
|
|
|
| 28 |
A |
457 |
458 |
8.56 |
|
|
|
| 29 |
A |
367 |
368 |
4.52 |
|
|
|
| 30 |
A |
276 |
276 |
4.43 |
|
|
|
| 31 |
A |
202 |
203 |
2.23 |
|
|
|
| 32 |
A |
140 |
140 |
1.86 |
|
|
|
| 33 |
A |
78 |
78 |
5.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22875.9 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 22912.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.208 |
|
|
|
| 2 |
C |
0.066 |
|
|
|
| 3 |
C |
0.073 |
|
|
|
| 4 |
C |
0.070 |
|
|
|
| 5 |
O |
-0.129 |
|
|
|
| 6 |
O |
-0.198 |
|
|
|
| 7 |
H |
0.085 |
|
|
|
| 8 |
H |
0.022 |
|
|
|
| 9 |
H |
0.009 |
|
|
|
| 10 |
H |
0.017 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
H |
0.029 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.083 |
-0.158 |
-0.546 |
5.114 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.542 |
-0.563 |
-0.565 |
| y |
-0.563 |
-37.389 |
-0.634 |
| z |
-0.565 |
-0.634 |
-33.402 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.146 |
-0.563 |
-0.565 |
| y |
-0.563 |
0.582 |
-0.634 |
| z |
-0.565 |
-0.634 |
6.564 |
|
| Polar |
|---|
| 3z2-r2 | 13.127 |
|---|
| x2-y2 | -5.152 |
|---|
| xy | -0.563 |
|---|
| xz | -0.565 |
|---|
| yz | -0.634 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.085 |
0.349 |
0.033 |
| y |
0.349 |
7.051 |
0.042 |
| z |
0.033 |
0.042 |
5.391 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |