return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

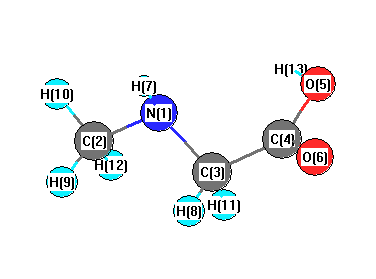
All results from a given calculation for CH3NHCH2COOH (Sarcosine)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-322.884176
Energy at 298.15K-322.893757
HF Energy-321.882459
Nuclear repulsion energy242.944038
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3626 3478 267.93      
2 A 3520 3377 2.91      
3 A 3157 3029 22.91      
4 A 3116 2989 17.27      
5 A 3112 2986 27.15      
6 A 3059 2935 20.50      
7 A 3023 2901 60.30      
8 A 1901 1824 311.02      
9 A 1522 1460 15.79      
10 A 1502 1441 12.53      
11 A 1496 1435 43.10      
12 A 1483 1423 49.52      
13 A 1468 1409 197.39      
14 A 1457 1397 117.04      
15 A 1361 1306 13.65      
16 A 1299 1246 6.89      
17 A 1264 1213 11.71      
18 A 1201 1152 27.56      
19 A 1165 1118 17.38      
20 A 1144 1097 13.91      
21 A 1012 971 17.38      
22 A 1000 960 13.44      
23 A 903 866 58.32      
24 A 897 861 14.52      
25 A 778 746 69.74      
26 A 660 633 3.03      
27 A 589 565 10.33      
28 A 477 458 9.78      
29 A 381 366 3.45      
30 A 278 267 2.91      
31 A 210 202 2.59      
32 A 134 128 2.32      
33 A 88 84 6.49      

Unscaled Zero Point Vibrational Energy (zpe) 24139.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23159.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.30142 0.06664 0.05712

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.197 0.285 -0.305
C2 2.547 -0.078 0.146
C3 0.178 -0.702 0.068
C4 -1.224 -0.074 0.023
O5 -1.210 1.270 0.118
O6 -2.238 -0.724 -0.055
H7 1.209 0.362 -1.326
H8 0.170 -1.626 -0.541
H9 2.876 -1.079 -0.203
H10 3.267 0.675 -0.217
H11 0.355 -1.003 1.118
H12 2.573 -0.072 1.250
H13 -0.257 1.501 0.127

Atom - Atom Distances (Å)
  N1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13
N11.46811.46712.46932.63483.58911.02432.18322.16572.10752.09602.10641.9440
C21.46812.45073.77223.99094.83212.03682.91851.11051.10322.56991.10443.2178
C31.46712.45071.53642.41142.41912.03381.10732.73773.39361.10672.74392.2457
C42.46933.77221.53641.34721.20692.81532.16114.22704.55862.13413.99001.8504
O52.63483.99092.41141.34722.24952.95993.27524.72384.52812.93474.17020.9800
O63.58914.83212.41911.20692.24953.83072.61735.12835.68182.85955.02702.9837
H71.02432.03682.03382.81532.95993.83072.37652.47302.35852.92652.94692.3575
H82.18322.91851.10732.16113.27522.61732.37652.78113.87181.78193.37583.2260
H92.16571.11052.73774.22704.72385.12832.47302.78111.79762.84721.79364.0721
H102.10751.10323.39364.55864.52815.68182.35853.87181.79763.61601.78653.6356
H112.09602.56991.10672.13412.93472.85952.92651.78192.84723.61602.40892.7614
H122.10641.10442.74393.99004.17025.02702.94693.37581.79361.78652.40893.4274
H131.94403.21782.24571.85040.98002.98372.35753.22604.07213.63562.76143.4274

picture of Sarcosine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 H9 113.533 N1 C2 H10 109.274
N1 C2 H12 109.112 N1 C3 C4 110.581
N1 C3 H8 115.290 N1 C3 H11 108.237
C2 N1 C3 113.218 C2 N1 H7 108.294
C3 N1 H7 108.128 C3 C4 O5 113.330
C3 C4 O6 123.279 C4 C3 H8 108.576
C4 C3 H11 106.562 C4 O5 H13 104.217
O5 C4 O6 123.363 H8 C3 H11 107.194
H9 C2 H10 108.591 H9 C2 H12 108.148
H10 C2 H12 108.039
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability