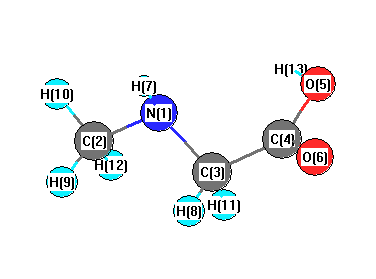
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3406 |
3386 |
3.11 |
|
|
|
| 2 |
A |
3121 |
3103 |
426.40 |
|
|
|
| 3 |
A |
3062 |
3044 |
15.29 |
|
|
|
| 4 |
A |
3019 |
3001 |
5.43 |
|
|
|
| 5 |
A |
3015 |
2998 |
26.05 |
|
|
|
| 6 |
A |
2961 |
2944 |
20.82 |
|
|
|
| 7 |
A |
2919 |
2902 |
88.98 |
|
|
|
| 8 |
A |
1821 |
1811 |
338.12 |
|
|
|
| 9 |
A |
1448 |
1440 |
12.39 |
|
|
|
| 10 |
A |
1432 |
1423 |
201.88 |
|
|
|
| 11 |
A |
1418 |
1410 |
26.05 |
|
|
|
| 12 |
A |
1409 |
1401 |
91.38 |
|
|
|
| 13 |
A |
1395 |
1387 |
26.79 |
|
|
|
| 14 |
A |
1379 |
1371 |
33.16 |
|
|
|
| 15 |
A |
1272 |
1264 |
5.83 |
|
|
|
| 16 |
A |
1231 |
1224 |
3.41 |
|
|
|
| 17 |
A |
1189 |
1182 |
26.17 |
|
|
|
| 18 |
A |
1141 |
1135 |
40.59 |
|
|
|
| 19 |
A |
1109 |
1102 |
11.31 |
|
|
|
| 20 |
A |
1086 |
1080 |
11.90 |
|
|
|
| 21 |
A |
979 |
974 |
24.71 |
|
|
|
| 22 |
A |
947 |
942 |
26.86 |
|
|
|
| 23 |
A |
919 |
913 |
23.35 |
|
|
|
| 24 |
A |
854 |
849 |
14.39 |
|
|
|
| 25 |
A |
772 |
768 |
54.96 |
|
|
|
| 26 |
A |
626 |
622 |
3.03 |
|
|
|
| 27 |
A |
558 |
554 |
4.37 |
|
|
|
| 28 |
A |
463 |
460 |
8.00 |
|
|
|
| 29 |
A |
375 |
373 |
6.00 |
|
|
|
| 30 |
A |
284 |
282 |
5.57 |
|
|
|
| 31 |
A |
204 |
203 |
2.14 |
|
|
|
| 32 |
A |
143 |
142 |
1.96 |
|
|
|
| 33 |
A |
76 |
75 |
5.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23014.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22881.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.246 |
|
|
|
| 2 |
C |
0.001 |
|
|
|
| 3 |
C |
0.016 |
|
|
|
| 4 |
C |
0.087 |
|
|
|
| 5 |
O |
-0.138 |
|
|
|
| 6 |
O |
-0.212 |
|
|
|
| 7 |
H |
0.105 |
|
|
|
| 8 |
H |
0.050 |
|
|
|
| 9 |
H |
0.033 |
|
|
|
| 10 |
H |
0.044 |
|
|
|
| 11 |
H |
0.070 |
|
|
|
| 12 |
H |
0.054 |
|
|
|
| 13 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.220 |
-0.236 |
-0.619 |
5.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.216 |
-0.493 |
-0.488 |
| y |
-0.493 |
-37.251 |
-0.609 |
| z |
-0.488 |
-0.609 |
-33.180 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.001 |
-0.493 |
-0.488 |
| y |
-0.493 |
0.447 |
-0.609 |
| z |
-0.488 |
-0.609 |
6.554 |
|
| Polar |
|---|
| 3z2-r2 | 13.108 |
|---|
| x2-y2 | -4.966 |
|---|
| xy | -0.493 |
|---|
| xz | -0.488 |
|---|
| yz | -0.609 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.999 |
0.345 |
0.025 |
| y |
0.345 |
6.969 |
0.039 |
| z |
0.025 |
0.039 |
5.359 |
<r2> (average value of r
2) Å
2
| <r2> |
194.251 |
| (<r2>)1/2 |
13.937 |