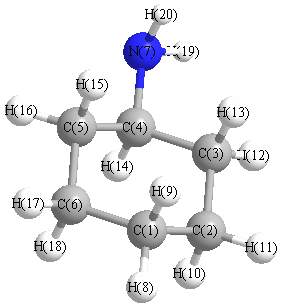
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3540 |
3417 |
0.37 |
|
|
|
| 2 |
A |
3460 |
3339 |
2.46 |
|
|
|
| 3 |
A |
3092 |
2984 |
41.16 |
|
|
|
| 4 |
A |
3083 |
2975 |
60.86 |
|
|
|
| 5 |
A |
3078 |
2970 |
42.43 |
|
|
|
| 6 |
A |
3077 |
2969 |
49.04 |
|
|
|
| 7 |
A |
3062 |
2955 |
58.19 |
|
|
|
| 8 |
A |
3025 |
2919 |
55.15 |
|
|
|
| 9 |
A |
3022 |
2916 |
14.98 |
|
|
|
| 10 |
A |
3020 |
2914 |
17.69 |
|
|
|
| 11 |
A |
3014 |
2908 |
53.66 |
|
|
|
| 12 |
A |
3004 |
2899 |
17.71 |
|
|
|
| 13 |
A |
2998 |
2893 |
3.01 |
|
|
|
| 14 |
A |
1641 |
1583 |
21.83 |
|
|
|
| 15 |
A |
1478 |
1426 |
2.05 |
|
|
|
| 16 |
A |
1459 |
1408 |
11.01 |
|
|
|
| 17 |
A |
1457 |
1406 |
5.30 |
|
|
|
| 18 |
A |
1454 |
1403 |
1.32 |
|
|
|
| 19 |
A |
1447 |
1396 |
0.08 |
|
|
|
| 20 |
A |
1414 |
1365 |
4.96 |
|
|
|
| 21 |
A |
1378 |
1330 |
3.14 |
|
|
|
| 22 |
A |
1374 |
1326 |
0.30 |
|
|
|
| 23 |
A |
1369 |
1321 |
3.84 |
|
|
|
| 24 |
A |
1358 |
1310 |
0.59 |
|
|
|
| 25 |
A |
1349 |
1302 |
3.11 |
|
|
|
| 26 |
A |
1315 |
1269 |
0.69 |
|
|
|
| 27 |
A |
1293 |
1248 |
0.81 |
|
|
|
| 28 |
A |
1277 |
1232 |
2.59 |
|
|
|
| 29 |
A |
1275 |
1230 |
2.12 |
|
|
|
| 30 |
A |
1221 |
1178 |
6.07 |
|
|
|
| 31 |
A |
1195 |
1153 |
1.46 |
|
|
|
| 32 |
A |
1143 |
1103 |
8.37 |
|
|
|
| 33 |
A |
1127 |
1088 |
6.60 |
|
|
|
| 34 |
A |
1098 |
1059 |
2.11 |
|
|
|
| 35 |
A |
1068 |
1031 |
0.42 |
|
|
|
| 36 |
A |
1054 |
1017 |
4.86 |
|
|
|
| 37 |
A |
1047 |
1010 |
0.33 |
|
|
|
| 38 |
A |
989 |
954 |
1.41 |
|
|
|
| 39 |
A |
944 |
911 |
45.92 |
|
|
|
| 40 |
A |
911 |
880 |
57.15 |
|
|
|
| 41 |
A |
895 |
863 |
24.59 |
|
|
|
| 42 |
A |
875 |
845 |
36.45 |
|
|
|
| 43 |
A |
861 |
831 |
0.64 |
|
|
|
| 44 |
A |
792 |
765 |
2.06 |
|
|
|
| 45 |
A |
788 |
761 |
0.22 |
|
|
|
| 46 |
A |
551 |
531 |
1.66 |
|
|
|
| 47 |
A |
459 |
443 |
0.85 |
|
|
|
| 48 |
A |
450 |
434 |
1.81 |
|
|
|
| 49 |
A |
406 |
392 |
0.10 |
|
|
|
| 50 |
A |
344 |
332 |
13.83 |
|
|
|
| 51 |
A |
333 |
321 |
1.28 |
|
|
|
| 52 |
A |
269 |
260 |
27.60 |
|
|
|
| 53 |
A |
228 |
220 |
5.04 |
|
|
|
| 54 |
A |
160 |
155 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41009.9 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 39574.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.050 |
|
|
|
| 2 |
C |
-0.047 |
|
|
|
| 3 |
C |
0.001 |
|
|
|
| 4 |
C |
-0.079 |
|
|
|
| 5 |
C |
-0.014 |
|
|
|
| 6 |
C |
-0.050 |
|
|
|
| 7 |
N |
-0.179 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
0.029 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.024 |
|
|
|
| 12 |
H |
0.030 |
|
|
|
| 13 |
H |
0.017 |
|
|
|
| 14 |
H |
0.022 |
|
|
|
| 15 |
H |
0.016 |
|
|
|
| 16 |
H |
0.015 |
|
|
|
| 17 |
H |
0.023 |
|
|
|
| 18 |
H |
0.029 |
|
|
|
| 19 |
H |
0.080 |
|
|
|
| 20 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.419 |
1.020 |
0.441 |
1.188 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.096 |
-3.675 |
-1.519 |
| y |
-3.675 |
-45.949 |
-0.560 |
| z |
-1.519 |
-0.560 |
-44.189 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.026 |
-3.675 |
-1.519 |
| y |
-3.675 |
-0.307 |
-0.560 |
| z |
-1.519 |
-0.560 |
2.333 |
|
| Polar |
|---|
| 3z2-r2 | 4.666 |
|---|
| x2-y2 | -1.146 |
|---|
| xy | -3.675 |
|---|
| xz | -1.519 |
|---|
| yz | -0.560 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.730 |
-0.205 |
-0.129 |
| y |
-0.205 |
10.781 |
-0.094 |
| z |
-0.129 |
-0.094 |
9.692 |
<r2> (average value of r
2) Å
2
| <r2> |
232.650 |
| (<r2>)1/2 |
15.253 |