return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (cyclohexanamine)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-290.358941
Energy at 298.15K-290.375468
HF Energy-289.252510
Nuclear repulsion energy328.610524
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3566 3412 0.83      
2 A 3483 3332 2.21      
3 A 3122 2987 43.51      
4 A 3114 2979 56.45      
5 A 3108 2974 35.18      
6 A 3107 2973 49.19      
7 A 3093 2959 54.03      
8 A 3054 2921 44.43      
9 A 3052 2919 15.60      
10 A 3051 2919 26.21      
11 A 3045 2913 45.34      
12 A 3035 2904 11.73      
13 A 3034 2903 9.37      
14 A 1667 1595 21.20      
15 A 1508 1443 1.42      
16 A 1492 1427 8.13      
17 A 1489 1425 4.01      
18 A 1486 1421 1.67      
19 A 1479 1415 0.03      
20 A 1447 1384 4.63      
21 A 1407 1346 4.69      
22 A 1403 1342 0.69      
23 A 1400 1340 2.13      
24 A 1383 1323 0.97      
25 A 1374 1314 4.41      
26 A 1340 1282 1.08      
27 A 1311 1254 0.93      
28 A 1294 1238 0.72      
29 A 1293 1237 3.46      
30 A 1240 1187 7.12      
31 A 1218 1165 2.07      
32 A 1164 1114 4.64      
33 A 1154 1104 9.00      
34 A 1111 1063 2.11      
35 A 1082 1035 0.90      
36 A 1070 1024 3.99      
37 A 1060 1014 0.72      
38 A 1002 959 1.60      
39 A 977 935 100.64      
40 A 936 895 27.39      
41 A 908 869 6.04      
42 A 893 854 10.10      
43 A 869 831 0.82      
44 A 803 768 1.84      
45 A 800 765 0.13      
46 A 555 531 1.14      
47 A 464 444 1.30      
48 A 452 432 1.81      
49 A 408 390 0.09      
50 A 348 333 15.23      
51 A 337 322 1.39      
52 A 277 265 29.59      
53 A 230 220 2.99      
54 A 161 154 0.35      

Unscaled Zero Point Vibrational Energy (zpe) 41575.2 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 39775.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.14077 0.07304 0.05305

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.884 0.016 0.293
C2 1.187 -1.256 -0.219
C3 -0.303 -1.271 0.163
C4 -1.035 -0.015 -0.332
C5 -0.326 1.254 0.178
C6 1.163 1.279 -0.207
N7 -2.457 -0.110 0.031
H8 2.944 0.028 -0.023
H9 1.881 0.012 1.402
H10 1.279 -1.301 -1.323
H11 1.691 -2.157 0.177
H12 -0.809 -2.167 -0.240
H13 -0.396 -1.320 1.268
H14 -0.989 -0.011 -1.442
H15 -0.422 1.287 1.284
H16 -0.843 2.152 -0.213
H17 1.648 2.187 0.196
H18 1.252 1.335 -1.311
H19 -2.929 0.741 -0.290
H20 -2.521 -0.063 1.054

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.53832.54092.98582.53601.53814.35101.10581.10912.17082.18473.50712.81733.35572.81303.50072.18602.17094.90244.4712
C21.53831.53782.54772.95782.53543.82822.18462.17171.10891.10562.19382.17352.78863.36343.96693.49872.81274.57554.0980
C32.54091.53781.53612.52572.96492.45113.50152.81972.17062.18121.10451.11082.15222.79573.48643.97113.37393.33962.6788
C42.98582.54771.53611.54122.55421.47053.99123.39342.82703.50382.16502.16161.11022.16462.17953.51182.83092.03972.0324
C52.53602.95782.52571.54121.53842.53463.49802.81343.37053.96273.47992.79642.15951.11061.10752.18402.17152.69422.7054
C61.53812.53542.96492.55421.53843.88482.18422.17102.81363.49773.97023.37122.79602.17602.18741.10571.10904.12834.1187
N74.35103.82822.45111.47052.53463.88485.40284.55174.14874.62762.64942.69122.08182.76882.79014.70754.20081.02471.0260
H81.10582.18463.50153.99123.49802.18425.40281.77772.49512.52664.35243.82644.18023.82344.34572.52752.49595.92205.5706
H91.10912.17172.81973.39342.81342.17104.55171.77773.08412.49753.83112.64164.04012.63513.82262.49863.08395.15154.4169
H102.17081.10892.17062.82703.37052.81364.14872.49513.08411.77582.50653.08632.61154.04814.20243.82252.63624.79034.6503
H112.18471.10562.18123.50383.96273.49774.62762.52662.49751.77582.53412.49943.79504.18875.01394.34443.82145.47344.7846
H123.50712.19381.10452.16503.47993.97022.64944.35243.83112.50652.53411.77852.47413.79464.31915.01834.20193.59883.0052
H132.81732.17351.11082.16162.79643.37122.69123.82642.64163.08632.49941.77853.06722.60683.80144.19904.05233.61832.4781
H143.35572.78862.15221.11022.15952.79602.08184.18024.04012.61153.79502.47413.06723.07172.49223.80362.61712.37842.9287
H152.81303.36342.79572.16461.11062.17602.76883.82342.63514.04814.18873.79462.60683.07171.78012.50593.08853.01102.5067
H163.50073.96693.48642.17951.10752.18742.79014.34573.82264.20245.01394.31913.80142.49221.78012.52472.50222.52033.0544
H172.18603.49873.97113.51182.18401.10574.70752.52752.49863.82254.34445.01834.19903.80362.50592.52471.77624.82534.8150
H182.17092.81273.37392.83092.17151.10904.20082.49593.08392.63623.82144.20194.05232.61713.08852.50221.77624.34484.6673
H194.90244.57553.33962.03972.69424.12831.02475.92205.15154.79035.47343.59883.61832.37843.01102.52034.82534.34481.6183
H204.47124.09802.67882.03242.70544.11871.02605.57064.41694.65034.78463.00522.47812.92872.50673.05444.81504.66731.6183

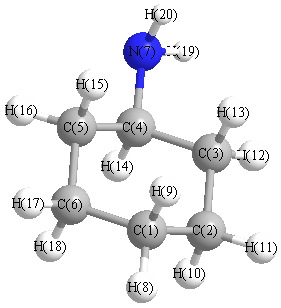
picture of cyclohexanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.382 C1 C2 H10 109.098
C1 C2 H11 110.370 C1 C6 C5 111.038
C1 C6 H17 110.486 C1 C6 H18 109.116
C2 C1 C6 111.006 C2 C1 H8 110.355
C2 C1 H9 109.158 C2 C3 C4 111.952
C2 C3 H12 111.190 C2 C3 H13 109.232
C3 C2 H10 109.114 C3 C2 H11 110.125
C3 C4 C5 110.324 C3 C4 N7 109.208
C3 C4 H14 107.757 C4 C3 H12 109.058
C4 C3 H13 108.438 C4 C5 C6 112.073
C4 C5 H15 108.333 C4 C5 H16 109.660
C4 N7 H19 108.337 C4 N7 H20 107.669
C5 C4 N7 114.593 C5 C4 H14 107.965
C5 C6 H17 110.303 C5 C6 H18 109.140
C6 C1 H8 110.336 C6 C1 H9 109.117
C6 C5 H15 109.397 C6 C5 H16 110.467
N7 C4 H14 106.717 H8 C1 H9 106.761
H10 C2 H11 106.622 H12 C3 H13 106.802
H15 C5 H16 106.744 H17 C6 H18 106.641
H19 N7 H20 104.211
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability