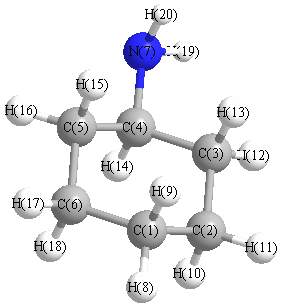
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3419 |
3399 |
2.04 |
|
|
|
| 2 |
A |
3339 |
3319 |
5.96 |
|
|
|
| 3 |
A |
3015 |
2998 |
38.56 |
|
|
|
| 4 |
A |
3007 |
2990 |
55.36 |
|
|
|
| 5 |
A |
3004 |
2986 |
46.68 |
|
|
|
| 6 |
A |
3001 |
2984 |
45.64 |
|
|
|
| 7 |
A |
2985 |
2968 |
57.13 |
|
|
|
| 8 |
A |
2950 |
2933 |
54.87 |
|
|
|
| 9 |
A |
2947 |
2930 |
12.79 |
|
|
|
| 10 |
A |
2944 |
2926 |
16.49 |
|
|
|
| 11 |
A |
2936 |
2919 |
54.30 |
|
|
|
| 12 |
A |
2927 |
2910 |
16.29 |
|
|
|
| 13 |
A |
2917 |
2901 |
3.92 |
|
|
|
| 14 |
A |
1591 |
1581 |
18.03 |
|
|
|
| 15 |
A |
1427 |
1419 |
1.79 |
|
|
|
| 16 |
A |
1409 |
1401 |
10.78 |
|
|
|
| 17 |
A |
1406 |
1398 |
4.24 |
|
|
|
| 18 |
A |
1404 |
1396 |
1.15 |
|
|
|
| 19 |
A |
1397 |
1389 |
0.07 |
|
|
|
| 20 |
A |
1365 |
1358 |
3.95 |
|
|
|
| 21 |
A |
1329 |
1321 |
3.47 |
|
|
|
| 22 |
A |
1325 |
1318 |
0.04 |
|
|
|
| 23 |
A |
1319 |
1312 |
3.47 |
|
|
|
| 24 |
A |
1312 |
1305 |
1.19 |
|
|
|
| 25 |
A |
1305 |
1298 |
4.09 |
|
|
|
| 26 |
A |
1271 |
1264 |
0.68 |
|
|
|
| 27 |
A |
1253 |
1246 |
1.01 |
|
|
|
| 28 |
A |
1238 |
1230 |
3.31 |
|
|
|
| 29 |
A |
1233 |
1226 |
1.73 |
|
|
|
| 30 |
A |
1182 |
1175 |
6.43 |
|
|
|
| 31 |
A |
1158 |
1151 |
1.48 |
|
|
|
| 32 |
A |
1107 |
1101 |
5.26 |
|
|
|
| 33 |
A |
1092 |
1086 |
4.82 |
|
|
|
| 34 |
A |
1067 |
1061 |
3.01 |
|
|
|
| 35 |
A |
1036 |
1030 |
0.24 |
|
|
|
| 36 |
A |
1024 |
1018 |
4.26 |
|
|
|
| 37 |
A |
1019 |
1013 |
0.45 |
|
|
|
| 38 |
A |
960 |
955 |
2.11 |
|
|
|
| 39 |
A |
920 |
914 |
55.39 |
|
|
|
| 40 |
A |
887 |
882 |
60.46 |
|
|
|
| 41 |
A |
871 |
866 |
18.06 |
|
|
|
| 42 |
A |
852 |
847 |
21.80 |
|
|
|
| 43 |
A |
838 |
834 |
0.51 |
|
|
|
| 44 |
A |
771 |
766 |
1.76 |
|
|
|
| 45 |
A |
765 |
761 |
0.16 |
|
|
|
| 46 |
A |
537 |
534 |
1.57 |
|
|
|
| 47 |
A |
450 |
447 |
0.77 |
|
|
|
| 48 |
A |
439 |
436 |
1.80 |
|
|
|
| 49 |
A |
396 |
393 |
0.08 |
|
|
|
| 50 |
A |
336 |
334 |
13.63 |
|
|
|
| 51 |
A |
327 |
325 |
1.46 |
|
|
|
| 52 |
A |
267 |
265 |
24.67 |
|
|
|
| 53 |
A |
224 |
223 |
4.70 |
|
|
|
| 54 |
A |
157 |
157 |
0.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39827.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 39596.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.024 |
|
|
|
| 2 |
C |
-0.020 |
|
|
|
| 3 |
C |
0.027 |
|
|
|
| 4 |
C |
-0.076 |
|
|
|
| 5 |
C |
0.012 |
|
|
|
| 6 |
C |
-0.023 |
|
|
|
| 7 |
N |
-0.139 |
|
|
|
| 8 |
H |
0.008 |
|
|
|
| 9 |
H |
0.016 |
|
|
|
| 10 |
H |
0.017 |
|
|
|
| 11 |
H |
0.009 |
|
|
|
| 12 |
H |
0.013 |
|
|
|
| 13 |
H |
0.005 |
|
|
|
| 14 |
H |
0.008 |
|
|
|
| 15 |
H |
0.003 |
|
|
|
| 16 |
H |
-0.001 |
|
|
|
| 17 |
H |
0.008 |
|
|
|
| 18 |
H |
0.017 |
|
|
|
| 19 |
H |
0.069 |
|
|
|
| 20 |
H |
0.069 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.399 |
0.985 |
0.429 |
1.147 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.446 |
-3.588 |
-1.498 |
| y |
-3.588 |
-46.382 |
-0.536 |
| z |
-1.498 |
-0.536 |
-44.748 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.881 |
-3.588 |
-1.498 |
| y |
-3.588 |
-0.285 |
-0.536 |
| z |
-1.498 |
-0.536 |
2.166 |
|
| Polar |
|---|
| 3z2-r2 | 4.332 |
|---|
| x2-y2 | -1.064 |
|---|
| xy | -3.588 |
|---|
| xz | -1.498 |
|---|
| yz | -0.536 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.287 |
-0.216 |
-0.138 |
| y |
-0.216 |
11.244 |
-0.091 |
| z |
-0.138 |
-0.091 |
10.026 |
<r2> (average value of r
2) Å
2
| <r2> |
234.869 |
| (<r2>)1/2 |
15.325 |