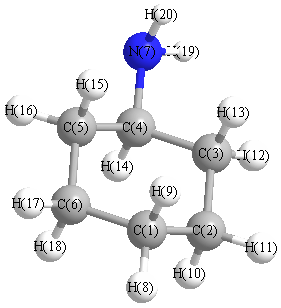
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3775 |
3249 |
0.14 |
|
|
|
| 2 |
A |
3698 |
3183 |
0.38 |
|
|
|
| 3 |
A |
3229 |
2779 |
64.70 |
|
|
|
| 4 |
A |
3214 |
2766 |
109.61 |
|
|
|
| 5 |
A |
3208 |
2761 |
74.29 |
|
|
|
| 6 |
A |
3207 |
2760 |
66.20 |
|
|
|
| 7 |
A |
3193 |
2748 |
83.52 |
|
|
|
| 8 |
A |
3160 |
2720 |
83.80 |
|
|
|
| 9 |
A |
3156 |
2717 |
19.76 |
|
|
|
| 10 |
A |
3156 |
2716 |
15.34 |
|
|
|
| 11 |
A |
3155 |
2715 |
17.30 |
|
|
|
| 12 |
A |
3144 |
2706 |
21.20 |
|
|
|
| 13 |
A |
3140 |
2703 |
14.44 |
|
|
|
| 14 |
A |
1785 |
1537 |
28.72 |
|
|
|
| 15 |
A |
1613 |
1389 |
1.75 |
|
|
|
| 16 |
A |
1595 |
1373 |
5.72 |
|
|
|
| 17 |
A |
1593 |
1371 |
5.13 |
|
|
|
| 18 |
A |
1588 |
1367 |
2.86 |
|
|
|
| 19 |
A |
1581 |
1361 |
0.06 |
|
|
|
| 20 |
A |
1550 |
1334 |
6.90 |
|
|
|
| 21 |
A |
1510 |
1300 |
5.09 |
|
|
|
| 22 |
A |
1505 |
1295 |
0.59 |
|
|
|
| 23 |
A |
1501 |
1292 |
1.68 |
|
|
|
| 24 |
A |
1481 |
1275 |
1.52 |
|
|
|
| 25 |
A |
1471 |
1266 |
4.13 |
|
|
|
| 26 |
A |
1436 |
1236 |
1.30 |
|
|
|
| 27 |
A |
1399 |
1204 |
0.83 |
|
|
|
| 28 |
A |
1382 |
1189 |
0.92 |
|
|
|
| 29 |
A |
1381 |
1189 |
3.16 |
|
|
|
| 30 |
A |
1322 |
1138 |
6.90 |
|
|
|
| 31 |
A |
1295 |
1115 |
2.40 |
|
|
|
| 32 |
A |
1224 |
1053 |
20.10 |
|
|
|
| 33 |
A |
1210 |
1041 |
7.34 |
|
|
|
| 34 |
A |
1164 |
1002 |
0.33 |
|
|
|
| 35 |
A |
1150 |
990 |
1.25 |
|
|
|
| 36 |
A |
1125 |
968 |
9.60 |
|
|
|
| 37 |
A |
1107 |
952 |
1.20 |
|
|
|
| 38 |
A |
1061 |
913 |
2.81 |
|
|
|
| 39 |
A |
1010 |
869 |
75.45 |
|
|
|
| 40 |
A |
988 |
850 |
50.77 |
|
|
|
| 41 |
A |
951 |
819 |
9.07 |
|
|
|
| 42 |
A |
940 |
809 |
30.86 |
|
|
|
| 43 |
A |
905 |
779 |
3.08 |
|
|
|
| 44 |
A |
852 |
733 |
0.24 |
|
|
|
| 45 |
A |
838 |
721 |
2.04 |
|
|
|
| 46 |
A |
591 |
508 |
1.24 |
|
|
|
| 47 |
A |
492 |
424 |
1.01 |
|
|
|
| 48 |
A |
483 |
415 |
1.54 |
|
|
|
| 49 |
A |
436 |
376 |
0.16 |
|
|
|
| 50 |
A |
370 |
319 |
13.43 |
|
|
|
| 51 |
A |
356 |
307 |
0.89 |
|
|
|
| 52 |
A |
287 |
247 |
34.37 |
|
|
|
| 53 |
A |
243 |
209 |
3.59 |
|
|
|
| 54 |
A |
170 |
146 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43688.8 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 37602.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.038 |
|
|
|
| 2 |
C |
-0.037 |
|
|
|
| 3 |
C |
0.012 |
|
|
|
| 4 |
C |
-0.019 |
|
|
|
| 5 |
C |
-0.003 |
|
|
|
| 6 |
C |
-0.040 |
|
|
|
| 7 |
N |
-0.248 |
|
|
|
| 8 |
H |
0.021 |
|
|
|
| 9 |
H |
0.021 |
|
|
|
| 10 |
H |
0.022 |
|
|
|
| 11 |
H |
0.022 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.010 |
|
|
|
| 14 |
H |
0.017 |
|
|
|
| 15 |
H |
0.010 |
|
|
|
| 16 |
H |
0.013 |
|
|
|
| 17 |
H |
0.021 |
|
|
|
| 18 |
H |
0.022 |
|
|
|
| 19 |
H |
0.083 |
|
|
|
| 20 |
H |
0.082 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.465 |
1.073 |
0.438 |
1.249 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.178 |
-3.830 |
-1.494 |
| y |
-3.830 |
-46.771 |
-0.584 |
| z |
-1.494 |
-0.584 |
-44.686 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.450 |
-3.830 |
-1.494 |
| y |
-3.830 |
-0.339 |
-0.584 |
| z |
-1.494 |
-0.584 |
2.789 |
|
| Polar |
|---|
| 3z2-r2 | 5.578 |
|---|
| x2-y2 | -1.407 |
|---|
| xy | -3.830 |
|---|
| xz | -1.494 |
|---|
| yz | -0.584 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
232.486 |
| (<r2>)1/2 |
15.247 |