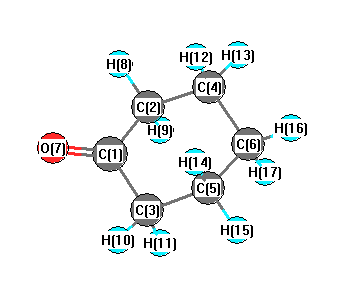
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3046 |
3028 |
14.50 |
|
|
|
| 2 |
A' |
3015 |
2997 |
42.14 |
|
|
|
| 3 |
A' |
3010 |
2992 |
38.37 |
|
|
|
| 4 |
A' |
2961 |
2944 |
43.34 |
|
|
|
| 5 |
A' |
2949 |
2932 |
16.38 |
|
|
|
| 6 |
A' |
2948 |
2931 |
12.47 |
|
|
|
| 7 |
A' |
1751 |
1741 |
177.28 |
|
|
|
| 8 |
A' |
1424 |
1416 |
0.79 |
|
|
|
| 9 |
A' |
1407 |
1399 |
8.63 |
|
|
|
| 10 |
A' |
1385 |
1377 |
7.51 |
|
|
|
| 11 |
A' |
1321 |
1313 |
7.80 |
|
|
|
| 12 |
A' |
1286 |
1279 |
2.69 |
|
|
|
| 13 |
A' |
1225 |
1218 |
0.20 |
|
|
|
| 14 |
A' |
1194 |
1187 |
0.98 |
|
|
|
| 15 |
A' |
1086 |
1080 |
0.31 |
|
|
|
| 16 |
A' |
1008 |
1003 |
3.38 |
|
|
|
| 17 |
A' |
973 |
968 |
0.90 |
|
|
|
| 18 |
A' |
843 |
839 |
3.50 |
|
|
|
| 19 |
A' |
836 |
831 |
0.63 |
|
|
|
| 20 |
A' |
736 |
732 |
4.03 |
|
|
|
| 21 |
A' |
639 |
635 |
1.41 |
|
|
|
| 22 |
A' |
461 |
459 |
0.23 |
|
|
|
| 23 |
A' |
395 |
393 |
0.92 |
|
|
|
| 24 |
A' |
299 |
297 |
0.09 |
|
|
|
| 25 |
A' |
94 |
93 |
1.74 |
|
|
|
| 26 |
A" |
3045 |
3027 |
19.49 |
|
|
|
| 27 |
A" |
3010 |
2993 |
32.16 |
|
|
|
| 28 |
A" |
2959 |
2942 |
19.90 |
|
|
|
| 29 |
A" |
2943 |
2926 |
2.23 |
|
|
|
| 30 |
A" |
1407 |
1399 |
5.99 |
|
|
|
| 31 |
A" |
1377 |
1369 |
7.58 |
|
|
|
| 32 |
A" |
1328 |
1320 |
1.83 |
|
|
|
| 33 |
A" |
1307 |
1300 |
12.00 |
|
|
|
| 34 |
A" |
1283 |
1275 |
16.79 |
|
|
|
| 35 |
A" |
1237 |
1229 |
0.18 |
|
|
|
| 36 |
A" |
1189 |
1183 |
21.93 |
|
|
|
| 37 |
A" |
1105 |
1099 |
17.34 |
|
|
|
| 38 |
A" |
1063 |
1057 |
5.23 |
|
|
|
| 39 |
A" |
1020 |
1014 |
4.57 |
|
|
|
| 40 |
A" |
901 |
896 |
7.68 |
|
|
|
| 41 |
A" |
870 |
865 |
0.10 |
|
|
|
| 42 |
A" |
729 |
725 |
5.31 |
|
|
|
| 43 |
A" |
475 |
472 |
8.01 |
|
|
|
| 44 |
A" |
400 |
398 |
2.34 |
|
|
|
| 45 |
A" |
171 |
170 |
0.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32054.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31868.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.013 |
|
|
|
| 2 |
C |
-0.001 |
|
|
|
| 3 |
C |
-0.001 |
|
|
|
| 4 |
C |
-0.033 |
|
|
|
| 5 |
C |
-0.033 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
O |
-0.181 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
0.042 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
H |
0.030 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
| 15 |
H |
0.020 |
|
|
|
| 16 |
H |
0.018 |
|
|
|
| 17 |
H |
0.020 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.863 |
-2.172 |
0.000 |
2.862 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.026 |
4.228 |
0.000 |
| y |
4.228 |
-47.905 |
0.000 |
| z |
0.000 |
0.000 |
-41.848 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.149 |
4.228 |
0.000 |
| y |
4.228 |
-4.469 |
0.000 |
| z |
0.000 |
0.000 |
4.618 |
|
| Polar |
|---|
| 3z2-r2 | 9.236 |
|---|
| x2-y2 | 2.880 |
|---|
| xy | 4.228 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.137 |
-1.128 |
0.000 |
| y |
-1.128 |
10.446 |
0.000 |
| z |
0.000 |
0.000 |
10.521 |
<r2> (average value of r
2) Å
2
| <r2> |
211.197 |
| (<r2>)1/2 |
14.533 |