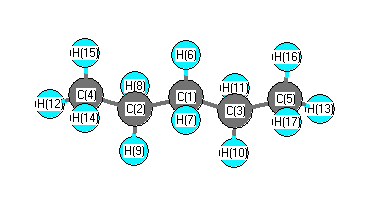
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3050 |
3032 |
38.45 |
|
|
|
| 2 |
A1 |
2965 |
2947 |
25.64 |
|
|
|
| 3 |
A1 |
2954 |
2937 |
85.47 |
|
|
|
| 4 |
A1 |
2936 |
2919 |
8.43 |
|
|
|
| 5 |
A1 |
1437 |
1429 |
5.14 |
|
|
|
| 6 |
A1 |
1421 |
1413 |
0.44 |
|
|
|
| 7 |
A1 |
1410 |
1402 |
0.00 |
|
|
|
| 8 |
A1 |
1350 |
1342 |
1.50 |
|
|
|
| 9 |
A1 |
1308 |
1301 |
0.03 |
|
|
|
| 10 |
A1 |
1126 |
1120 |
1.08 |
|
|
|
| 11 |
A1 |
1045 |
1038 |
0.72 |
|
|
|
| 12 |
A1 |
854 |
849 |
1.40 |
|
|
|
| 13 |
A1 |
392 |
389 |
0.00 |
|
|
|
| 14 |
A1 |
171 |
170 |
0.03 |
|
|
|
| 15 |
A2 |
3040 |
3023 |
0.00 |
|
|
|
| 16 |
A2 |
2984 |
2967 |
0.00 |
|
|
|
| 17 |
A2 |
1421 |
1413 |
0.00 |
|
|
|
| 18 |
A2 |
1279 |
1271 |
0.00 |
|
|
|
| 19 |
A2 |
1213 |
1206 |
0.00 |
|
|
|
| 20 |
A2 |
956 |
950 |
0.00 |
|
|
|
| 21 |
A2 |
745 |
740 |
0.00 |
|
|
|
| 22 |
A2 |
251 |
249 |
0.00 |
|
|
|
| 23 |
A2 |
117 |
116 |
0.00 |
|
|
|
| 24 |
B1 |
3041 |
3024 |
91.69 |
|
|
|
| 25 |
B1 |
2999 |
2982 |
56.85 |
|
|
|
| 26 |
B1 |
2964 |
2947 |
0.44 |
|
|
|
| 27 |
B1 |
1420 |
1411 |
12.90 |
|
|
|
| 28 |
B1 |
1275 |
1267 |
0.44 |
|
|
|
| 29 |
B1 |
1154 |
1148 |
0.22 |
|
|
|
| 30 |
B1 |
837 |
832 |
1.79 |
|
|
|
| 31 |
B1 |
722 |
717 |
4.99 |
|
|
|
| 32 |
B1 |
244 |
242 |
0.00 |
|
|
|
| 33 |
B1 |
117 |
116 |
0.00 |
|
|
|
| 34 |
B2 |
3050 |
3032 |
27.37 |
|
|
|
| 35 |
B2 |
2964 |
2947 |
44.22 |
|
|
|
| 36 |
B2 |
2948 |
2931 |
0.10 |
|
|
|
| 37 |
B2 |
1430 |
1422 |
1.09 |
|
|
|
| 38 |
B2 |
1411 |
1403 |
0.78 |
|
|
|
| 39 |
B2 |
1349 |
1341 |
2.67 |
|
|
|
| 40 |
B2 |
1344 |
1337 |
0.29 |
|
|
|
| 41 |
B2 |
1231 |
1224 |
6.40 |
|
|
|
| 42 |
B2 |
1068 |
1062 |
0.38 |
|
|
|
| 43 |
B2 |
1014 |
1008 |
0.81 |
|
|
|
| 44 |
B2 |
915 |
909 |
2.94 |
|
|
|
| 45 |
B2 |
383 |
381 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34149.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33951.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.043 |
|
|
|
| 2 |
C |
-0.070 |
|
|
|
| 3 |
C |
-0.070 |
|
|
|
| 4 |
C |
-0.047 |
|
|
|
| 5 |
C |
-0.047 |
|
|
|
| 6 |
H |
0.015 |
|
|
|
| 7 |
H |
0.015 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.019 |
|
|
|
| 10 |
H |
0.019 |
|
|
|
| 11 |
H |
0.019 |
|
|
|
| 12 |
H |
0.025 |
|
|
|
| 13 |
H |
0.025 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
| 15 |
H |
0.030 |
|
|
|
| 16 |
H |
0.030 |
|
|
|
| 17 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.064 |
0.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.656 |
0.000 |
0.000 |
| y |
0.000 |
-35.744 |
0.000 |
| z |
0.000 |
0.000 |
-35.409 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.921 |
0.000 |
0.000 |
| y |
0.000 |
-0.712 |
0.000 |
| z |
0.000 |
0.000 |
-0.209 |
|
| Polar |
|---|
| 3z2-r2 | -0.418 |
|---|
| x2-y2 | 1.088 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.978 |
0.000 |
0.000 |
| y |
0.000 |
10.678 |
0.000 |
| z |
0.000 |
0.000 |
8.397 |
<r2> (average value of r
2) Å
2
| <r2> |
203.243 |
| (<r2>)1/2 |
14.256 |