Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -195.930721 |
| Energy at 298.15K | -195.941000 |
| HF Energy | -195.154258 |
| Nuclear repulsion energy | 172.110655 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3256 |
3124 |
16.74 |
|
|
|
| 2 |
A |
3166 |
3037 |
4.07 |
|
|
|
| 3 |
A |
3151 |
3023 |
25.31 |
|
|
|
| 4 |
A |
3145 |
3017 |
35.35 |
|
|
|
| 5 |
A |
3132 |
3005 |
38.10 |
|
|
|
| 6 |
A |
3096 |
2970 |
31.06 |
|
|
|
| 7 |
A |
3088 |
2962 |
24.75 |
|
|
|
| 8 |
A |
3055 |
2931 |
11.40 |
|
|
|
| 9 |
A |
3049 |
2925 |
46.60 |
|
|
|
| 10 |
A |
3041 |
2917 |
23.14 |
|
|
|
| 11 |
A |
1725 |
1655 |
5.25 |
|
|
|
| 12 |
A |
1505 |
1444 |
2.15 |
|
|
|
| 13 |
A |
1497 |
1436 |
7.77 |
|
|
|
| 14 |
A |
1489 |
1429 |
0.30 |
|
|
|
| 15 |
A |
1480 |
1420 |
4.92 |
|
|
|
| 16 |
A |
1457 |
1398 |
0.80 |
|
|
|
| 17 |
A |
1425 |
1367 |
3.63 |
|
|
|
| 18 |
A |
1383 |
1327 |
2.57 |
|
|
|
| 19 |
A |
1364 |
1309 |
2.49 |
|
|
|
| 20 |
A |
1318 |
1264 |
1.33 |
|
|
|
| 21 |
A |
1293 |
1241 |
0.38 |
|
|
|
| 22 |
A |
1262 |
1211 |
1.42 |
|
|
|
| 23 |
A |
1199 |
1150 |
0.33 |
|
|
|
| 24 |
A |
1113 |
1067 |
1.29 |
|
|
|
| 25 |
A |
1092 |
1048 |
4.05 |
|
|
|
| 26 |
A |
1023 |
982 |
8.98 |
|
|
|
| 27 |
A |
1015 |
973 |
4.58 |
|
|
|
| 28 |
A |
947 |
908 |
0.86 |
|
|
|
| 29 |
A |
929 |
892 |
30.90 |
|
|
|
| 30 |
A |
889 |
852 |
0.10 |
|
|
|
| 31 |
A |
877 |
841 |
1.58 |
|
|
|
| 32 |
A |
778 |
746 |
2.25 |
|
|
|
| 33 |
A |
638 |
612 |
6.89 |
|
|
|
| 34 |
A |
434 |
417 |
0.32 |
|
|
|
| 35 |
A |
391 |
375 |
1.10 |
|
|
|
| 36 |
A |
301 |
289 |
0.21 |
|
|
|
| 37 |
A |
230 |
221 |
0.03 |
|
|
|
| 38 |
A |
121 |
116 |
0.03 |
|
|
|
| 39 |
A |
90 |
87 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30219.3 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 28992.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.114 |
0.567 |
0.124 |
| C2 |
-1.183 |
-0.250 |
-0.408 |
| C3 |
-0.061 |
-0.907 |
0.365 |
| C4 |
1.337 |
-0.521 |
-0.158 |
| C5 |
1.660 |
0.969 |
0.024 |
| H6 |
-2.906 |
1.012 |
-0.491 |
| H7 |
-2.114 |
0.813 |
1.194 |
| H8 |
-1.219 |
-0.468 |
-1.487 |
| H9 |
-0.176 |
-2.007 |
0.298 |
| H10 |
-0.146 |
-0.640 |
1.437 |
| H11 |
2.096 |
-1.131 |
0.368 |
| H12 |
1.410 |
-0.791 |
-1.231 |
| H13 |
2.673 |
1.206 |
-0.348 |
| H14 |
0.937 |
1.604 |
-0.519 |
| H15 |
1.617 |
1.251 |
1.093 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3477 | 2.5388 | 3.6300 | 3.7967 | 1.0965 | 1.0982 | 2.1136 | 3.2273 | 2.6566 | 4.5470 | 4.0124 | 4.8527 | 3.2865 | 3.9149 |
C2 | 1.3477 | | 1.5126 | 2.5475 | 3.1231 | 2.1366 | 2.1354 | 1.1018 | 2.1453 | 2.1523 | 3.4837 | 2.7740 | 4.1222 | 2.8190 | 3.5132 | C3 | 2.5388 | 1.5126 | | 1.5426 | 2.5684 | 3.5362 | 2.8028 | 2.2281 | 1.1084 | 1.1078 | 2.1696 | 2.1740 | 3.5283 | 2.8435 | 2.8285 | C4 | 3.6300 | 2.5475 | 1.5426 | | 1.5346 | 4.5237 | 3.9389 | 2.8819 | 2.1699 | 2.1812 | 1.1071 | 1.1088 | 2.1910 | 2.1923 | 2.1866 | C5 | 3.7967 | 3.1231 | 2.5684 | 1.5346 | | 4.5949 | 3.9537 | 3.5547 | 3.5073 | 2.8007 | 2.1721 | 2.1753 | 1.1051 | 1.1049 | 1.1068 | H6 | 1.0965 | 2.1366 | 3.5362 | 4.5237 | 4.5949 | | 1.8728 | 2.4547 | 4.1460 | 3.7503 | 5.5094 | 4.7352 | 5.5841 | 3.8888 | 4.7979 | H7 | 1.0982 | 2.1354 | 2.8028 | 3.9389 | 3.9537 | 1.8728 | | 3.1030 | 3.5369 | 2.4581 | 4.7102 | 4.5680 | 5.0443 | 3.5872 | 3.7570 | H8 | 2.1136 | 1.1018 | 2.2281 | 2.8819 | 3.5547 | 2.4547 | 3.1030 | | 2.5778 | 3.1198 | 3.8569 | 2.6613 | 4.3871 | 3.1438 | 4.2016 | H9 | 3.2273 | 2.1453 | 1.1084 | 2.1699 | 3.5073 | 4.1460 | 3.5369 | 2.5778 | | 1.7798 | 2.4366 | 2.5167 | 4.3426 | 3.8667 | 3.8026 | H10 | 2.6566 | 2.1523 | 1.1078 | 2.1812 | 2.8007 | 3.7503 | 2.4581 | 3.1198 | 1.7798 | | 2.5320 | 3.0921 | 3.8130 | 3.1677 | 2.6073 | H11 | 4.5470 | 3.4837 | 2.1696 | 1.1071 | 2.1721 | 5.5094 | 4.7102 | 3.8569 | 2.4366 | 2.5320 | | 1.7732 | 2.5113 | 3.1006 | 2.5356 | H12 | 4.0124 | 2.7740 | 2.1740 | 1.1088 | 2.1753 | 4.7352 | 4.5680 | 2.6613 | 2.5167 | 3.0921 | 1.7732 | | 2.5218 | 2.5431 | 3.1003 | H13 | 4.8527 | 4.1222 | 3.5283 | 2.1910 | 1.1051 | 5.5841 | 5.0443 | 4.3871 | 4.3426 | 3.8130 | 2.5113 | 2.5218 | | 1.7890 | 1.7879 | H14 | 3.2865 | 2.8190 | 2.8435 | 2.1923 | 1.1049 | 3.8888 | 3.5872 | 3.1438 | 3.8667 | 3.1677 | 3.1006 | 2.5431 | 1.7890 | | 1.7845 | H15 | 3.9149 | 3.5132 | 2.8285 | 2.1866 | 1.1068 | 4.7979 | 3.7570 | 4.2016 | 3.8026 | 2.6073 | 2.5356 | 3.1003 | 1.7879 | 1.7845 | |
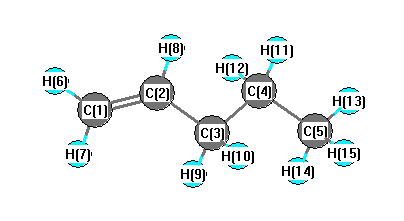 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.039 |
|
C1 |
C2 |
H8 |
118.940 |
| C2 |
C1 |
H6 |
121.546 |
|
C2 |
C1 |
H7 |
121.295 |
| C2 |
C3 |
C4 |
112.980 |
|
C2 |
C3 |
H9 |
108.894 |
| C2 |
C3 |
H10 |
109.474 |
|
C3 |
C2 |
H8 |
116.021 |
| C3 |
C4 |
C5 |
113.161 |
|
C3 |
C4 |
H11 |
108.818 |
| C3 |
C4 |
H12 |
109.065 |
|
C4 |
C3 |
H9 |
108.772 |
| C4 |
C3 |
H10 |
109.679 |
|
C4 |
C5 |
H13 |
111.158 |
| C4 |
C5 |
H14 |
111.279 |
|
C4 |
C5 |
H15 |
110.715 |
| C5 |
C4 |
H11 |
109.554 |
|
C5 |
C4 |
H12 |
109.710 |
| H6 |
C1 |
H7 |
117.158 |
|
H9 |
C3 |
H10 |
106.851 |
| H11 |
C4 |
H12 |
106.299 |
|
H13 |
C5 |
H14 |
108.095 |
| H13 |
C5 |
H15 |
107.859 |
|
H14 |
C5 |
H15 |
107.583 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
