Jump to
S1C2
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -195.925668 |
| Energy at 298.15K | -195.935961 |
| HF Energy | -195.154492 |
| Nuclear repulsion energy | 172.207474 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3266 |
3124 |
15.85 |
|
|
|
| 2 |
A |
3175 |
3037 |
3.95 |
|
|
|
| 3 |
A |
3159 |
3023 |
25.24 |
|
|
|
| 4 |
A |
3157 |
3020 |
32.81 |
|
|
|
| 5 |
A |
3145 |
3009 |
35.54 |
|
|
|
| 6 |
A |
3108 |
2973 |
29.48 |
|
|
|
| 7 |
A |
3100 |
2966 |
23.27 |
|
|
|
| 8 |
A |
3065 |
2932 |
10.73 |
|
|
|
| 9 |
A |
3060 |
2927 |
44.56 |
|
|
|
| 10 |
A |
3052 |
2919 |
23.05 |
|
|
|
| 11 |
A |
1739 |
1664 |
3.89 |
|
|
|
| 12 |
A |
1507 |
1442 |
2.35 |
|
|
|
| 13 |
A |
1499 |
1434 |
8.06 |
|
|
|
| 14 |
A |
1492 |
1427 |
0.31 |
|
|
|
| 15 |
A |
1483 |
1419 |
4.86 |
|
|
|
| 16 |
A |
1460 |
1397 |
0.84 |
|
|
|
| 17 |
A |
1428 |
1366 |
3.84 |
|
|
|
| 18 |
A |
1387 |
1326 |
2.31 |
|
|
|
| 19 |
A |
1368 |
1309 |
1.97 |
|
|
|
| 20 |
A |
1322 |
1265 |
1.57 |
|
|
|
| 21 |
A |
1296 |
1240 |
0.35 |
|
|
|
| 22 |
A |
1265 |
1211 |
1.30 |
|
|
|
| 23 |
A |
1201 |
1149 |
0.33 |
|
|
|
| 24 |
A |
1116 |
1067 |
1.27 |
|
|
|
| 25 |
A |
1095 |
1048 |
4.18 |
|
|
|
| 26 |
A |
1027 |
983 |
9.81 |
|
|
|
| 27 |
A |
1018 |
974 |
3.48 |
|
|
|
| 28 |
A |
950 |
909 |
1.22 |
|
|
|
| 29 |
A |
940 |
899 |
30.91 |
|
|
|
| 30 |
A |
890 |
852 |
0.09 |
|
|
|
| 31 |
A |
879 |
841 |
1.75 |
|
|
|
| 32 |
A |
779 |
745 |
2.34 |
|
|
|
| 33 |
A |
641 |
613 |
7.13 |
|
|
|
| 34 |
A |
436 |
417 |
0.31 |
|
|
|
| 35 |
A |
392 |
375 |
1.13 |
|
|
|
| 36 |
A |
301 |
288 |
0.20 |
|
|
|
| 37 |
A |
230 |
220 |
0.03 |
|
|
|
| 38 |
A |
121 |
115 |
0.03 |
|
|
|
| 39 |
A |
91 |
87 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30319.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29006.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.111 |
0.568 |
0.124 |
| C2 |
-1.184 |
-0.250 |
-0.407 |
| C3 |
-0.062 |
-0.907 |
0.365 |
| C4 |
1.336 |
-0.521 |
-0.158 |
| C5 |
1.659 |
0.968 |
0.023 |
| H6 |
-2.903 |
1.013 |
-0.490 |
| H7 |
-2.108 |
0.815 |
1.193 |
| H8 |
-1.222 |
-0.470 |
-1.486 |
| H9 |
-0.176 |
-2.007 |
0.299 |
| H10 |
-0.146 |
-0.640 |
1.437 |
| H11 |
2.095 |
-1.131 |
0.368 |
| H12 |
1.409 |
-0.791 |
-1.230 |
| H13 |
2.671 |
1.206 |
-0.349 |
| H14 |
0.936 |
1.604 |
-0.518 |
| H15 |
1.616 |
1.251 |
1.092 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3452 | 2.5360 | 3.6261 | 3.7919 | 1.0961 | 1.0977 | 2.1112 | 3.2255 | 2.6542 | 4.5428 | 4.0089 | 4.8472 | 3.2812 | 3.9107 |
C2 | 1.3452 | | 1.5122 | 2.5472 | 3.1224 | 2.1341 | 2.1330 | 1.1014 | 2.1450 | 2.1516 | 3.4829 | 2.7741 | 4.1209 | 2.8179 | 3.5127 | C3 | 2.5360 | 1.5122 | | 1.5423 | 2.5680 | 3.5334 | 2.7998 | 2.2274 | 1.1079 | 1.1073 | 2.1690 | 2.1734 | 3.5275 | 2.8425 | 2.8286 | C4 | 3.6261 | 2.5472 | 1.5423 | | 1.5343 | 4.5200 | 3.9340 | 2.8828 | 2.1693 | 2.1804 | 1.1066 | 1.1083 | 2.1904 | 2.1916 | 2.1861 | C5 | 3.7919 | 3.1224 | 2.5680 | 1.5343 | | 4.5901 | 3.9472 | 3.5557 | 3.5065 | 2.8002 | 2.1715 | 2.1746 | 1.1046 | 1.1044 | 1.1063 | H6 | 1.0961 | 2.1341 | 3.5334 | 4.5200 | 4.5901 | | 1.8718 | 2.4522 | 4.1441 | 3.7476 | 5.5054 | 4.7321 | 5.5785 | 3.8835 | 4.7935 | H7 | 1.0977 | 2.1330 | 2.7998 | 3.9340 | 3.9472 | 1.8718 | | 3.1005 | 3.5355 | 2.4555 | 4.7051 | 4.5636 | 5.0370 | 3.5800 | 3.7508 | H8 | 2.1112 | 1.1014 | 2.2274 | 2.8828 | 3.5557 | 2.4522 | 3.1005 | | 2.5766 | 3.1186 | 3.8570 | 2.6630 | 4.3877 | 3.1448 | 4.2023 | H9 | 3.2255 | 2.1450 | 1.1079 | 2.1693 | 3.5065 | 4.1441 | 3.5355 | 2.5766 | | 1.7788 | 2.4359 | 2.5160 | 4.3414 | 3.8653 | 3.8020 | H10 | 2.6542 | 2.1516 | 1.1073 | 2.1804 | 2.8002 | 3.7476 | 2.4555 | 3.1186 | 1.7788 | | 2.5308 | 3.0908 | 3.8121 | 3.1666 | 2.6075 | H11 | 4.5428 | 3.4829 | 2.1690 | 1.1066 | 2.1715 | 5.5054 | 4.7051 | 3.8570 | 2.4359 | 2.5308 | | 1.7723 | 2.5108 | 3.0995 | 2.5347 | H12 | 4.0089 | 2.7741 | 2.1734 | 1.1083 | 2.1746 | 4.7321 | 4.5636 | 2.6630 | 2.5160 | 3.0908 | 1.7723 | | 2.5206 | 2.5425 | 3.0992 | H13 | 4.8472 | 4.1209 | 3.5275 | 2.1904 | 1.1046 | 5.5785 | 5.0370 | 4.3877 | 4.3414 | 3.8121 | 2.5108 | 2.5206 | | 1.7882 | 1.7870 | H14 | 3.2812 | 2.8179 | 2.8425 | 2.1916 | 1.1044 | 3.8835 | 3.5800 | 3.1448 | 3.8653 | 3.1666 | 3.0995 | 2.5425 | 1.7882 | | 1.7837 | H15 | 3.9107 | 3.5127 | 2.8286 | 2.1861 | 1.1063 | 4.7935 | 3.7508 | 4.2023 | 3.8020 | 2.6075 | 2.5347 | 3.0992 | 1.7870 | 1.7837 | |
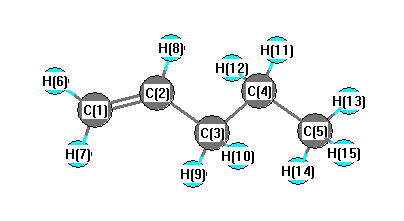 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.024 |
|
C1 |
C2 |
H8 |
118.957 |
| C2 |
C1 |
H6 |
121.557 |
|
C2 |
C1 |
H7 |
121.318 |
| C2 |
C3 |
C4 |
112.998 |
|
C2 |
C3 |
H9 |
108.927 |
| C2 |
C3 |
H10 |
109.469 |
|
C3 |
C2 |
H8 |
116.019 |
| C3 |
C4 |
C5 |
113.166 |
|
C3 |
C4 |
H11 |
108.822 |
| C3 |
C4 |
H12 |
109.067 |
|
C4 |
C3 |
H9 |
108.769 |
| C4 |
C3 |
H10 |
109.657 |
|
C4 |
C5 |
H13 |
111.161 |
| C4 |
C5 |
H14 |
111.271 |
|
C4 |
C5 |
H15 |
110.723 |
| C5 |
C4 |
H11 |
109.555 |
|
C5 |
C4 |
H12 |
109.704 |
| H6 |
C1 |
H7 |
117.125 |
|
H9 |
C3 |
H10 |
106.828 |
| H11 |
C4 |
H12 |
106.291 |
|
H13 |
C5 |
H14 |
108.096 |
| H13 |
C5 |
H15 |
107.858 |
|
H14 |
C5 |
H15 |
107.582 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
