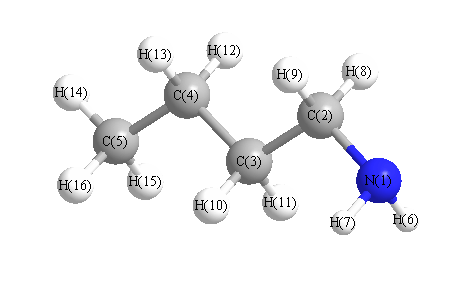
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3469 |
3348 |
1.55 |
|
|
|
| 2 |
A' |
3121 |
3012 |
34.48 |
|
|
|
| 3 |
A' |
3035 |
2929 |
30.18 |
|
|
|
| 4 |
A' |
3029 |
2923 |
96.15 |
|
|
|
| 5 |
A' |
3022 |
2916 |
0.28 |
|
|
|
| 6 |
A' |
3005 |
2900 |
18.90 |
|
|
|
| 7 |
A' |
1651 |
1593 |
16.68 |
|
|
|
| 8 |
A' |
1482 |
1430 |
4.33 |
|
|
|
| 9 |
A' |
1471 |
1420 |
0.45 |
|
|
|
| 10 |
A' |
1461 |
1409 |
0.47 |
|
|
|
| 11 |
A' |
1456 |
1405 |
0.31 |
|
|
|
| 12 |
A' |
1398 |
1349 |
7.71 |
|
|
|
| 13 |
A' |
1394 |
1345 |
3.70 |
|
|
|
| 14 |
A' |
1363 |
1315 |
3.83 |
|
|
|
| 15 |
A' |
1270 |
1226 |
8.62 |
|
|
|
| 16 |
A' |
1149 |
1109 |
5.45 |
|
|
|
| 17 |
A' |
1108 |
1070 |
13.17 |
|
|
|
| 18 |
A' |
1082 |
1044 |
4.91 |
|
|
|
| 19 |
A' |
1030 |
994 |
21.87 |
|
|
|
| 20 |
A' |
922 |
890 |
62.14 |
|
|
|
| 21 |
A' |
884 |
853 |
105.86 |
|
|
|
| 22 |
A' |
428 |
413 |
4.56 |
|
|
|
| 23 |
A' |
391 |
377 |
0.47 |
|
|
|
| 24 |
A' |
179 |
172 |
1.86 |
|
|
|
| 25 |
A" |
3548 |
3423 |
0.09 |
|
|
|
| 26 |
A" |
3115 |
3006 |
49.03 |
|
|
|
| 27 |
A" |
3078 |
2970 |
75.58 |
|
|
|
| 28 |
A" |
3062 |
2955 |
0.74 |
|
|
|
| 29 |
A" |
3038 |
2932 |
2.14 |
|
|
|
| 30 |
A" |
1468 |
1417 |
6.90 |
|
|
|
| 31 |
A" |
1385 |
1336 |
1.18 |
|
|
|
| 32 |
A" |
1317 |
1271 |
0.40 |
|
|
|
| 33 |
A" |
1297 |
1251 |
0.02 |
|
|
|
| 34 |
A" |
1215 |
1173 |
0.03 |
|
|
|
| 35 |
A" |
1045 |
1008 |
0.54 |
|
|
|
| 36 |
A" |
916 |
884 |
0.03 |
|
|
|
| 37 |
A" |
784 |
757 |
0.31 |
|
|
|
| 38 |
A" |
740 |
714 |
4.45 |
|
|
|
| 39 |
A" |
317 |
306 |
43.11 |
|
|
|
| 40 |
A" |
253 |
245 |
0.08 |
|
|
|
| 41 |
A" |
126 |
122 |
0.39 |
|
|
|
| 42 |
A" |
113 |
109 |
2.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32807.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 31659.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.213 |
|
|
|
| 2 |
C |
0.008 |
|
|
|
| 3 |
C |
-0.090 |
|
|
|
| 4 |
C |
-0.109 |
|
|
|
| 5 |
C |
-0.072 |
|
|
|
| 6 |
H |
0.083 |
|
|
|
| 7 |
H |
0.083 |
|
|
|
| 8 |
H |
0.038 |
|
|
|
| 9 |
H |
0.038 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
H |
0.034 |
|
|
|
| 13 |
H |
0.034 |
|
|
|
| 14 |
H |
0.037 |
|
|
|
| 15 |
H |
0.040 |
|
|
|
| 16 |
H |
0.040 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.978 |
0.798 |
0.000 |
1.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.900 |
2.820 |
0.000 |
| y |
2.820 |
-32.568 |
0.000 |
| z |
0.000 |
0.000 |
-32.091 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.571 |
2.820 |
0.000 |
| y |
2.820 |
3.428 |
0.000 |
| z |
0.000 |
0.000 |
4.143 |
|
| Polar |
|---|
| 3z2-r2 | 8.286 |
|---|
| x2-y2 | -7.332 |
|---|
| xy | 2.820 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.164 |
0.329 |
0.000 |
| y |
0.329 |
7.632 |
0.000 |
| z |
0.000 |
0.000 |
7.291 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |