return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-213.158861
Energy at 298.15K-213.171612
Nuclear repulsion energy184.560348
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3493 3342 1.75      
2 A' 3148 3012 35.59      
3 A' 3061 2929 22.22      
4 A' 3059 2927 79.10      
5 A' 3053 2921 9.39      
6 A' 3036 2904 22.07      
7 A' 1675 1603 16.59      
8 A' 1512 1446 2.90      
9 A' 1503 1438 0.31      
10 A' 1492 1427 0.56      
11 A' 1487 1423 0.17      
12 A' 1435 1373 8.20      
13 A' 1425 1363 2.86      
14 A' 1389 1329 4.15      
15 A' 1291 1236 12.06      
16 A' 1170 1119 7.78      
17 A' 1125 1076 13.95      
18 A' 1100 1053 4.93      
19 A' 1054 1008 25.73      
20 A' 950 909 105.43      
21 A' 911 872 28.40      
22 A' 434 416 4.30      
23 A' 396 379 0.48      
24 A' 181 173 2.03      
25 A" 3574 3420 0.54      
26 A" 3144 3008 53.26      
27 A" 3113 2978 70.71      
28 A" 3097 2963 2.80      
29 A" 3074 2941 3.15      
30 A" 1500 1435 5.66      
31 A" 1410 1349 1.27      
32 A" 1333 1275 0.40      
33 A" 1315 1258 0.00      
34 A" 1239 1185 0.00      
35 A" 1057 1011 0.80      
36 A" 926 886 0.06      
37 A" 791 757 0.38      
38 A" 742 710 1.93      
39 A" 313 300 44.48      
40 A" 259 247 0.15      
41 A" 125 120 0.07      
42 A" 111 106 2.62      

Unscaled Zero Point Vibrational Energy (zpe) 33249.9 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 31810.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.58800 0.06440 0.06128

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.544 0.440 0.000
C2 1.348 -0.413 0.000
C3 0.000 0.324 0.000
C4 -1.211 -0.618 0.000
C5 -2.551 0.130 0.000
H6 2.487 1.068 0.808
H7 2.487 1.068 -0.808
H8 1.402 -1.076 0.885
H9 1.402 -1.076 -0.885
H10 -0.049 0.988 -0.888
H11 -0.049 0.988 0.888
H12 -1.157 -1.281 0.886
H13 -1.157 -1.281 -0.886
H14 -3.405 -0.571 0.000
H15 -2.643 0.776 0.893
H16 -2.643 0.776 -0.893

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46842.54623.90095.10381.02471.02472.09402.09402.79482.79484.17604.17606.03355.27315.2731
C21.46841.53652.56743.93652.03472.03471.10741.10742.16872.16872.79522.79524.75534.25854.2585
C32.54621.53651.53452.55822.71822.71822.17052.17051.10991.10992.16802.16803.52042.82552.8255
C43.90092.56741.53451.53424.14334.14332.79682.79682.17242.17241.10811.10812.19402.18832.1883
C55.10383.93652.55821.53425.18735.18734.22654.22652.79012.79012.17242.17241.10451.10581.1058
H61.02472.03472.71824.14335.18731.61632.40362.93903.05172.53814.33534.65406.16805.13815.4117
H71.02472.03472.71824.14335.18731.61632.93902.40362.53813.05174.65404.33536.16805.41175.1381
H82.09401.10742.17052.79684.22652.40362.93901.76973.08412.52372.56703.11864.91364.44864.7906
H92.09401.10742.17052.79684.22652.93902.40361.76972.52373.08413.11862.56704.91364.79064.4486
H102.79482.16871.10992.17242.79013.05172.53813.08412.52371.77593.08622.52533.80533.15312.6022
H112.79482.16871.10992.17242.79012.53813.05172.52373.08411.77592.52533.08623.80532.60223.1531
H124.17602.79522.16801.10812.17244.33534.65402.56703.11863.08622.52531.77222.51852.53753.0988
H134.17602.79522.16801.10812.17244.65404.33533.11862.56702.52533.08621.77222.51853.09882.5375
H146.03354.75533.52042.19401.10456.16806.16804.91364.91363.80533.80532.51852.51851.78651.7865
H155.27314.25852.82552.18831.10585.13815.41174.44864.79063.15312.60222.53753.09881.78651.7853
H165.27314.25852.82552.18831.10585.41175.13814.79064.44862.60223.15313.09882.53751.78651.7853

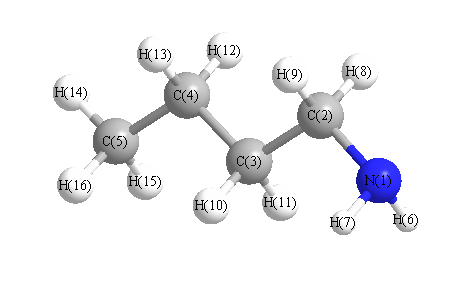
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 115.834 N1 C2 H8 107.952
N1 C2 H9 107.952 C2 N1 H6 108.079
C2 N1 H7 108.079 C2 C3 C4 113.446
C2 C3 H10 109.012 C2 C3 H11 109.012
C3 C2 H8 109.289 C3 C2 H9 109.289
C3 C4 C5 112.948 C3 C4 H12 109.190
C3 C4 H13 109.190 C4 C3 H10 109.425
C4 C3 H11 109.425 C4 C5 H14 111.460
C4 C5 H15 110.928 C4 C5 H16 110.928
C5 C4 H12 109.545 C5 C4 H13 109.545
H6 N1 H7 104.122 H8 C2 H9 106.081
H10 C3 H11 106.271 H12 C4 H13 106.197
H14 C5 H15 107.854 H14 C5 H16 107.854
H15 C5 H16 107.657
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability