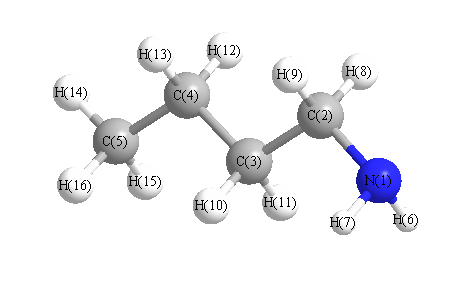
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3348 |
3329 |
4.44 |
|
|
|
| 2 |
A' |
3048 |
3030 |
34.01 |
|
|
|
| 3 |
A' |
2963 |
2946 |
34.27 |
|
|
|
| 4 |
A' |
2952 |
2934 |
82.39 |
|
|
|
| 5 |
A' |
2945 |
2927 |
10.55 |
|
|
|
| 6 |
A' |
2928 |
2911 |
16.85 |
|
|
|
| 7 |
A' |
1602 |
1593 |
13.50 |
|
|
|
| 8 |
A' |
1434 |
1426 |
3.24 |
|
|
|
| 9 |
A' |
1423 |
1415 |
0.36 |
|
|
|
| 10 |
A' |
1411 |
1403 |
0.41 |
|
|
|
| 11 |
A' |
1405 |
1397 |
0.29 |
|
|
|
| 12 |
A' |
1347 |
1339 |
1.05 |
|
|
|
| 13 |
A' |
1343 |
1335 |
9.60 |
|
|
|
| 14 |
A' |
1317 |
1310 |
3.84 |
|
|
|
| 15 |
A' |
1229 |
1222 |
10.13 |
|
|
|
| 16 |
A' |
1119 |
1112 |
5.02 |
|
|
|
| 17 |
A' |
1075 |
1069 |
7.39 |
|
|
|
| 18 |
A' |
1055 |
1048 |
5.57 |
|
|
|
| 19 |
A' |
1003 |
998 |
25.83 |
|
|
|
| 20 |
A' |
898 |
893 |
77.82 |
|
|
|
| 21 |
A' |
866 |
861 |
78.75 |
|
|
|
| 22 |
A' |
418 |
415 |
4.41 |
|
|
|
| 23 |
A' |
381 |
379 |
0.46 |
|
|
|
| 24 |
A' |
174 |
173 |
1.81 |
|
|
|
| 25 |
A" |
3425 |
3405 |
1.43 |
|
|
|
| 26 |
A" |
3040 |
3022 |
45.81 |
|
|
|
| 27 |
A" |
2998 |
2981 |
76.34 |
|
|
|
| 28 |
A" |
2984 |
2966 |
0.00 |
|
|
|
| 29 |
A" |
2960 |
2942 |
1.88 |
|
|
|
| 30 |
A" |
1420 |
1412 |
6.37 |
|
|
|
| 31 |
A" |
1346 |
1338 |
1.57 |
|
|
|
| 32 |
A" |
1276 |
1268 |
0.45 |
|
|
|
| 33 |
A" |
1257 |
1250 |
0.05 |
|
|
|
| 34 |
A" |
1177 |
1170 |
0.03 |
|
|
|
| 35 |
A" |
1016 |
1010 |
0.50 |
|
|
|
| 36 |
A" |
889 |
884 |
0.04 |
|
|
|
| 37 |
A" |
763 |
759 |
0.25 |
|
|
|
| 38 |
A" |
723 |
719 |
4.54 |
|
|
|
| 39 |
A" |
324 |
322 |
40.33 |
|
|
|
| 40 |
A" |
249 |
248 |
0.04 |
|
|
|
| 41 |
A" |
125 |
124 |
0.53 |
|
|
|
| 42 |
A" |
112 |
112 |
1.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31882.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31697.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.169 |
|
|
|
| 2 |
C |
0.005 |
|
|
|
| 3 |
C |
-0.058 |
|
|
|
| 4 |
C |
-0.079 |
|
|
|
| 5 |
C |
-0.047 |
|
|
|
| 6 |
H |
0.072 |
|
|
|
| 7 |
H |
0.072 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
| 9 |
H |
0.027 |
|
|
|
| 10 |
H |
0.010 |
|
|
|
| 11 |
H |
0.010 |
|
|
|
| 12 |
H |
0.022 |
|
|
|
| 13 |
H |
0.022 |
|
|
|
| 14 |
H |
0.026 |
|
|
|
| 15 |
H |
0.030 |
|
|
|
| 16 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.933 |
0.783 |
0.000 |
1.218 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.013 |
2.773 |
0.000 |
| y |
2.773 |
-32.953 |
0.000 |
| z |
0.000 |
0.000 |
-32.531 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.270 |
2.773 |
0.000 |
| y |
2.773 |
3.318 |
0.000 |
| z |
0.000 |
0.000 |
3.952 |
|
| Polar |
|---|
| 3z2-r2 | 7.904 |
|---|
| x2-y2 | -7.059 |
|---|
| xy | 2.773 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.670 |
0.354 |
0.000 |
| y |
0.354 |
7.919 |
0.000 |
| z |
0.000 |
0.000 |
7.536 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |