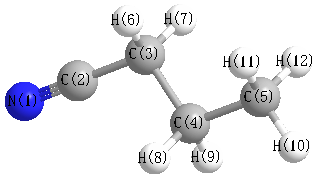
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3135 |
3025 |
20.69 |
|
|
|
| 2 |
A' |
3061 |
2954 |
21.85 |
|
|
|
| 3 |
A' |
3049 |
2942 |
2.09 |
|
|
|
| 4 |
A' |
3042 |
2936 |
21.40 |
|
|
|
| 5 |
A' |
2369 |
2286 |
12.34 |
|
|
|
| 6 |
A' |
1479 |
1428 |
6.72 |
|
|
|
| 7 |
A' |
1466 |
1415 |
1.00 |
|
|
|
| 8 |
A' |
1439 |
1389 |
4.00 |
|
|
|
| 9 |
A' |
1401 |
1352 |
2.10 |
|
|
|
| 10 |
A' |
1376 |
1328 |
2.77 |
|
|
|
| 11 |
A' |
1286 |
1241 |
3.00 |
|
|
|
| 12 |
A' |
1113 |
1074 |
2.32 |
|
|
|
| 13 |
A' |
1069 |
1032 |
0.19 |
|
|
|
| 14 |
A' |
977 |
943 |
1.62 |
|
|
|
| 15 |
A' |
887 |
856 |
2.16 |
|
|
|
| 16 |
A' |
535 |
516 |
0.94 |
|
|
|
| 17 |
A' |
348 |
336 |
0.05 |
|
|
|
| 18 |
A' |
166 |
160 |
5.29 |
|
|
|
| 19 |
A" |
3127 |
3018 |
40.07 |
|
|
|
| 20 |
A" |
3106 |
2998 |
0.74 |
|
|
|
| 21 |
A" |
3087 |
2979 |
0.04 |
|
|
|
| 22 |
A" |
1470 |
1419 |
8.26 |
|
|
|
| 23 |
A" |
1315 |
1269 |
0.01 |
|
|
|
| 24 |
A" |
1244 |
1200 |
0.00 |
|
|
|
| 25 |
A" |
1112 |
1073 |
0.23 |
|
|
|
| 26 |
A" |
867 |
837 |
0.25 |
|
|
|
| 27 |
A" |
747 |
721 |
4.76 |
|
|
|
| 28 |
A" |
393 |
380 |
0.17 |
|
|
|
| 29 |
A" |
253 |
244 |
0.03 |
|
|
|
| 30 |
A" |
101 |
97 |
3.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22510.2 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 21722.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.140 |
|
|
|
| 2 |
C |
-0.078 |
|
|
|
| 3 |
C |
-0.030 |
|
|
|
| 4 |
C |
-0.109 |
|
|
|
| 5 |
C |
-0.080 |
|
|
|
| 6 |
H |
0.086 |
|
|
|
| 7 |
H |
0.086 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
H |
0.051 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.007 |
0.028 |
0.000 |
4.008 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.783 |
0.997 |
0.000 |
| y |
0.997 |
-29.808 |
0.000 |
| z |
0.000 |
0.000 |
-30.074 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.842 |
0.997 |
0.000 |
| y |
0.997 |
6.120 |
0.000 |
| z |
0.000 |
0.000 |
5.721 |
|
| Polar |
|---|
| 3z2-r2 | 11.443 |
|---|
| x2-y2 | -11.975 |
|---|
| xy | 0.997 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.218 |
-0.296 |
0.000 |
| y |
-0.296 |
6.006 |
0.000 |
| z |
0.000 |
0.000 |
5.581 |
<r2> (average value of r
2) Å
2
| <r2> |
159.624 |
| (<r2>)1/2 |
12.634 |