return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: QCISD(T)=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)=FULL/cc-pVDZ
 hartrees
Energy at 0K-210.809394
Energy at 298.15K-210.816618
HF Energy-210.008941
Nuclear repulsion energy152.657361
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3141 3012        
2 A' 3066 2941        
3 A' 3061 2935        
4 A' 3046 2921        
5 A' 2277 2183        
6 A' 1499 1437        
7 A' 1484 1423        
8 A' 1464 1404        
9 A' 1416 1358        
10 A' 1390 1333        
11 A' 1294 1241        
12 A' 1119 1073        
13 A' 1074 1030        
14 A' 963 924        
15 A' 881 845        
16 A' 515 494        
17 A' 342 328        
18 A' 161 155        
19 A" 3134 3006        
20 A" 3119 2991        
21 A" 3101 2974        
22 A" 1490 1429        
23 A" 1321 1267        
24 A" 1255 1203        
25 A" 1122 1076        
26 A" 867 831        
27 A" 741 711        
28 A" 367 352        
29 A" 246 236        
30 A" 99 95        

Unscaled Zero Point Vibrational Energy (zpe) 22527.2 cm-1
Scaled (by 0.959) Zero Point Vibrational Energy (zpe) 21603.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/cc-pVDZ
ABC
0.78205 0.07414 0.07044

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.641 0.330 0.000
C2 -1.473 0.481 0.000
C3 0.000 0.639 0.000
C4 0.736 -0.716 0.000
C5 2.258 -0.528 0.000
H6 0.283 1.232 0.890
H7 0.283 1.232 -0.890
H8 0.421 -1.296 0.888
H9 0.421 -1.296 -0.888
H10 2.776 -1.503 0.000
H11 2.588 0.034 0.894
H12 2.588 0.034 -0.894

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.17752.65903.53514.97363.18693.18693.57893.57895.71895.31335.3133
C21.17751.48172.51263.86532.10732.10732.74512.74514.68984.18274.1827
C32.65901.48171.54202.54181.10621.10622.17042.17043.50652.80452.8045
C43.53512.51261.54201.53392.18912.18911.10641.10642.18692.18932.1893
C54.97363.86532.54181.53392.79092.79092.18042.18041.10421.10621.1062
H63.18692.10731.10622.18912.79091.77942.53213.09393.80622.59803.1516
H73.18692.10731.10622.18912.79091.77943.09392.53213.80623.15162.5980
H83.57892.74512.17041.10642.18042.53213.09391.77602.52562.54263.1050
H93.57892.74512.17041.10642.18043.09392.53211.77602.52563.10502.5426
H105.71894.68983.50652.18691.10423.80623.80622.52562.52561.78761.7876
H115.31334.18272.80452.18931.10622.59803.15162.54263.10501.78761.7886
H125.31334.18272.80452.18931.10623.15162.59803.10502.54261.78761.7886

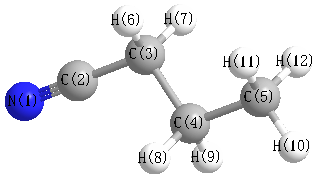
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.747 C2 C3 C4 112.381
C2 C3 H6 108.163 C2 C3 H7 108.163
C3 C4 C5 111.448 C3 C4 H8 108.965
C3 C4 H9 108.965 C4 C3 H6 110.431
C4 C3 H7 110.431 C4 C5 H10 110.932
C4 C5 H11 111.001 C4 C5 H12 111.001
C5 C4 H8 110.288 C5 C4 H9 110.288
H6 C3 H7 107.088 H8 C4 H9 106.759
H10 C5 H11 107.943 H10 C5 H12 107.943
H11 C5 H12 107.884
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability