return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: MP3=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3=FULL/cc-pVDZ
 hartrees
Energy at 0K-210.769817
Energy at 298.15K-210.777125
HF Energy-210.011834
Nuclear repulsion energy153.274139
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3184 2994 22.89      
2 A' 3112 2926 21.83      
3 A' 3105 2920 1.75      
4 A' 3088 2904 21.54      
5 A' 2430 2285 3.28      
6 A' 1513 1423 5.31      
7 A' 1499 1409 0.81      
8 A' 1481 1392 2.88      
9 A' 1435 1349 1.51      
10 A' 1410 1326 1.86      
11 A' 1312 1234 4.20      
12 A' 1134 1066 1.00      
13 A' 1088 1023 0.03      
14 A' 985 926 2.34      
15 A' 896 842 1.82      
16 A' 532 501 0.64      
17 A' 348 327 0.09      
18 A' 166 156 5.45      
19 A" 3177 2987 45.52      
20 A" 3163 2974 0.01      
21 A" 3145 2957 0.13      
22 A" 1504 1414 7.22      
23 A" 1337 1257 0.04      
24 A" 1270 1195 0.01      
25 A" 1138 1070 0.03      
26 A" 878 826 0.19      
27 A" 751 706 2.50      
28 A" 384 361 0.31      
29 A" 251 236 0.02      
30 A" 102 96 3.72      

Unscaled Zero Point Vibrational Energy (zpe) 22909.2 cm-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 21541.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/cc-pVDZ
ABC
0.78803 0.07467 0.07094

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.624 0.348 0.000
C2 -1.470 0.488 0.000
C3 0.000 0.637 0.000
C4 0.728 -0.720 0.000
C5 2.249 -0.541 0.000
H6 0.285 1.227 0.887
H7 0.285 1.227 -0.887
H8 0.412 -1.297 0.885
H9 0.412 -1.297 -0.885
H10 2.760 -1.516 0.000
H11 2.582 0.017 0.892
H12 2.582 0.017 -0.892

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.16272.63963.51804.95333.16553.16553.56393.56395.69735.29245.2924
C21.16271.47702.50793.85842.10022.10022.74012.74014.68044.17564.1756
C32.63961.47701.53962.53871.10281.10282.16592.16593.50022.80142.8014
C43.51802.50791.53961.53142.18472.18471.10301.10302.18212.18532.1853
C54.95333.85842.53871.53142.78732.78732.17542.17541.10111.10311.1031
H63.16552.10021.10282.18472.78731.77362.52703.08633.79952.59663.1472
H73.16552.10021.10282.18472.78731.77363.08632.52703.79953.14722.5966
H83.56392.74012.16591.10302.17542.52703.08631.77022.51932.53753.0977
H93.56392.74012.16591.10302.17543.08632.52701.77022.51933.09772.5375
H105.69734.68043.50022.18211.10113.79953.79952.51932.51931.78221.7822
H115.29244.17562.80142.18531.10312.59663.14722.53753.09771.78221.7832
H125.29244.17562.80142.18531.10313.14722.59663.09772.53751.78221.7832

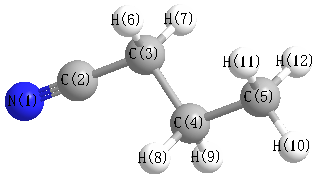
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.841 C2 C3 C4 112.460
C2 C3 H6 108.118 C2 C3 H7 108.118
C3 C4 C5 111.511 C3 C4 H8 108.969
C3 C4 H9 108.969 C4 C3 H6 110.450
C4 C3 H7 110.450 C4 C5 H10 110.922
C4 C5 H11 111.049 C4 C5 H12 111.049
C5 C4 H8 110.269 C5 C4 H9 110.269
H6 C3 H7 107.053 H8 C4 H9 106.722
H10 C5 H11 107.914 H10 C5 H12 107.914
H11 C5 H12 107.851
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability