return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-555.652334
Energy at 298.15K 
HF Energy-554.836787
Nuclear repulsion energy221.498338
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3126 2980        
2 A' 3056 2913        
3 A' 3038 2896        
4 A' 3033 2890        
5 A' 3027 2885        
6 A' 2711 2583        
7 A' 1498 1428        
8 A' 1488 1418        
9 A' 1479 1410        
10 A' 1477 1407        
11 A' 1412 1346        
12 A' 1398 1333        
13 A' 1329 1267        
14 A' 1241 1183        
15 A' 1136 1083        
16 A' 1091 1040        
17 A' 1052 1003        
18 A' 929 886        
19 A' 855 815        
20 A' 754 719        
21 A' 388 370        
22 A' 321 306        
23 A' 149 142        
24 A" 3121 2975        
25 A" 3115 2969        
26 A" 3083 2939        
27 A" 3064 2920        
28 A" 1488 1418        
29 A" 1321 1259        
30 A" 1299 1239        
31 A" 1226 1169        
32 A" 1070 1019        
33 A" 920 877        
34 A" 789 752        
35 A" 739 704        
36 A" 253 241        
37 A" 186 177        
38 A" 113 107        
39 A" 97 92        

Unscaled Zero Point Vibrational Energy (zpe) 28935.1 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 27578.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.52310 0.04390 0.04179

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.390 -1.865 0.000
C2 -0.237 -0.991 0.000
C3 0.000 0.525 0.000
C4 -1.316 1.321 0.000
C5 -1.083 2.839 0.000
H6 0.884 -3.121 0.000
H7 -0.812 -1.287 0.898
H8 -0.812 -1.287 -0.898
H9 0.601 0.803 -0.890
H10 0.601 0.803 0.890
H11 -1.916 1.035 0.889
H12 -1.916 1.035 -0.889
H13 -2.040 3.394 0.000
H14 -0.509 3.150 0.895
H15 -0.509 3.150 -0.895

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.84642.76514.17985.31491.35422.44752.44752.92102.92104.48674.48676.27865.43675.4367
C21.84641.53512.55163.92332.40591.10641.10642.17132.17132.77812.77814.74174.24584.2458
C32.76511.53511.53762.55513.75152.17922.17921.10971.10972.17272.17273.52012.81942.8194
C44.17982.55161.53761.53644.95612.80322.80322.17642.17641.10991.10992.19612.19042.1904
C55.31493.92332.55511.53646.27624.23134.23132.78902.78902.17672.17671.10591.10741.1074
H61.35422.40593.75154.95616.27622.65412.65414.03324.03325.08905.08907.14056.48556.4855
H72.44751.10642.17922.80324.23132.65411.79513.09222.52282.57093.13084.92154.44714.7946
H82.44751.10642.17922.80324.23132.65411.79512.52283.09223.13082.57094.92154.79464.4471
H92.92102.17131.10972.17642.78904.03323.09222.52281.78093.09142.52783.80573.15102.5966
H102.92102.17131.10972.17642.78904.03322.52283.09221.78092.52783.09143.80572.59663.1510
H114.48672.77812.17271.10992.17675.08902.57093.13083.09142.52781.77842.52372.53983.1036
H124.48672.77812.17271.10992.17675.08903.13082.57092.52783.09141.77842.52373.10362.5398
H136.27864.74173.52012.19611.10597.14054.92154.92153.80573.80572.52372.52371.78991.7899
H145.43674.24582.81942.19041.10746.48554.44714.79463.15102.59662.53983.10361.78991.7891
H155.43674.24582.81942.19041.10746.48554.79464.44712.59663.15103.10362.53981.78991.7891

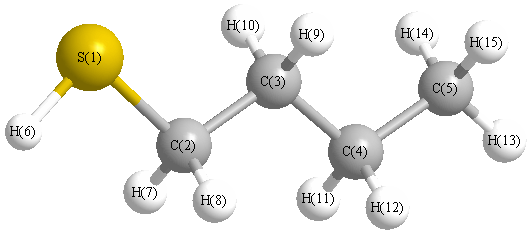
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 109.367 S1 C2 H7 109.399
S1 C2 H8 109.399 C2 S1 H6 96.257
C2 C3 C4 112.283 C2 C3 H9 109.307
C2 C3 H10 109.307 C3 C2 H7 110.113
C3 C2 H8 110.113 C3 C4 C5 112.443
C3 C4 H11 109.240 C3 C4 H12 109.240
C4 C3 H9 109.534 C4 C3 H10 109.534
C4 C5 H13 111.385 C4 C5 H14 110.843
C4 C5 H15 110.843 C5 C4 H11 109.630
C5 C4 H12 109.630 H7 C2 H8 108.431
H9 C3 H10 106.718 H11 C4 H12 106.480
H13 C5 H14 107.936 H13 C5 H15 107.936
H14 C5 H15 107.754
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability