return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-555.629295
Energy at 298.15K 
HF Energy-554.837198
Nuclear repulsion energy221.791039
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3139 3011 35.07      
2 A' 3076 2951 34.66      
3 A' 3054 2930 29.72      
4 A' 3051 2927 22.85      
5 A' 3045 2921 18.14      
6 A' 2730 2619 14.06      
7 A' 1509 1448 3.13      
8 A' 1499 1438 0.93      
9 A' 1491 1431 1.44      
10 A' 1488 1428 0.27      
11 A' 1427 1369 2.17      
12 A' 1413 1356 3.96      
13 A' 1344 1290 6.44      
14 A' 1256 1205 23.36      
15 A' 1148 1101 0.79      
16 A' 1100 1056 0.23      
17 A' 1062 1019 0.01      
18 A' 939 901 1.42      
19 A' 866 831 1.72      
20 A' 765 734 2.20      
21 A' 392 376 0.64      
22 A' 325 312 0.69      
23 A' 152 146 1.08      
24 A" 3136 3009 78.06      
25 A" 3132 3005 1.24      
26 A" 3100 2974 17.00      
27 A" 3081 2956 2.30      
28 A" 1497 1436 5.76      
29 A" 1333 1279 0.15      
30 A" 1311 1258 0.98      
31 A" 1237 1187 0.44      
32 A" 1081 1037 1.03      
33 A" 929 891 1.05      
34 A" 796 764 0.00      
35 A" 745 715 2.02      
36 A" 256 246 0.02      
37 A" 188 180 15.00      
38 A" 113 109 1.59      
39 A" 97 93 3.03      

Unscaled Zero Point Vibrational Energy (zpe) 29150.2 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27966.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.52571 0.04398 0.04188

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.390 -1.862 0.000
C2 -0.233 -0.990 0.000
C3 0.000 0.525 0.000
C4 -1.315 1.318 0.000
C5 -1.087 2.835 0.000
H6 0.890 -3.118 0.000
H7 -0.808 -1.286 0.896
H8 -0.808 -1.286 -0.896
H9 0.601 0.803 -0.889
H10 0.601 0.803 0.889
H11 -1.915 1.032 0.887
H12 -1.915 1.032 -0.887
H13 -2.045 3.386 0.000
H14 -0.515 3.148 0.893
H15 -0.515 3.148 -0.893

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.84202.76194.17465.30981.35232.44272.44272.91772.91774.48094.48096.27175.43345.4334
C21.84201.53332.54943.91972.40541.10471.10472.16822.16822.77562.77564.73674.24324.2432
C32.76191.53331.53582.55293.74992.17592.17591.10811.10812.17012.17013.51652.81832.8183
C44.17462.54941.53581.53444.95342.79942.79942.17392.17391.10831.10832.19322.18852.1885
C55.30983.91972.55291.53446.27244.22614.22612.78692.78692.17372.17371.10481.10611.1061
H61.35232.40543.74994.95346.27242.65362.65364.03074.03075.08615.08617.13516.48316.4831
H72.44271.10472.17592.79944.22612.65361.79123.08742.51962.56783.12614.91494.44354.7900
H82.44271.10472.17592.79944.22612.65361.79122.51963.08743.12612.56784.91494.79004.4435
H92.91772.16821.10812.17392.78694.03073.08742.51961.77743.08752.52553.80253.14932.5967
H102.91772.16821.10812.17392.78694.03072.51963.08741.77742.52553.08753.80252.59673.1493
H114.48092.77562.17011.10832.17375.08612.56783.12613.08752.52551.77492.51962.53763.1000
H124.48092.77562.17011.10832.17375.08613.12612.56782.52553.08751.77492.51963.10002.5376
H136.27174.73673.51652.19321.10487.13514.91494.91493.80253.80252.51962.51961.78751.7875
H145.43344.24322.81832.18851.10616.48314.44354.79003.14932.59672.53763.10001.78751.7863
H155.43344.24322.81832.18851.10616.48314.79004.44352.59673.14933.10002.53761.78751.7863

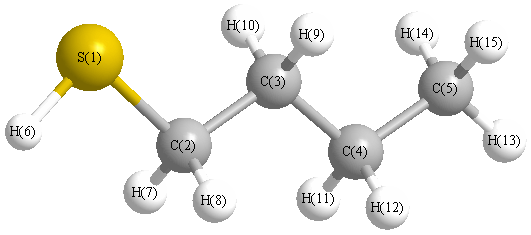
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 109.487 S1 C2 H7 109.425
S1 C2 H8 109.425 C2 S1 H6 96.503
C2 C3 C4 112.334 C2 C3 H9 109.286
C2 C3 H10 109.286 C3 C2 H7 110.078
C3 C2 H8 110.078 C3 C4 C5 112.511
C3 C4 H11 109.254 C3 C4 H12 109.254
C4 C3 H9 109.561 C4 C3 H10 109.561
C4 C5 H13 111.359 C4 C5 H14 110.911
C4 C5 H15 110.911 C5 C4 H11 109.618
C5 C4 H12 109.618 H7 C2 H8 108.327
H9 C3 H10 106.648 H11 C4 H12 106.400
H13 C5 H14 107.900 H13 C5 H15 107.900
H14 C5 H15 107.707
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability