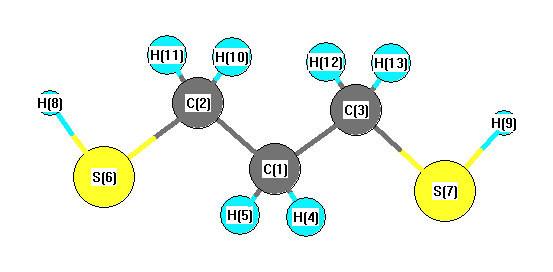
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
2985 |
2968 |
52.55 |
|
|
|
| 2 |
A1 |
2971 |
2954 |
0.02 |
|
|
|
| 3 |
A1 |
2613 |
2598 |
2.93 |
|
|
|
| 4 |
A1 |
1429 |
1421 |
2.27 |
|
|
|
| 5 |
A1 |
1416 |
1408 |
0.11 |
|
|
|
| 6 |
A1 |
1225 |
1217 |
22.07 |
|
|
|
| 7 |
A1 |
1073 |
1066 |
0.94 |
|
|
|
| 8 |
A1 |
825 |
820 |
0.18 |
|
|
|
| 9 |
A1 |
767 |
762 |
0.45 |
|
|
|
| 10 |
A1 |
289 |
287 |
0.25 |
|
|
|
| 11 |
A1 |
122 |
121 |
0.12 |
|
|
|
| 12 |
A2 |
3035 |
3017 |
0.00 |
|
|
|
| 13 |
A2 |
1270 |
1262 |
0.00 |
|
|
|
| 14 |
A2 |
1055 |
1049 |
0.00 |
|
|
|
| 15 |
A2 |
804 |
799 |
0.00 |
|
|
|
| 16 |
A2 |
172 |
171 |
0.00 |
|
|
|
| 17 |
A2 |
88 |
88 |
0.00 |
|
|
|
| 18 |
B1 |
3049 |
3031 |
33.10 |
|
|
|
| 19 |
B1 |
3012 |
2994 |
0.07 |
|
|
|
| 20 |
B1 |
1221 |
1214 |
2.59 |
|
|
|
| 21 |
B1 |
968 |
962 |
3.84 |
|
|
|
| 22 |
B1 |
742 |
738 |
4.22 |
|
|
|
| 23 |
B1 |
163 |
162 |
35.68 |
|
|
|
| 24 |
B1 |
98 |
98 |
0.05 |
|
|
|
| 25 |
B2 |
2978 |
2961 |
2.74 |
|
|
|
| 26 |
B2 |
2613 |
2598 |
12.13 |
|
|
|
| 27 |
B2 |
1411 |
1403 |
0.83 |
|
|
|
| 28 |
B2 |
1312 |
1305 |
17.62 |
|
|
|
| 29 |
B2 |
1169 |
1162 |
17.62 |
|
|
|
| 30 |
B2 |
1048 |
1042 |
0.02 |
|
|
|
| 31 |
B2 |
861 |
856 |
0.18 |
|
|
|
| 32 |
B2 |
686 |
682 |
4.58 |
|
|
|
| 33 |
B2 |
318 |
316 |
6.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21892.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21765.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.031 |
|
|
|
| 2 |
C |
-0.148 |
|
|
|
| 3 |
C |
-0.148 |
|
|
|
| 4 |
H |
0.053 |
|
|
|
| 5 |
H |
0.053 |
|
|
|
| 6 |
S |
-0.105 |
|
|
|
| 7 |
S |
-0.105 |
|
|
|
| 8 |
H |
0.086 |
|
|
|
| 9 |
H |
0.086 |
|
|
|
| 10 |
H |
0.064 |
|
|
|
| 11 |
H |
0.064 |
|
|
|
| 12 |
H |
0.064 |
|
|
|
| 13 |
H |
0.064 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.670 |
2.670 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.059 |
0.000 |
0.000 |
| y |
0.000 |
-46.622 |
0.000 |
| z |
0.000 |
0.000 |
-45.871 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.813 |
0.000 |
0.000 |
| y |
0.000 |
0.843 |
0.000 |
| z |
0.000 |
0.000 |
1.970 |
|
| Polar |
|---|
| 3z2-r2 | 3.939 |
|---|
| x2-y2 | -2.437 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.837 |
0.000 |
0.000 |
| y |
0.000 |
14.322 |
0.000 |
| z |
0.000 |
0.000 |
9.179 |
<r2> (average value of r
2) Å
2
| <r2> |
350.044 |
| (<r2>)1/2 |
18.709 |