return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-213.180634
Energy at 298.15K-213.193335
HF Energy-212.329739
Nuclear repulsion energy187.266834
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3461 3299        
2 A' 3141 2994        
3 A' 3120 2973        
4 A' 3057 2914        
5 A' 3040 2897        
6 A' 2943 2805        
7 A' 1518 1446        
8 A' 1490 1420        
9 A' 1479 1410        
10 A' 1429 1362        
11 A' 1384 1319        
12 A' 1319 1257        
13 A' 1236 1178        
14 A' 1164 1110        
15 A' 1077 1026        
16 A' 915 872        
17 A' 825 787        
18 A' 805 767        
19 A' 427 407        
20 A' 266 253        
21 A' 184 176        
22 A' 111 106        
23 A" 3141 2994        
24 A" 3119 2973        
25 A" 3054 2911        
26 A" 3039 2897        
27 A" 2938 2801        
28 A" 1509 1438        
29 A" 1497 1427        
30 A" 1486 1417        
31 A" 1471 1402        
32 A" 1404 1338        
33 A" 1349 1286        
34 A" 1277 1217        
35 A" 1173 1118        
36 A" 1107 1055        
37 A" 1067 1017        
38 A" 951 906        
39 A" 802 764        
40 A" 422 402        
41 A" 265 253        
42 A" 107 102        

Unscaled Zero Point Vibrational Energy (zpe) 32785.3 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 31247.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.58136 0.06958 0.06560

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.018 -0.297 0.000
C2 0.018 0.521 1.219
C3 0.018 0.521 -1.219
C4 0.018 -0.372 2.463
C5 0.018 -0.372 -2.463
H6 -0.847 -0.852 0.000
H7 -0.841 1.236 1.263
H8 0.938 1.140 1.205
H9 -0.841 1.236 -1.263
H10 0.938 1.140 -1.205
H11 0.074 0.234 3.387
H12 -0.908 -0.979 2.512
H13 0.879 -1.064 2.433
H14 0.074 0.234 -3.387
H15 -0.908 -0.979 -2.512
H16 0.879 -1.064 -2.433

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46801.46802.46382.46381.02802.16392.08932.16392.08933.42852.76262.69293.42852.76262.6929
C21.46802.43761.53133.78832.03001.11831.10912.72182.66572.18772.18622.17504.61494.12624.0736
C31.46802.43763.78831.53132.03002.72182.66571.11831.10914.61494.12624.07362.18772.18622.1750
C42.46381.53133.78834.92542.65412.18212.17134.14774.07271.10621.10821.10545.88105.09645.0192
C52.46383.78831.53134.92542.65414.14774.07272.18212.17135.88105.09645.01921.10621.10821.1054
H61.02802.03002.03002.65412.65412.44052.93442.44052.93443.67402.51592.99093.67402.51592.9909
H72.16391.11832.72182.18214.14772.44051.78292.52613.04432.52042.54313.10144.84384.37704.6811
H82.08931.10912.66572.17134.07272.93441.78293.04432.41042.51553.09942.52444.75974.66004.2548
H92.16392.72181.11834.14772.18212.44052.52613.04431.78294.84384.37704.68112.52042.54313.1014
H102.08932.66571.10914.07272.17132.93443.04432.41041.78294.75974.66004.25482.51553.09942.5244
H113.42852.18774.61491.10625.88103.67402.52042.51554.84384.75971.78871.80046.77366.10166.0172
H122.76262.18624.12621.10825.09642.51592.54313.09944.37704.66001.78871.79096.10165.02395.2591
H132.69292.17504.07361.10545.01922.99093.10142.52444.68114.25481.80041.79096.01725.25914.8668
H143.42854.61492.18775.88101.10623.67404.84384.75972.52042.51556.77366.10166.01721.78871.8004
H152.76264.12622.18625.09641.10822.51594.37704.66002.54313.09946.10165.02395.25911.78871.7909
H162.69294.07362.17505.01921.10542.99094.68114.25483.10142.52446.01725.25914.86681.80041.7909

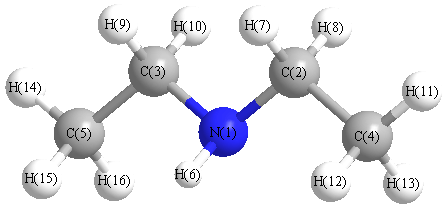
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.450 N1 C2 H7 112.885
N1 C2 H8 107.513 N1 C3 C5 110.450
N1 C3 H9 112.885 N1 C3 H10 107.513
C2 N1 C3 112.253 C2 N1 H6 107.532
C2 C4 H11 111.063 C2 C4 H12 110.821
C2 C4 H13 110.112 C3 N1 H6 107.532
C3 C5 H14 111.063 C3 C5 H15 110.821
C3 C5 H16 110.112 C4 C2 H7 109.904
C4 C2 H8 109.602 C5 C3 H8 150.941
C5 C3 H10 109.602 H7 C2 H8 106.341
H9 C3 H10 106.341 H11 C4 H12 107.756
H11 C4 H13 108.994 H12 C4 H13 108.003
H14 C5 H15 107.756 H14 C5 H16 108.994
H15 C5 H16 108.003
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability