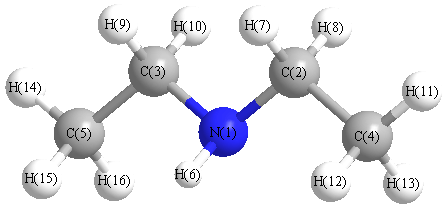
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3333 |
3.92 |
|
|
|
| 2 |
A' |
3033 |
3037 |
46.58 |
|
|
|
| 3 |
A' |
3011 |
3016 |
72.96 |
|
|
|
| 4 |
A' |
2948 |
2953 |
51.41 |
|
|
|
| 5 |
A' |
2943 |
2948 |
31.76 |
|
|
|
| 6 |
A' |
2805 |
2810 |
210.73 |
|
|
|
| 7 |
A' |
1463 |
1466 |
0.13 |
|
|
|
| 8 |
A' |
1434 |
1437 |
1.58 |
|
|
|
| 9 |
A' |
1421 |
1424 |
10.81 |
|
|
|
| 10 |
A' |
1372 |
1374 |
2.63 |
|
|
|
| 11 |
A' |
1344 |
1347 |
0.02 |
|
|
|
| 12 |
A' |
1272 |
1274 |
3.27 |
|
|
|
| 13 |
A' |
1187 |
1189 |
1.04 |
|
|
|
| 14 |
A' |
1117 |
1119 |
9.61 |
|
|
|
| 15 |
A' |
1018 |
1020 |
6.21 |
|
|
|
| 16 |
A' |
885 |
886 |
8.66 |
|
|
|
| 17 |
A' |
804 |
806 |
1.13 |
|
|
|
| 18 |
A' |
755 |
756 |
63.72 |
|
|
|
| 19 |
A' |
413 |
414 |
0.11 |
|
|
|
| 20 |
A' |
258 |
258 |
0.38 |
|
|
|
| 21 |
A' |
181 |
181 |
0.59 |
|
|
|
| 22 |
A' |
108 |
108 |
0.77 |
|
|
|
| 23 |
A" |
3032 |
3037 |
14.78 |
|
|
|
| 24 |
A" |
3011 |
3016 |
16.51 |
|
|
|
| 25 |
A" |
2948 |
2952 |
32.47 |
|
|
|
| 26 |
A" |
2940 |
2945 |
2.21 |
|
|
|
| 27 |
A" |
2798 |
2802 |
21.05 |
|
|
|
| 28 |
A" |
1456 |
1458 |
20.84 |
|
|
|
| 29 |
A" |
1446 |
1449 |
2.15 |
|
|
|
| 30 |
A" |
1431 |
1433 |
5.19 |
|
|
|
| 31 |
A" |
1415 |
1417 |
2.17 |
|
|
|
| 32 |
A" |
1354 |
1356 |
5.71 |
|
|
|
| 33 |
A" |
1302 |
1304 |
51.01 |
|
|
|
| 34 |
A" |
1235 |
1237 |
5.37 |
|
|
|
| 35 |
A" |
1112 |
1114 |
49.84 |
|
|
|
| 36 |
A" |
1069 |
1070 |
17.86 |
|
|
|
| 37 |
A" |
1028 |
1030 |
0.99 |
|
|
|
| 38 |
A" |
915 |
916 |
1.02 |
|
|
|
| 39 |
A" |
790 |
792 |
0.23 |
|
|
|
| 40 |
A" |
414 |
414 |
0.42 |
|
|
|
| 41 |
A" |
258 |
259 |
0.77 |
|
|
|
| 42 |
A" |
118 |
118 |
1.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31585.8 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 31636.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.173 |
|
|
|
| 2 |
C |
0.057 |
|
|
|
| 3 |
C |
0.057 |
|
|
|
| 4 |
C |
0.012 |
|
|
|
| 5 |
C |
0.012 |
|
|
|
| 6 |
H |
0.057 |
|
|
|
| 7 |
H |
-0.028 |
|
|
|
| 8 |
H |
-0.000 |
|
|
|
| 9 |
H |
-0.028 |
|
|
|
| 10 |
H |
-0.000 |
|
|
|
| 11 |
H |
0.002 |
|
|
|
| 12 |
H |
0.003 |
|
|
|
| 13 |
H |
0.012 |
|
|
|
| 14 |
H |
0.002 |
|
|
|
| 15 |
H |
0.003 |
|
|
|
| 16 |
H |
0.012 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.664 |
0.257 |
0.000 |
0.712 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.098 |
1.553 |
0.000 |
| y |
1.553 |
-34.880 |
0.000 |
| z |
0.000 |
0.000 |
-33.918 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.301 |
1.553 |
0.000 |
| y |
1.553 |
-0.872 |
0.000 |
| z |
0.000 |
0.000 |
0.571 |
|
| Polar |
|---|
| 3z2-r2 | 1.141 |
|---|
| x2-y2 | 0.782 |
|---|
| xy | 1.553 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.446 |
0.033 |
0.000 |
| y |
0.033 |
8.003 |
0.000 |
| z |
0.000 |
0.000 |
10.502 |
<r2> (average value of r
2) Å
2
| <r2> |
191.322 |
| (<r2>)1/2 |
13.832 |