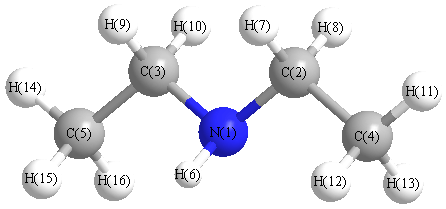
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3487 |
3364 |
0.65 |
|
|
|
| 2 |
A' |
3141 |
3031 |
35.12 |
|
|
|
| 3 |
A' |
3119 |
3010 |
60.43 |
|
|
|
| 4 |
A' |
3046 |
2939 |
65.13 |
|
|
|
| 5 |
A' |
3040 |
2934 |
10.50 |
|
|
|
| 6 |
A' |
2919 |
2817 |
187.46 |
|
|
|
| 7 |
A' |
1503 |
1450 |
1.08 |
|
|
|
| 8 |
A' |
1473 |
1421 |
2.37 |
|
|
|
| 9 |
A' |
1459 |
1408 |
14.30 |
|
|
|
| 10 |
A' |
1417 |
1368 |
2.26 |
|
|
|
| 11 |
A' |
1373 |
1325 |
0.50 |
|
|
|
| 12 |
A' |
1308 |
1262 |
3.62 |
|
|
|
| 13 |
A' |
1223 |
1181 |
1.42 |
|
|
|
| 14 |
A' |
1159 |
1119 |
15.24 |
|
|
|
| 15 |
A' |
1075 |
1038 |
7.69 |
|
|
|
| 16 |
A' |
906 |
875 |
7.57 |
|
|
|
| 17 |
A' |
824 |
795 |
1.52 |
|
|
|
| 18 |
A' |
782 |
755 |
68.94 |
|
|
|
| 19 |
A' |
426 |
411 |
0.21 |
|
|
|
| 20 |
A' |
264 |
255 |
0.56 |
|
|
|
| 21 |
A' |
184 |
178 |
0.69 |
|
|
|
| 22 |
A' |
110 |
106 |
0.95 |
|
|
|
| 23 |
A" |
3141 |
3031 |
12.22 |
|
|
|
| 24 |
A" |
3119 |
3010 |
11.35 |
|
|
|
| 25 |
A" |
3044 |
2938 |
3.41 |
|
|
|
| 26 |
A" |
3039 |
2933 |
29.63 |
|
|
|
| 27 |
A" |
2913 |
2811 |
15.56 |
|
|
|
| 28 |
A" |
1499 |
1447 |
30.95 |
|
|
|
| 29 |
A" |
1485 |
1433 |
0.86 |
|
|
|
| 30 |
A" |
1469 |
1418 |
5.50 |
|
|
|
| 31 |
A" |
1454 |
1404 |
3.68 |
|
|
|
| 32 |
A" |
1389 |
1341 |
14.90 |
|
|
|
| 33 |
A" |
1339 |
1292 |
37.22 |
|
|
|
| 34 |
A" |
1270 |
1225 |
3.71 |
|
|
|
| 35 |
A" |
1186 |
1145 |
46.93 |
|
|
|
| 36 |
A" |
1103 |
1064 |
6.76 |
|
|
|
| 37 |
A" |
1064 |
1027 |
4.32 |
|
|
|
| 38 |
A" |
953 |
920 |
0.16 |
|
|
|
| 39 |
A" |
806 |
778 |
0.30 |
|
|
|
| 40 |
A" |
424 |
409 |
0.59 |
|
|
|
| 41 |
A" |
263 |
254 |
0.73 |
|
|
|
| 42 |
A" |
118 |
114 |
1.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32658.9 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 31515.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.252 |
|
|
|
| 2 |
C |
-0.009 |
|
|
|
| 3 |
C |
-0.009 |
|
|
|
| 4 |
C |
-0.085 |
|
|
|
| 5 |
C |
-0.085 |
|
|
|
| 6 |
H |
0.086 |
|
|
|
| 7 |
H |
0.011 |
|
|
|
| 8 |
H |
0.039 |
|
|
|
| 9 |
H |
0.011 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.040 |
|
|
|
| 12 |
H |
0.037 |
|
|
|
| 13 |
H |
0.050 |
|
|
|
| 14 |
H |
0.040 |
|
|
|
| 15 |
H |
0.037 |
|
|
|
| 16 |
H |
0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.716 |
0.298 |
0.000 |
0.776 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.451 |
1.574 |
0.000 |
| y |
1.574 |
-34.199 |
0.000 |
| z |
0.000 |
0.000 |
-33.178 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.237 |
1.574 |
0.000 |
| y |
1.574 |
-0.884 |
0.000 |
| z |
0.000 |
0.000 |
0.647 |
|
| Polar |
|---|
| 3z2-r2 | 1.294 |
|---|
| x2-y2 | 0.748 |
|---|
| xy | 1.574 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.163 |
0.057 |
0.000 |
| y |
0.057 |
7.674 |
0.000 |
| z |
0.000 |
0.000 |
9.817 |
<r2> (average value of r
2) Å
2
| <r2> |
187.227 |
| (<r2>)1/2 |
13.683 |