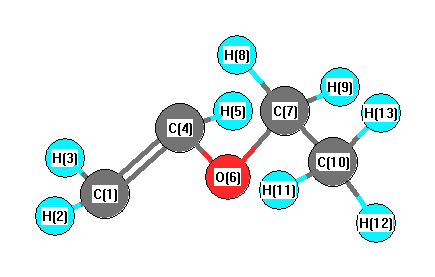
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3214 |
3196 |
4.28 |
|
|
|
| 2 |
A |
3107 |
3089 |
0.02 |
|
|
|
| 3 |
A |
3075 |
3058 |
17.79 |
|
|
|
| 4 |
A |
3074 |
3057 |
19.34 |
|
|
|
| 5 |
A |
3055 |
3038 |
20.87 |
|
|
|
| 6 |
A |
2986 |
2969 |
14.82 |
|
|
|
| 7 |
A |
2958 |
2940 |
46.41 |
|
|
|
| 8 |
A |
2916 |
2899 |
59.47 |
|
|
|
| 9 |
A |
1672 |
1662 |
154.08 |
|
|
|
| 10 |
A |
1446 |
1437 |
8.03 |
|
|
|
| 11 |
A |
1426 |
1418 |
2.05 |
|
|
|
| 12 |
A |
1404 |
1395 |
6.17 |
|
|
|
| 13 |
A |
1376 |
1368 |
11.62 |
|
|
|
| 14 |
A |
1348 |
1341 |
54.56 |
|
|
|
| 15 |
A |
1324 |
1316 |
36.73 |
|
|
|
| 16 |
A |
1285 |
1277 |
4.07 |
|
|
|
| 17 |
A |
1245 |
1238 |
0.20 |
|
|
|
| 18 |
A |
1196 |
1189 |
234.35 |
|
|
|
| 19 |
A |
1126 |
1119 |
4.30 |
|
|
|
| 20 |
A |
1110 |
1104 |
115.38 |
|
|
|
| 21 |
A |
1067 |
1060 |
20.93 |
|
|
|
| 22 |
A |
960 |
954 |
4.53 |
|
|
|
| 23 |
A |
933 |
928 |
17.78 |
|
|
|
| 24 |
A |
828 |
823 |
4.04 |
|
|
|
| 25 |
A |
794 |
789 |
0.14 |
|
|
|
| 26 |
A |
778 |
773 |
42.17 |
|
|
|
| 27 |
A |
675 |
671 |
4.61 |
|
|
|
| 28 |
A |
483 |
480 |
3.74 |
|
|
|
| 29 |
A |
424 |
422 |
1.44 |
|
|
|
| 30 |
A |
247 |
246 |
0.44 |
|
|
|
| 31 |
A |
202 |
201 |
0.24 |
|
|
|
| 32 |
A |
87 |
86 |
1.99 |
|
|
|
| 33 |
A |
22 |
22 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23920.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23781.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.069 |
|
|
|
| 2 |
H |
0.024 |
|
|
|
| 3 |
H |
0.027 |
|
|
|
| 4 |
C |
0.061 |
|
|
|
| 5 |
H |
-0.004 |
|
|
|
| 6 |
O |
-0.219 |
|
|
|
| 7 |
C |
0.049 |
|
|
|
| 8 |
H |
0.031 |
|
|
|
| 9 |
H |
0.031 |
|
|
|
| 10 |
C |
-0.063 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.048 |
|
|
|
| 13 |
H |
0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.276 |
1.171 |
0.000 |
1.732 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.056 |
-0.144 |
0.000 |
| y |
-0.144 |
-30.618 |
-0.001 |
| z |
0.000 |
-0.001 |
-33.158 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.832 |
-0.144 |
0.000 |
| y |
-0.144 |
0.989 |
-0.001 |
| z |
0.000 |
-0.001 |
-2.821 |
|
| Polar |
|---|
| 3z2-r2 | -5.642 |
|---|
| x2-y2 | 0.562 |
|---|
| xy | -0.144 |
|---|
| xz | 0.000 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.117 |
0.767 |
0.000 |
| y |
0.767 |
7.130 |
-0.000 |
| z |
0.000 |
-0.000 |
5.093 |
<r2> (average value of r
2) Å
2
| <r2> |
164.077 |
| (<r2>)1/2 |
12.809 |