Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -230.918266 |
| Energy at 298.15K | -230.924156 |
| HF Energy | -230.918266 |
| Nuclear repulsion energy | 159.990306 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3213 |
3195 |
2.56 |
|
|
|
| 2 |
A |
3209 |
3190 |
4.46 |
|
|
|
| 3 |
A |
3133 |
3115 |
8.33 |
|
|
|
| 4 |
A |
3108 |
3090 |
1.54 |
|
|
|
| 5 |
A |
3107 |
3089 |
2.96 |
|
|
|
| 6 |
A |
3082 |
3064 |
10.36 |
|
|
|
| 7 |
A |
1667 |
1657 |
84.20 |
|
|
|
| 8 |
A |
1646 |
1636 |
185.79 |
|
|
|
| 9 |
A |
1364 |
1356 |
1.15 |
|
|
|
| 10 |
A |
1346 |
1338 |
22.80 |
|
|
|
| 11 |
A |
1310 |
1302 |
60.65 |
|
|
|
| 12 |
A |
1265 |
1257 |
4.13 |
|
|
|
| 13 |
A |
1200 |
1193 |
248.74 |
|
|
|
| 14 |
A |
1097 |
1091 |
13.67 |
|
|
|
| 15 |
A |
975 |
969 |
22.64 |
|
|
|
| 16 |
A |
950 |
945 |
8.25 |
|
|
|
| 17 |
A |
937 |
932 |
35.82 |
|
|
|
| 18 |
A |
850 |
845 |
14.23 |
|
|
|
| 19 |
A |
817 |
812 |
31.98 |
|
|
|
| 20 |
A |
793 |
789 |
46.13 |
|
|
|
| 21 |
A |
704 |
700 |
4.25 |
|
|
|
| 22 |
A |
688 |
684 |
2.56 |
|
|
|
| 23 |
A |
572 |
569 |
2.15 |
|
|
|
| 24 |
A |
436 |
433 |
2.02 |
|
|
|
| 25 |
A |
267 |
266 |
1.11 |
|
|
|
| 26 |
A |
210 |
209 |
7.94 |
|
|
|
| 27 |
A |
93 |
93 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19018.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 18908.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.052 |
-0.793 |
0.045 |
| C2 |
1.292 |
-0.501 |
0.013 |
| C3 |
-0.932 |
0.243 |
0.282 |
| C4 |
1.870 |
0.703 |
-0.149 |
| C5 |
-2.189 |
0.211 |
-0.184 |
| H6 |
1.868 |
-1.431 |
0.124 |
| H7 |
-0.534 |
1.048 |
0.923 |
| H8 |
2.966 |
0.767 |
-0.139 |
| H9 |
1.298 |
1.626 |
-0.313 |
| H10 |
-2.891 |
1.007 |
0.095 |
| H11 |
-2.537 |
-0.609 |
-0.828 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3752 | 1.3789 | 2.4432 | 2.3721 | 2.0254 | 2.0957 | 3.4018 | 2.7926 | 3.3613 | 2.6399 |
C2 | 1.3752 | | 2.3596 | 1.3456 | 3.5583 | 1.1002 | 2.5612 | 2.1055 | 2.1516 | 4.4464 | 3.9209 | C3 | 1.3789 | 2.3596 | | 2.8720 | 1.3416 | 3.2660 | 1.1040 | 3.9550 | 2.6904 | 2.1111 | 2.1291 | C4 | 2.4432 | 1.3456 | 2.8720 | | 4.0893 | 2.1516 | 2.6545 | 1.0975 | 1.0982 | 4.7766 | 4.6479 | C5 | 2.3721 | 3.5583 | 1.3416 | 4.0893 | | 4.3881 | 2.1607 | 5.1851 | 3.7654 | 1.0971 | 1.0987 | H6 | 2.0254 | 1.1002 | 3.2660 | 2.1516 | 4.3881 | | 3.5434 | 2.4707 | 3.1403 | 5.3473 | 4.5812 | H7 | 2.0957 | 2.5612 | 1.1040 | 2.6545 | 2.1607 | 3.5434 | | 3.6677 | 2.2839 | 2.4987 | 3.1345 | H8 | 3.4018 | 2.1055 | 3.9550 | 1.0975 | 5.1851 | 2.4707 | 3.6677 | | 1.8842 | 5.8658 | 5.7135 | H9 | 2.7926 | 2.1516 | 2.6904 | 1.0982 | 3.7654 | 3.1403 | 2.2839 | 1.8842 | | 4.2534 | 4.4679 | H10 | 3.3613 | 4.4464 | 2.1111 | 4.7766 | 1.0971 | 5.3473 | 2.4987 | 5.8658 | 4.2534 | | 1.8942 | H11 | 2.6399 | 3.9209 | 2.1291 | 4.6479 | 1.0987 | 4.5812 | 3.1345 | 5.7135 | 4.4679 | 1.8942 | |
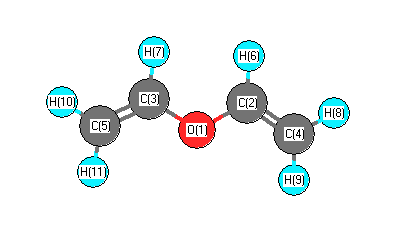 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.782 |
|
O1 |
C2 |
H6 |
109.308 |
| O1 |
C3 |
C5 |
121.365 |
|
O1 |
C3 |
H7 |
114.687 |
| C2 |
O1 |
C3 |
117.909 |
|
C2 |
C4 |
H8 |
118.699 |
| C2 |
C4 |
H9 |
123.069 |
|
C3 |
C5 |
H10 |
119.589 |
| C3 |
C5 |
H11 |
121.175 |
|
C4 |
C2 |
H6 |
122.900 |
| C5 |
C3 |
H7 |
123.856 |
|
H8 |
C4 |
H9 |
118.217 |
| H10 |
C5 |
H11 |
119.228 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.164 |
|
|
|
| 2 |
C |
0.065 |
|
|
|
| 3 |
C |
0.055 |
|
|
|
| 4 |
C |
-0.089 |
|
|
|
| 5 |
C |
-0.045 |
|
|
|
| 6 |
H |
0.016 |
|
|
|
| 7 |
H |
0.022 |
|
|
|
| 8 |
H |
0.037 |
|
|
|
| 9 |
H |
0.035 |
|
|
|
| 10 |
H |
0.036 |
|
|
|
| 11 |
H |
0.033 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.309 |
0.518 |
0.252 |
0.654 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.560 |
-1.531 |
0.491 |
| y |
-1.531 |
-29.269 |
0.846 |
| z |
0.491 |
0.846 |
-32.383 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.266 |
-1.531 |
0.491 |
| y |
-1.531 |
0.703 |
0.846 |
| z |
0.491 |
0.846 |
-3.969 |
|
| Polar |
|---|
| 3z2-r2 | -7.937 |
|---|
| x2-y2 | 1.709 |
|---|
| xy | -1.531 |
|---|
| xz | 0.491 |
|---|
| yz | 0.846 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.519 |
0.347 |
0.549 |
| y |
0.347 |
6.746 |
0.280 |
| z |
0.549 |
0.280 |
4.015 |
<r2> (average value of r
2) Å
2
| <r2> |
132.989 |
| (<r2>)1/2 |
11.532 |
