return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-267.695988
Energy at 298.15K-267.703194
HF Energy-266.845377
Nuclear repulsion energy176.720101
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3144 2996        
2 A' 3068 2924        
3 A' 3064 2921        
4 A' 3054 2911        
5 A' 1810 1725        
6 A' 1510 1439        
7 A' 1489 1419        
8 A' 1427 1360        
9 A' 1395 1329        
10 A' 1381 1316        
11 A' 1215 1158        
12 A' 1136 1083        
13 A' 1053 1004        
14 A' 867 827        
15 A' 787 750        
16 A' 381 363        
17 A' 234 223        
18 A" 3152 3005        
19 A" 3116 2970        
20 A" 1476 1406        
21 A" 1290 1230        
22 A" 1180 1124        
23 A" 1036 988        
24 A" 807 769        
25 A" 345 329        
26 A" 244 232        
27 A" 69 66        

Unscaled Zero Point Vibrational Energy (zpe) 19864.0 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 18932.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.58090 0.09653 0.08549

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.183 -0.255 0.000
C2 -0.692 -0.558 0.000
O3 0.000 0.718 0.000
C4 1.347 0.613 0.000
O5 1.979 -0.420 0.000
H6 -2.758 -1.200 0.000
H7 -2.462 0.327 0.897
H8 -2.462 0.327 -0.897
H9 -0.385 -1.135 -0.893
H10 -0.385 -1.135 0.893
H11 1.785 1.635 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.52152.39003.63544.16541.10591.10511.10512.19152.19154.3957
C21.52151.45142.35162.67472.16342.17242.17241.10611.10613.3087
O32.39001.45141.35112.28283.35922.64902.64902.09202.09202.0073
C43.63542.35161.35111.21134.48773.92343.92342.61792.61791.1117
O54.16542.67472.28281.21134.80094.59154.59152.62642.62642.0640
H61.10592.16343.35924.48774.80091.79551.79552.53602.53605.3554
H71.10512.17242.64903.92344.59151.79551.79443.10652.53904.5336
H81.10512.17242.64903.92344.59151.79551.79442.53903.10654.5336
H92.19151.10612.09202.61792.62642.53603.10652.53901.78543.6304
H102.19151.10612.09202.61792.62642.53602.53903.10651.78543.6304
H114.39573.30872.00731.11172.06405.35544.53364.53363.63043.6304

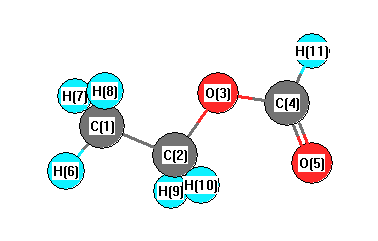
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 106.996 C1 C2 H9 112.059
C1 C2 H10 112.059 C2 C1 H6 109.844
C2 C1 H7 110.597 C2 C1 H8 110.597
C2 O3 C4 114.049 O3 C2 H9 109.021
O3 C2 H10 109.021 O3 C4 O5 125.881
O3 C4 H11 108.799 O5 C4 H11 125.321
H6 C1 H7 108.593 H6 C1 H8 108.593
H7 C1 H8 108.560 H9 C2 H10 107.625
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability