return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-267.671985
Energy at 298.15K-267.679231
HF Energy-266.846495
Nuclear repulsion energy177.070950
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3159 3030 21.85      
2 A' 3090 2965 53.65      
3 A' 3086 2960 32.42      
4 A' 3070 2945 17.83      
5 A' 1836 1762 276.14      
6 A' 1522 1460 6.08      
7 A' 1499 1438 1.94      
8 A' 1441 1383 18.73      
9 A' 1411 1354 1.42      
10 A' 1394 1338 2.98      
11 A' 1240 1189 417.33      
12 A' 1147 1100 8.17      
13 A' 1065 1022 14.56      
14 A' 879 844 11.71      
15 A' 797 764 3.59      
16 A' 386 370 6.18      
17 A' 236 226 6.09      
18 A" 3166 3037 34.71      
19 A" 3134 3006 11.57      
20 A" 1484 1424 4.88      
21 A" 1302 1250 1.46      
22 A" 1190 1141 4.52      
23 A" 1056 1013 0.36      
24 A" 814 781 0.19      
25 A" 344 330 19.95      
26 A" 245 235 2.64      
27 A" 74 71 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 20032.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19219.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.58652 0.09666 0.08570

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.181 -0.258 0.000
C2 -0.691 -0.560 0.000
O3 0.000 0.711 0.000
C4 1.343 0.616 0.000
O5 1.980 -0.410 0.000
H6 -2.755 -1.201 0.000
H7 -2.459 0.324 0.896
H8 -2.459 0.324 -0.896
H9 -0.386 -1.137 -0.891
H10 -0.386 -1.137 0.891
H11 1.776 1.638 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.51992.38603.63034.16321.10441.10381.10382.18792.18794.3874
C21.51991.44672.34962.67522.16132.17022.17021.10441.10443.3043
O32.38601.44671.34642.27513.35352.64542.64542.08722.08722.0035
C43.63032.34961.34641.20754.48293.91703.91702.61842.61841.1098
O54.16322.67522.27511.20754.80074.58744.58742.63072.63072.0578
H61.10442.16133.35354.48294.80071.79321.79322.53202.53205.3472
H71.10382.17022.64543.91704.58741.79321.79143.10222.53574.5238
H81.10382.17022.64543.91704.58741.79321.79142.53573.10224.5238
H92.18791.10442.08722.61842.63072.53203.10222.53571.78283.6286
H102.18791.10442.08722.61842.63072.53202.53573.10221.78283.6286
H114.38743.30432.00351.10982.05785.34724.52384.52383.62863.6286

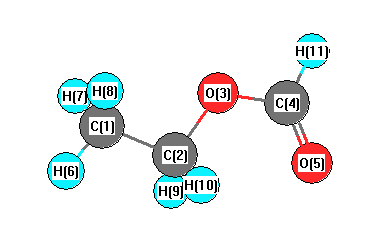
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.060 C1 C2 H9 111.984
C1 C2 H10 111.984 C2 C1 H6 109.880
C2 C1 H7 110.613 C2 C1 H8 110.613
C2 O3 C4 114.491 O3 C2 H9 109.064
O3 C2 H10 109.064 O3 C4 O5 125.874
O3 C4 H11 108.934 O5 C4 H11 125.192
H6 C1 H7 108.599 H6 C1 H8 108.599
H7 C1 H8 108.478 H9 C2 H10 107.631
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability