return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-267.661638
Energy at 298.15K-267.668913
HF Energy-266.847375
Nuclear repulsion energy177.432485
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3171 3033 20.37      
2 A' 3100 2966 57.19      
3 A' 3092 2958 25.43      
4 A' 3079 2946 16.99      
5 A' 1882 1801 293.14      
6 A' 1527 1461 6.26      
7 A' 1502 1437 1.89      
8 A' 1447 1385 18.16      
9 A' 1426 1364 3.09      
10 A' 1400 1340 1.51      
11 A' 1264 1209 442.25      
12 A' 1153 1103 9.04      
13 A' 1076 1029 14.06      
14 A' 891 852 10.80      
15 A' 807 772 4.76      
16 A' 389 372 6.21      
17 A' 238 227 6.13      
18 A" 3177 3039 31.65      
19 A" 3141 3005 13.59      
20 A" 1486 1422 4.95      
21 A" 1310 1253 1.63      
22 A" 1194 1143 4.65      
23 A" 1070 1023 0.40      
24 A" 817 781 0.18      
25 A" 344 329 20.25      
26 A" 246 235 2.76      
27 A" 77 74 0.38      

Unscaled Zero Point Vibrational Energy (zpe) 20152.2 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 19279.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.59099 0.09690 0.08598

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.178 -0.263 0.000
C2 -0.688 -0.561 0.000
O3 0.000 0.706 0.000
C4 1.339 0.618 0.000
O5 1.978 -0.401 0.000
H6 -2.750 -1.207 0.000
H7 -2.458 0.318 0.895
H8 -2.458 0.318 -0.895
H9 -0.384 -1.138 -0.891
H10 -0.384 -1.138 0.891
H11 1.771 1.640 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.52002.38413.62604.15801.10391.10331.10332.18612.18614.3835
C21.52001.44182.34472.66992.16152.16992.16991.10421.10423.2996
O32.38411.44181.34232.26653.35032.64462.64462.08422.08422.0019
C43.62602.34471.34231.20224.47823.91293.91292.61672.61671.1094
O54.15802.66992.26651.20224.79604.58184.58182.62952.62952.0512
H61.10392.16153.35034.47824.79601.79241.79242.52932.52935.3427
H71.10332.16992.64463.91294.58181.79241.79053.10052.53434.5202
H81.10332.16992.64463.91294.58181.79241.79052.53433.10054.5202
H92.18611.10422.08422.61672.62952.52933.10052.53431.78183.6270
H102.18611.10422.08422.61672.62952.52932.53433.10051.78183.6270
H114.38353.29962.00191.10942.05125.34274.52024.52023.62703.6270

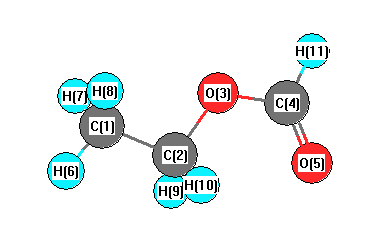
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.178 C1 C2 H9 111.846
C1 C2 H10 111.846 C2 C1 H6 109.912
C2 C1 H7 110.605 C2 C1 H8 110.605
C2 O3 C4 114.696 O3 C2 H9 109.179
O3 C2 H10 109.179 O3 C4 O5 125.853
O3 C4 H11 109.109 O5 C4 H11 125.038
H6 C1 H7 108.597 H6 C1 H8 108.597
H7 C1 H8 108.467 H9 C2 H10 107.571
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability