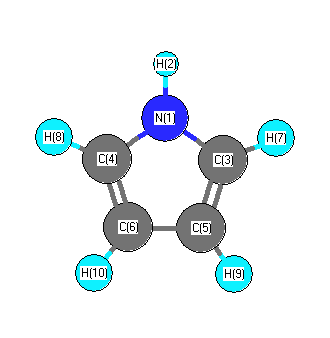
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3548 |
3554 |
43.25 |
|
|
|
| 2 |
A1 |
3189 |
3194 |
0.20 |
|
|
|
| 3 |
A1 |
3167 |
3172 |
6.70 |
|
|
|
| 4 |
A1 |
1449 |
1452 |
7.24 |
|
|
|
| 5 |
A1 |
1375 |
1377 |
2.40 |
|
|
|
| 6 |
A1 |
1138 |
1140 |
2.53 |
|
|
|
| 7 |
A1 |
1055 |
1057 |
5.90 |
|
|
|
| 8 |
A1 |
996 |
997 |
25.01 |
|
|
|
| 9 |
A1 |
866 |
867 |
0.01 |
|
|
|
| 10 |
A2 |
842 |
844 |
0.00 |
|
|
|
| 11 |
A2 |
646 |
647 |
0.00 |
|
|
|
| 12 |
A2 |
609 |
610 |
0.00 |
|
|
|
| 13 |
B1 |
786 |
788 |
4.12 |
|
|
|
| 14 |
B1 |
699 |
700 |
102.33 |
|
|
|
| 15 |
B1 |
621 |
622 |
2.08 |
|
|
|
| 16 |
B1 |
444 |
445 |
54.20 |
|
|
|
| 17 |
B2 |
3183 |
3188 |
7.10 |
|
|
|
| 18 |
B2 |
3156 |
3161 |
3.18 |
|
|
|
| 19 |
B2 |
1519 |
1522 |
1.80 |
|
|
|
| 20 |
B2 |
1408 |
1410 |
4.01 |
|
|
|
| 21 |
B2 |
1253 |
1255 |
1.65 |
|
|
|
| 22 |
B2 |
1116 |
1118 |
1.62 |
|
|
|
| 23 |
B2 |
1024 |
1026 |
23.72 |
|
|
|
| 24 |
B2 |
846 |
848 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17467.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 17495.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.003 |
|
|
|
| 2 |
H |
0.069 |
|
|
|
| 3 |
C |
0.046 |
|
|
|
| 4 |
C |
0.046 |
|
|
|
| 5 |
C |
0.019 |
|
|
|
| 6 |
C |
0.019 |
|
|
|
| 7 |
H |
-0.036 |
|
|
|
| 8 |
H |
-0.036 |
|
|
|
| 9 |
H |
-0.065 |
|
|
|
| 10 |
H |
-0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.981 |
1.981 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.125 |
0.000 |
0.000 |
| y |
0.000 |
-27.341 |
0.000 |
| z |
0.000 |
0.000 |
-23.911 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.499 |
0.000 |
0.000 |
| y |
0.000 |
1.176 |
0.000 |
| z |
0.000 |
0.000 |
6.322 |
|
| Polar |
|---|
| 3z2-r2 | 12.645 |
|---|
| x2-y2 | -5.783 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.112 |
0.000 |
0.000 |
| y |
0.000 |
8.577 |
0.000 |
| z |
0.000 |
0.000 |
8.285 |
<r2> (average value of r
2) Å
2
| <r2> |
87.133 |
| (<r2>)1/2 |
9.335 |