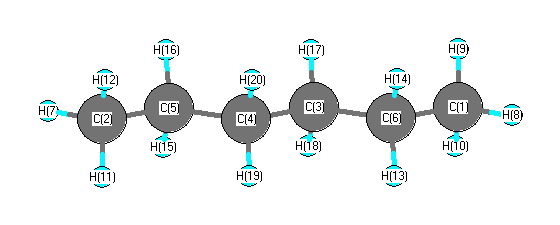
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3047 |
3030 |
0.00 |
|
|
|
| 2 |
Ag |
2962 |
2945 |
0.00 |
|
|
|
| 3 |
Ag |
2948 |
2931 |
0.00 |
|
|
|
| 4 |
Ag |
2933 |
2915 |
0.00 |
|
|
|
| 5 |
Ag |
1431 |
1423 |
0.00 |
|
|
|
| 6 |
Ag |
1414 |
1406 |
0.00 |
|
|
|
| 7 |
Ag |
1408 |
1400 |
0.00 |
|
|
|
| 8 |
Ag |
1351 |
1343 |
0.00 |
|
|
|
| 9 |
Ag |
1345 |
1338 |
0.00 |
|
|
|
| 10 |
Ag |
1278 |
1270 |
0.00 |
|
|
|
| 11 |
Ag |
1125 |
1119 |
0.00 |
|
|
|
| 12 |
Ag |
1067 |
1061 |
0.00 |
|
|
|
| 13 |
Ag |
1014 |
1008 |
0.00 |
|
|
|
| 14 |
Ag |
881 |
876 |
0.00 |
|
|
|
| 15 |
Ag |
360 |
358 |
0.00 |
|
|
|
| 16 |
Ag |
286 |
285 |
0.00 |
|
|
|
| 17 |
Au |
3039 |
3021 |
93.83 |
|
|
|
| 18 |
Au |
2999 |
2981 |
84.72 |
|
|
|
| 19 |
Au |
2972 |
2954 |
0.29 |
|
|
|
| 20 |
Au |
1420 |
1412 |
12.74 |
|
|
|
| 21 |
Au |
1281 |
1274 |
0.73 |
|
|
|
| 22 |
Au |
1198 |
1191 |
0.03 |
|
|
|
| 23 |
Au |
974 |
968 |
0.64 |
|
|
|
| 24 |
Au |
779 |
775 |
1.72 |
|
|
|
| 25 |
Au |
720 |
716 |
5.61 |
|
|
|
| 26 |
Au |
255 |
254 |
0.00 |
|
|
|
| 27 |
Au |
103 |
102 |
0.00 |
|
|
|
| 28 |
Au |
72 |
72 |
0.01 |
|
|
|
| 29 |
Bg |
3038 |
3021 |
0.00 |
|
|
|
| 30 |
Bg |
2988 |
2971 |
0.00 |
|
|
|
| 31 |
Bg |
2959 |
2942 |
0.00 |
|
|
|
| 32 |
Bg |
1421 |
1412 |
0.00 |
|
|
|
| 33 |
Bg |
1275 |
1268 |
0.00 |
|
|
|
| 34 |
Bg |
1254 |
1246 |
0.00 |
|
|
|
| 35 |
Bg |
1153 |
1146 |
0.00 |
|
|
|
| 36 |
Bg |
873 |
868 |
0.00 |
|
|
|
| 37 |
Bg |
728 |
724 |
0.00 |
|
|
|
| 38 |
Bg |
243 |
242 |
0.00 |
|
|
|
| 39 |
Bg |
149 |
148 |
0.00 |
|
|
|
| 40 |
Bu |
3047 |
3030 |
66.21 |
|
|
|
| 41 |
Bu |
2962 |
2944 |
76.00 |
|
|
|
| 42 |
Bu |
2951 |
2934 |
107.60 |
|
|
|
| 43 |
Bu |
2936 |
2919 |
14.44 |
|
|
|
| 44 |
Bu |
1437 |
1428 |
6.15 |
|
|
|
| 45 |
Bu |
1423 |
1415 |
1.46 |
|
|
|
| 46 |
Bu |
1407 |
1399 |
0.58 |
|
|
|
| 47 |
Bu |
1347 |
1339 |
3.37 |
|
|
|
| 48 |
Bu |
1328 |
1320 |
2.25 |
|
|
|
| 49 |
Bu |
1212 |
1205 |
6.49 |
|
|
|
| 50 |
Bu |
1062 |
1056 |
2.28 |
|
|
|
| 51 |
Bu |
1032 |
1026 |
1.35 |
|
|
|
| 52 |
Bu |
879 |
874 |
3.75 |
|
|
|
| 53 |
Bu |
451 |
449 |
0.09 |
|
|
|
| 54 |
Bu |
127 |
126 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40170.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 39937.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.046 |
|
|
|
| 2 |
C |
-0.046 |
|
|
|
| 3 |
C |
-0.039 |
|
|
|
| 4 |
C |
-0.039 |
|
|
|
| 5 |
C |
-0.069 |
|
|
|
| 6 |
C |
-0.069 |
|
|
|
| 7 |
H |
0.025 |
|
|
|
| 8 |
H |
0.025 |
|
|
|
| 9 |
H |
0.030 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.030 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.020 |
|
|
|
| 16 |
H |
0.020 |
|
|
|
| 17 |
H |
0.015 |
|
|
|
| 18 |
H |
0.015 |
|
|
|
| 19 |
H |
0.015 |
|
|
|
| 20 |
H |
0.015 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.949 |
-0.008 |
0.000 |
| y |
-0.008 |
-42.848 |
0.000 |
| z |
0.000 |
0.000 |
-41.253 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.101 |
-0.008 |
0.000 |
| y |
-0.008 |
-1.247 |
0.000 |
| z |
0.000 |
0.000 |
1.146 |
|
| Polar |
|---|
| 3z2-r2 | 2.292 |
|---|
| x2-y2 | 0.899 |
|---|
| xy | -0.008 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.632 |
1.379 |
0.000 |
| y |
1.379 |
12.396 |
0.000 |
| z |
0.000 |
0.000 |
9.392 |
<r2> (average value of r
2) Å
2
| <r2> |
321.725 |
| (<r2>)1/2 |
17.937 |