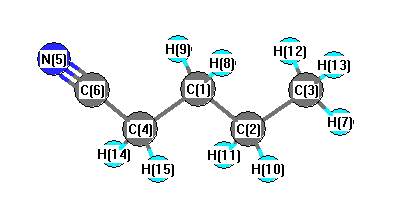
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3022 |
3026 |
34.40 |
|
|
|
| 2 |
A' |
2957 |
2961 |
47.98 |
|
|
|
| 3 |
A' |
2949 |
2954 |
16.95 |
|
|
|
| 4 |
A' |
2945 |
2950 |
3.26 |
|
|
|
| 5 |
A' |
2928 |
2932 |
23.85 |
|
|
|
| 6 |
A' |
2256 |
2260 |
8.52 |
|
|
|
| 7 |
A' |
1446 |
1448 |
3.66 |
|
|
|
| 8 |
A' |
1432 |
1434 |
0.18 |
|
|
|
| 9 |
A' |
1422 |
1424 |
0.95 |
|
|
|
| 10 |
A' |
1402 |
1404 |
2.38 |
|
|
|
| 11 |
A' |
1361 |
1363 |
0.56 |
|
|
|
| 12 |
A' |
1344 |
1346 |
0.31 |
|
|
|
| 13 |
A' |
1307 |
1309 |
3.62 |
|
|
|
| 14 |
A' |
1232 |
1234 |
6.46 |
|
|
|
| 15 |
A' |
1085 |
1086 |
2.21 |
|
|
|
| 16 |
A' |
1029 |
1031 |
0.11 |
|
|
|
| 17 |
A' |
985 |
987 |
0.85 |
|
|
|
| 18 |
A' |
911 |
913 |
0.18 |
|
|
|
| 19 |
A' |
877 |
878 |
1.80 |
|
|
|
| 20 |
A' |
522 |
523 |
1.28 |
|
|
|
| 21 |
A' |
372 |
372 |
0.60 |
|
|
|
| 22 |
A' |
272 |
272 |
0.70 |
|
|
|
| 23 |
A' |
122 |
123 |
4.43 |
|
|
|
| 24 |
A" |
3014 |
3019 |
63.28 |
|
|
|
| 25 |
A" |
2999 |
3004 |
18.95 |
|
|
|
| 26 |
A" |
2975 |
2980 |
0.32 |
|
|
|
| 27 |
A" |
2953 |
2957 |
10.37 |
|
|
|
| 28 |
A" |
1431 |
1433 |
5.53 |
|
|
|
| 29 |
A" |
1285 |
1287 |
0.15 |
|
|
|
| 30 |
A" |
1267 |
1269 |
0.29 |
|
|
|
| 31 |
A" |
1195 |
1196 |
0.04 |
|
|
|
| 32 |
A" |
1094 |
1095 |
0.08 |
|
|
|
| 33 |
A" |
903 |
904 |
0.58 |
|
|
|
| 34 |
A" |
773 |
774 |
0.08 |
|
|
|
| 35 |
A" |
725 |
727 |
3.70 |
|
|
|
| 36 |
A" |
374 |
375 |
0.05 |
|
|
|
| 37 |
A" |
243 |
244 |
0.08 |
|
|
|
| 38 |
A" |
115 |
115 |
0.36 |
|
|
|
| 39 |
A" |
85 |
85 |
3.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27802.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 27847.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.013 |
|
|
|
| 2 |
C |
-0.010 |
|
|
|
| 3 |
C |
0.026 |
|
|
|
| 4 |
C |
0.062 |
|
|
|
| 5 |
N |
-0.082 |
|
|
|
| 6 |
C |
-0.141 |
|
|
|
| 7 |
H |
0.005 |
|
|
|
| 8 |
H |
0.012 |
|
|
|
| 9 |
H |
0.012 |
|
|
|
| 10 |
H |
-0.003 |
|
|
|
| 11 |
H |
-0.003 |
|
|
|
| 12 |
H |
0.011 |
|
|
|
| 13 |
H |
0.011 |
|
|
|
| 14 |
H |
0.044 |
|
|
|
| 15 |
H |
0.044 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.514 |
1.811 |
0.000 |
3.953 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.275 |
-7.545 |
0.000 |
| y |
-7.545 |
-42.207 |
0.000 |
| z |
0.000 |
0.000 |
-37.130 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.606 |
-7.545 |
0.000 |
| y |
-7.545 |
-0.004 |
0.000 |
| z |
0.000 |
0.000 |
7.611 |
|
| Polar |
|---|
| 3z2-r2 | 15.222 |
|---|
| x2-y2 | -5.068 |
|---|
| xy | -7.545 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.221 |
1.963 |
0.000 |
| y |
1.963 |
9.608 |
0.000 |
| z |
0.000 |
0.000 |
7.201 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |