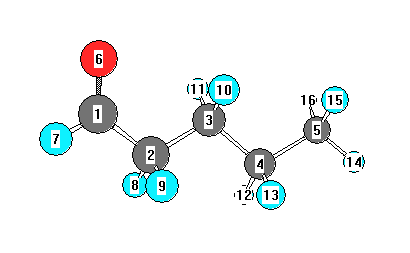
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3052 |
3034 |
29.61 |
|
|
|
| 2 |
A' |
2972 |
2955 |
34.88 |
|
|
|
| 3 |
A' |
2965 |
2948 |
20.06 |
|
|
|
| 4 |
A' |
2946 |
2929 |
28.68 |
|
|
|
| 5 |
A' |
2931 |
2914 |
20.33 |
|
|
|
| 6 |
A' |
2752 |
2736 |
192.75 |
|
|
|
| 7 |
A' |
1766 |
1755 |
141.98 |
|
|
|
| 8 |
A' |
1433 |
1425 |
5.73 |
|
|
|
| 9 |
A' |
1418 |
1409 |
0.33 |
|
|
|
| 10 |
A' |
1409 |
1401 |
1.19 |
|
|
|
| 11 |
A' |
1377 |
1369 |
8.95 |
|
|
|
| 12 |
A' |
1353 |
1346 |
8.84 |
|
|
|
| 13 |
A' |
1349 |
1341 |
1.03 |
|
|
|
| 14 |
A' |
1340 |
1332 |
11.08 |
|
|
|
| 15 |
A' |
1306 |
1298 |
22.70 |
|
|
|
| 16 |
A' |
1220 |
1213 |
2.11 |
|
|
|
| 17 |
A' |
1104 |
1098 |
9.05 |
|
|
|
| 18 |
A' |
1062 |
1056 |
0.51 |
|
|
|
| 19 |
A' |
1025 |
1019 |
1.43 |
|
|
|
| 20 |
A' |
902 |
897 |
2.72 |
|
|
|
| 21 |
A' |
874 |
869 |
14.94 |
|
|
|
| 22 |
A' |
675 |
671 |
10.74 |
|
|
|
| 23 |
A' |
386 |
383 |
1.77 |
|
|
|
| 24 |
A' |
281 |
280 |
3.87 |
|
|
|
| 25 |
A' |
132 |
131 |
4.44 |
|
|
|
| 26 |
A" |
3043 |
3025 |
42.91 |
|
|
|
| 27 |
A" |
3015 |
2997 |
28.04 |
|
|
|
| 28 |
A" |
2978 |
2961 |
7.21 |
|
|
|
| 29 |
A" |
2959 |
2941 |
7.66 |
|
|
|
| 30 |
A" |
1419 |
1410 |
6.63 |
|
|
|
| 31 |
A" |
1273 |
1265 |
0.23 |
|
|
|
| 32 |
A" |
1252 |
1245 |
0.01 |
|
|
|
| 33 |
A" |
1179 |
1173 |
0.00 |
|
|
|
| 34 |
A" |
1103 |
1097 |
0.15 |
|
|
|
| 35 |
A" |
938 |
933 |
1.10 |
|
|
|
| 36 |
A" |
823 |
818 |
0.10 |
|
|
|
| 37 |
A" |
725 |
721 |
3.09 |
|
|
|
| 38 |
A" |
642 |
638 |
3.34 |
|
|
|
| 39 |
A" |
240 |
239 |
0.00 |
|
|
|
| 40 |
A" |
195 |
194 |
0.32 |
|
|
|
| 41 |
A" |
112 |
111 |
0.84 |
|
|
|
| 42 |
A" |
61 |
60 |
1.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29992.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29818.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.126 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
C |
-0.034 |
|
|
|
| 4 |
C |
-0.075 |
|
|
|
| 5 |
C |
-0.046 |
|
|
|
| 6 |
O |
-0.167 |
|
|
|
| 7 |
H |
-0.013 |
|
|
|
| 8 |
H |
0.048 |
|
|
|
| 9 |
H |
0.048 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.031 |
|
|
|
| 12 |
H |
0.020 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
H |
0.027 |
|
|
|
| 15 |
H |
0.034 |
|
|
|
| 16 |
H |
0.034 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.345 |
0.059 |
0.000 |
2.345 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.960 |
3.741 |
0.000 |
| y |
3.741 |
-37.564 |
0.000 |
| z |
0.000 |
0.000 |
-37.400 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.478 |
3.741 |
0.000 |
| y |
3.741 |
3.115 |
0.000 |
| z |
0.000 |
0.000 |
3.362 |
|
| Polar |
|---|
| 3z2-r2 | 6.725 |
|---|
| x2-y2 | -6.396 |
|---|
| xy | 3.741 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.262 |
-0.523 |
0.000 |
| y |
-0.523 |
10.734 |
0.000 |
| z |
0.000 |
0.000 |
7.183 |
<r2> (average value of r
2) Å
2
| <r2> |
254.325 |
| (<r2>)1/2 |
15.948 |