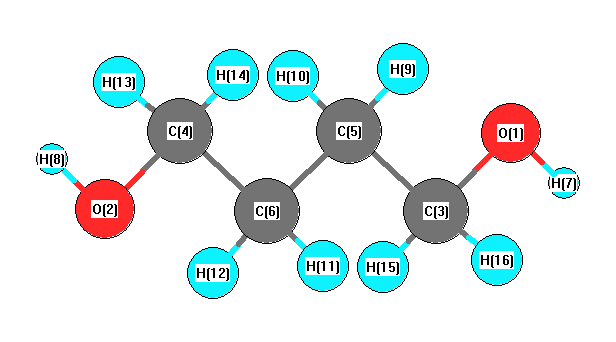
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3676 |
3654 |
0.00 |
|
|
|
| 2 |
Ag |
2956 |
2939 |
0.00 |
|
|
|
| 3 |
Ag |
2870 |
2853 |
0.00 |
|
|
|
| 4 |
Ag |
1448 |
1440 |
0.00 |
|
|
|
| 5 |
Ag |
1417 |
1409 |
0.00 |
|
|
|
| 6 |
Ag |
1407 |
1398 |
0.00 |
|
|
|
| 7 |
Ag |
1327 |
1319 |
0.00 |
|
|
|
| 8 |
Ag |
1230 |
1223 |
0.00 |
|
|
|
| 9 |
Ag |
1065 |
1059 |
0.00 |
|
|
|
| 10 |
Ag |
1047 |
1041 |
0.00 |
|
|
|
| 11 |
Ag |
1014 |
1008 |
0.00 |
|
|
|
| 12 |
Ag |
352 |
350 |
0.00 |
|
|
|
| 13 |
Ag |
327 |
325 |
0.00 |
|
|
|
| 14 |
Au |
3016 |
2999 |
72.77 |
|
|
|
| 15 |
Au |
2897 |
2880 |
126.85 |
|
|
|
| 16 |
Au |
1265 |
1258 |
1.73 |
|
|
|
| 17 |
Au |
1159 |
1152 |
1.70 |
|
|
|
| 18 |
Au |
906 |
901 |
2.22 |
|
|
|
| 19 |
Au |
742 |
738 |
1.24 |
|
|
|
| 20 |
Au |
269 |
268 |
203.23 |
|
|
|
| 21 |
Au |
95 |
95 |
14.02 |
|
|
|
| 22 |
Au |
79 |
78 |
2.21 |
|
|
|
| 23 |
Bg |
2993 |
2976 |
0.00 |
|
|
|
| 24 |
Bg |
2897 |
2881 |
0.00 |
|
|
|
| 25 |
Bg |
1255 |
1247 |
0.00 |
|
|
|
| 26 |
Bg |
1228 |
1221 |
0.00 |
|
|
|
| 27 |
Bg |
1126 |
1120 |
0.00 |
|
|
|
| 28 |
Bg |
790 |
786 |
0.00 |
|
|
|
| 29 |
Bg |
266 |
265 |
0.00 |
|
|
|
| 30 |
Bg |
153 |
152 |
0.00 |
|
|
|
| 31 |
Bu |
3676 |
3654 |
28.73 |
|
|
|
| 32 |
Bu |
2963 |
2946 |
63.17 |
|
|
|
| 33 |
Bu |
2870 |
2853 |
132.77 |
|
|
|
| 34 |
Bu |
1453 |
1445 |
6.28 |
|
|
|
| 35 |
Bu |
1426 |
1418 |
2.97 |
|
|
|
| 36 |
Bu |
1405 |
1397 |
13.04 |
|
|
|
| 37 |
Bu |
1259 |
1252 |
91.40 |
|
|
|
| 38 |
Bu |
1163 |
1156 |
18.02 |
|
|
|
| 39 |
Bu |
1052 |
1046 |
204.38 |
|
|
|
| 40 |
Bu |
983 |
977 |
0.97 |
|
|
|
| 41 |
Bu |
508 |
505 |
34.65 |
|
|
|
| 42 |
Bu |
133 |
133 |
4.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30081.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29906.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.242 |
|
|
|
| 2 |
O |
-0.242 |
|
|
|
| 3 |
C |
0.090 |
|
|
|
| 4 |
C |
0.090 |
|
|
|
| 5 |
C |
-0.058 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
| 9 |
H |
0.026 |
|
|
|
| 10 |
H |
0.026 |
|
|
|
| 11 |
H |
0.026 |
|
|
|
| 12 |
H |
0.026 |
|
|
|
| 13 |
H |
0.010 |
|
|
|
| 14 |
H |
0.010 |
|
|
|
| 15 |
H |
0.010 |
|
|
|
| 16 |
H |
0.010 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.318 |
6.637 |
0.000 |
| y |
6.637 |
-42.703 |
0.000 |
| z |
0.000 |
0.000 |
-38.415 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
13.241 |
6.637 |
0.000 |
| y |
6.637 |
-9.837 |
0.000 |
| z |
0.000 |
0.000 |
-3.404 |
|
| Polar |
|---|
| 3z2-r2 | -6.808 |
|---|
| x2-y2 | 15.385 |
|---|
| xy | 6.637 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.899 |
1.282 |
0.000 |
| y |
1.282 |
9.086 |
0.000 |
| z |
0.000 |
0.000 |
6.939 |
<r2> (average value of r
2) Å
2
| <r2> |
286.782 |
| (<r2>)1/2 |
16.935 |