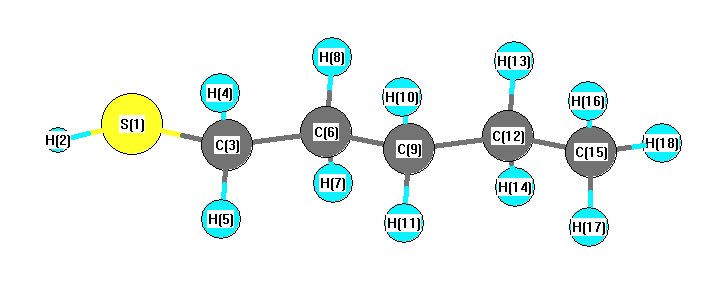
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3051 |
3034 |
30.55 |
|
|
|
| 2 |
A' |
2979 |
2962 |
39.21 |
|
|
|
| 3 |
A' |
2966 |
2948 |
34.42 |
|
|
|
| 4 |
A' |
2959 |
2942 |
41.68 |
|
|
|
| 5 |
A' |
2954 |
2936 |
12.06 |
|
|
|
| 6 |
A' |
2941 |
2924 |
9.19 |
|
|
|
| 7 |
A' |
2611 |
2596 |
10.04 |
|
|
|
| 8 |
A' |
1436 |
1427 |
4.90 |
|
|
|
| 9 |
A' |
1427 |
1418 |
0.18 |
|
|
|
| 10 |
A' |
1414 |
1406 |
1.19 |
|
|
|
| 11 |
A' |
1413 |
1405 |
0.66 |
|
|
|
| 12 |
A' |
1409 |
1401 |
0.05 |
|
|
|
| 13 |
A' |
1352 |
1344 |
1.81 |
|
|
|
| 14 |
A' |
1349 |
1342 |
0.00 |
|
|
|
| 15 |
A' |
1313 |
1306 |
9.49 |
|
|
|
| 16 |
A' |
1246 |
1239 |
4.56 |
|
|
|
| 17 |
A' |
1181 |
1174 |
22.06 |
|
|
|
| 18 |
A' |
1106 |
1100 |
2.36 |
|
|
|
| 19 |
A' |
1063 |
1056 |
0.22 |
|
|
|
| 20 |
A' |
1048 |
1042 |
0.86 |
|
|
|
| 21 |
A' |
1014 |
1008 |
0.14 |
|
|
|
| 22 |
A' |
890 |
884 |
0.88 |
|
|
|
| 23 |
A' |
834 |
829 |
0.86 |
|
|
|
| 24 |
A' |
723 |
719 |
4.20 |
|
|
|
| 25 |
A' |
426 |
424 |
1.23 |
|
|
|
| 26 |
A' |
339 |
337 |
0.23 |
|
|
|
| 27 |
A' |
240 |
239 |
1.05 |
|
|
|
| 28 |
A' |
113 |
112 |
0.88 |
|
|
|
| 29 |
A" |
3043 |
3025 |
53.57 |
|
|
|
| 30 |
A" |
3039 |
3021 |
20.54 |
|
|
|
| 31 |
A" |
3006 |
2989 |
36.23 |
|
|
|
| 32 |
A" |
2992 |
2975 |
4.18 |
|
|
|
| 33 |
A" |
2971 |
2954 |
1.25 |
|
|
|
| 34 |
A" |
1423 |
1415 |
6.60 |
|
|
|
| 35 |
A" |
1279 |
1271 |
0.60 |
|
|
|
| 36 |
A" |
1274 |
1267 |
0.14 |
|
|
|
| 37 |
A" |
1231 |
1224 |
0.36 |
|
|
|
| 38 |
A" |
1168 |
1162 |
0.08 |
|
|
|
| 39 |
A" |
1030 |
1024 |
1.95 |
|
|
|
| 40 |
A" |
926 |
920 |
0.02 |
|
|
|
| 41 |
A" |
813 |
808 |
1.79 |
|
|
|
| 42 |
A" |
742 |
738 |
0.22 |
|
|
|
| 43 |
A" |
725 |
720 |
5.02 |
|
|
|
| 44 |
A" |
253 |
252 |
0.02 |
|
|
|
| 45 |
A" |
177 |
176 |
13.38 |
|
|
|
| 46 |
A" |
150 |
149 |
1.64 |
|
|
|
| 47 |
A" |
100 |
100 |
3.15 |
|
|
|
| 48 |
A" |
67 |
67 |
1.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34101.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33904.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.114 |
|
|
|
| 2 |
H |
0.083 |
|
|
|
| 3 |
C |
-0.140 |
|
|
|
| 4 |
H |
0.061 |
|
|
|
| 5 |
H |
0.061 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
| 9 |
C |
-0.045 |
|
|
|
| 10 |
H |
0.020 |
|
|
|
| 11 |
H |
0.020 |
|
|
|
| 12 |
C |
-0.071 |
|
|
|
| 13 |
H |
0.023 |
|
|
|
| 14 |
H |
0.023 |
|
|
|
| 15 |
C |
-0.047 |
|
|
|
| 16 |
H |
0.031 |
|
|
|
| 17 |
H |
0.031 |
|
|
|
| 18 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.293 |
-1.575 |
0.000 |
1.602 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.813 |
3.129 |
0.000 |
| y |
3.129 |
-51.313 |
0.000 |
| z |
0.000 |
0.000 |
-48.421 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.055 |
3.129 |
0.000 |
| y |
3.129 |
-4.697 |
0.000 |
| z |
0.000 |
0.000 |
-0.358 |
|
| Polar |
|---|
| 3z2-r2 | -0.716 |
|---|
| x2-y2 | 6.501 |
|---|
| xy | 3.129 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.882 |
-2.095 |
0.000 |
| y |
-2.095 |
11.434 |
0.000 |
| z |
0.000 |
0.000 |
8.816 |
<r2> (average value of r
2) Å
2
| <r2> |
415.931 |
| (<r2>)1/2 |
20.394 |