Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.149339 |
| Energy at 298.15K | -235.163696 |
| Nuclear repulsion energy | 254.725200 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3116 |
2989 |
105.02 |
|
|
|
| 2 |
A |
3104 |
2978 |
0.00 |
|
|
|
| 3 |
A |
3088 |
2962 |
9.98 |
|
|
|
| 4 |
A |
3061 |
2937 |
0.00 |
|
|
|
| 5 |
A |
3053 |
2929 |
61.32 |
|
|
|
| 6 |
A |
3047 |
2923 |
0.00 |
|
|
|
| 7 |
A |
1517 |
1456 |
0.00 |
|
|
|
| 8 |
A |
1501 |
1440 |
7.96 |
|
|
|
| 9 |
A |
1480 |
1420 |
0.00 |
|
|
|
| 10 |
A |
1401 |
1344 |
0.00 |
|
|
|
| 11 |
A |
1374 |
1318 |
0.00 |
|
|
|
| 12 |
A |
1336 |
1282 |
0.37 |
|
|
|
| 13 |
A |
1298 |
1245 |
0.00 |
|
|
|
| 14 |
A |
1276 |
1224 |
0.00 |
|
|
|
| 15 |
A |
1261 |
1210 |
0.20 |
|
|
|
| 16 |
A |
1076 |
1032 |
0.31 |
|
|
|
| 17 |
A |
1053 |
1011 |
0.00 |
|
|
|
| 18 |
A |
945 |
907 |
0.00 |
|
|
|
| 19 |
A |
907 |
870 |
0.00 |
|
|
|
| 20 |
A |
827 |
794 |
0.00 |
|
|
|
| 21 |
A |
785 |
753 |
1.51 |
|
|
|
| 22 |
A |
467 |
448 |
0.00 |
|
|
|
| 23 |
A |
427 |
410 |
0.19 |
|
|
|
| 24 |
A |
284 |
273 |
0.00 |
|
|
|
| 25 |
A |
127 |
122 |
0.06 |
|
|
|
| 26 |
B |
3109 |
2983 |
69.56 |
|
|
|
| 27 |
B |
3101 |
2975 |
54.73 |
|
|
|
| 28 |
B |
3090 |
2964 |
35.06 |
|
|
|
| 29 |
B |
3059 |
2935 |
66.95 |
|
|
|
| 30 |
B |
3044 |
2921 |
27.76 |
|
|
|
| 31 |
B |
3044 |
2920 |
14.52 |
|
|
|
| 32 |
B |
1497 |
1436 |
2.69 |
|
|
|
| 33 |
B |
1493 |
1432 |
1.90 |
|
|
|
| 34 |
B |
1480 |
1420 |
0.22 |
|
|
|
| 35 |
B |
1388 |
1331 |
0.08 |
|
|
|
| 36 |
B |
1385 |
1329 |
2.31 |
|
|
|
| 37 |
B |
1385 |
1328 |
2.77 |
|
|
|
| 38 |
B |
1310 |
1256 |
1.45 |
|
|
|
| 39 |
B |
1184 |
1136 |
1.57 |
|
|
|
| 40 |
B |
1153 |
1106 |
0.05 |
|
|
|
| 41 |
B |
1141 |
1095 |
0.03 |
|
|
|
| 42 |
B |
1109 |
1064 |
0.10 |
|
|
|
| 43 |
B |
1026 |
984 |
0.82 |
|
|
|
| 44 |
B |
894 |
858 |
1.07 |
|
|
|
| 45 |
B |
887 |
851 |
0.65 |
|
|
|
| 46 |
B |
763 |
732 |
0.47 |
|
|
|
| 47 |
B |
550 |
528 |
0.26 |
|
|
|
| 48 |
B |
242 |
232 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37572.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 36046.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.222 |
-0.660 |
0.394 |
| C2 |
0.000 |
1.528 |
-0.000 |
| C3 |
1.222 |
-0.660 |
-0.393 |
| C4 |
1.222 |
0.660 |
0.394 |
| C5 |
-0.000 |
-1.528 |
-0.000 |
| C6 |
-1.222 |
0.660 |
-0.393 |
| H7 |
-1.189 |
-0.428 |
1.477 |
| H8 |
-0.267 |
2.192 |
0.845 |
| H9 |
2.159 |
-1.222 |
-0.224 |
| H10 |
1.189 |
0.428 |
1.477 |
| H11 |
0.267 |
-2.192 |
0.845 |
| H12 |
-2.159 |
1.222 |
-0.224 |
| H13 |
-2.159 |
-1.222 |
0.225 |
| H14 |
0.268 |
2.191 |
-0.845 |
| H15 |
1.189 |
-0.428 |
-1.477 |
| H16 |
2.159 |
1.222 |
0.225 |
| H17 |
-0.268 |
-2.191 |
-0.845 |
| H18 |
-1.189 |
0.428 |
-1.477 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.5375 | 2.5679 | 2.7782 | 1.5499 | 1.5373 | 1.1082 | 3.0411 | 3.4831 | 2.8585 | 2.1833 | 2.1916 | 1.1056 | 3.4474 | 3.0605 | 3.8738 | 2.1885 | 2.1640 |
C2 | 2.5375 | | 2.5375 | 1.5499 | 3.0568 | 1.5499 | 2.7244 | 1.1069 | 3.5040 | 2.1920 | 3.8241 | 2.1925 | 3.5040 | 1.1069 | 2.7243 | 2.1925 | 3.8237 | 2.1921 | C3 | 2.5679 | 2.5375 | | 1.5373 | 1.5499 | 2.7784 | 3.0602 | 3.4475 | 1.1056 | 2.1640 | 2.1886 | 3.8739 | 3.4832 | 3.0407 | 1.1082 | 2.1916 | 2.1833 | 2.8589 | C4 | 2.7782 | 1.5499 | 1.5373 | | 2.5375 | 2.5679 | 2.8585 | 2.1833 | 2.1916 | 1.1082 | 3.0411 | 3.4831 | 3.8738 | 2.1885 | 2.1640 | 1.1056 | 3.4474 | 3.0605 | C5 | 1.5499 | 3.0568 | 1.5499 | 2.5375 | | 2.5375 | 2.1920 | 3.8241 | 2.1925 | 2.7244 | 1.1069 | 3.5040 | 2.1925 | 3.8237 | 2.1921 | 3.5040 | 1.1069 | 2.7243 | C6 | 1.5373 | 1.5499 | 2.7784 | 2.5679 | 2.5375 | | 2.1640 | 2.1886 | 3.8739 | 3.0602 | 3.4475 | 1.1056 | 2.1916 | 2.1833 | 2.8589 | 3.4832 | 3.0407 | 1.1082 | H7 | 1.1082 | 2.7244 | 3.0602 | 2.8585 | 2.1920 | 2.1640 | | 2.8480 | 3.8386 | 2.5273 | 2.3727 | 2.5609 | 1.7718 | 3.7912 | 3.7918 | 3.9371 | 3.0577 | 3.0748 | H8 | 3.0411 | 1.1069 | 3.4475 | 2.1833 | 3.8241 | 2.1886 | 2.8480 | | 4.3225 | 2.3727 | 4.4156 | 2.3798 | 3.9518 | 1.7729 | 3.7912 | 2.6857 | 4.6973 | 3.0576 | H9 | 3.4831 | 3.5040 | 1.1056 | 2.1916 | 2.1925 | 3.8739 | 3.8386 | 4.3225 | | 2.5609 | 2.3798 | 4.9623 | 4.3420 | 3.9514 | 1.7718 | 2.4849 | 2.6860 | 3.9375 | H10 | 2.8585 | 2.1920 | 2.1640 | 1.1082 | 2.7244 | 3.0602 | 2.5273 | 2.3727 | 2.5609 | | 2.8480 | 3.8386 | 3.9371 | 3.0577 | 3.0748 | 1.7718 | 3.7912 | 3.7918 | H11 | 2.1833 | 3.8241 | 2.1886 | 3.0411 | 1.1069 | 3.4475 | 2.3727 | 4.4156 | 2.3798 | 2.8480 | | 4.3225 | 2.6857 | 4.6973 | 3.0576 | 3.9518 | 1.7729 | 3.7912 | H12 | 2.1916 | 2.1925 | 3.8739 | 3.4831 | 3.5040 | 1.1056 | 2.5609 | 2.3798 | 4.9623 | 3.8386 | 4.3225 | | 2.4849 | 2.6860 | 3.9375 | 4.3420 | 3.9514 | 1.7718 | H13 | 1.1056 | 3.5040 | 3.4832 | 3.8738 | 2.1925 | 2.1916 | 1.7718 | 3.9518 | 4.3420 | 3.9371 | 2.6857 | 2.4849 | | 4.3225 | 3.8391 | 4.9623 | 2.3798 | 2.5609 | H14 | 3.4474 | 1.1069 | 3.0407 | 2.1885 | 3.8237 | 2.1833 | 3.7912 | 1.7729 | 3.9514 | 3.0577 | 4.6973 | 2.6860 | 4.3225 | | 2.8474 | 2.3798 | 4.4146 | 2.3725 | H15 | 3.0605 | 2.7243 | 1.1082 | 2.1640 | 2.1921 | 2.8589 | 3.7918 | 3.7912 | 1.7718 | 3.0748 | 3.0576 | 3.9375 | 3.8391 | 2.8474 | | 2.5609 | 2.3725 | 2.5280 | H16 | 3.8738 | 2.1925 | 2.1916 | 1.1056 | 3.5040 | 3.4832 | 3.9371 | 2.6857 | 2.4849 | 1.7718 | 3.9518 | 4.3420 | 4.9623 | 2.3798 | 2.5609 | | 4.3225 | 3.8391 | H17 | 2.1885 | 3.8237 | 2.1833 | 3.4474 | 1.1069 | 3.0407 | 3.0577 | 4.6973 | 2.6860 | 3.7912 | 1.7729 | 3.9514 | 2.3798 | 4.4146 | 2.3725 | 4.3225 | | 2.8474 | H18 | 2.1640 | 2.1921 | 2.8589 | 3.0605 | 2.7243 | 1.1082 | 3.0748 | 3.0576 | 3.9375 | 3.7918 | 3.7912 | 1.7718 | 2.5609 | 2.3725 | 2.5280 | 3.8391 | 2.8474 | |
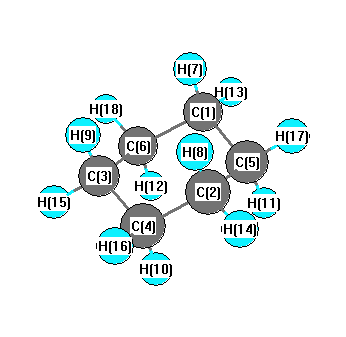 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
C2 |
55.938 |
|
C1 |
C5 |
H11 |
109.396 |
| C1 |
C5 |
H17 |
109.800 |
|
C1 |
C6 |
C3 |
65.899 |
| C1 |
C6 |
H12 |
110.991 |
|
C1 |
C6 |
H18 |
108.682 |
| C2 |
C4 |
C3 |
110.558 |
|
C2 |
C4 |
H10 |
109.998 |
| C2 |
C4 |
H16 |
110.187 |
|
C2 |
C5 |
H11 |
126.805 |
| C2 |
C5 |
H17 |
126.772 |
|
C3 |
C4 |
H10 |
108.684 |
| C3 |
C4 |
H16 |
110.991 |
|
C3 |
C6 |
H12 |
170.844 |
| C3 |
C6 |
H18 |
82.777 |
|
C4 |
C2 |
C5 |
55.938 |
| C4 |
C2 |
H8 |
109.396 |
|
C4 |
C2 |
H14 |
109.800 |
| C4 |
C3 |
C6 |
65.899 |
|
C4 |
C3 |
H9 |
110.991 |
| C4 |
C3 |
H15 |
108.682 |
|
C5 |
C1 |
C6 |
110.558 |
| C5 |
C1 |
H7 |
109.998 |
|
C5 |
C1 |
H13 |
110.187 |
| C5 |
C2 |
H8 |
126.805 |
|
C5 |
C2 |
H14 |
126.772 |
| C6 |
C1 |
H7 |
108.684 |
|
C6 |
C1 |
H13 |
110.991 |
| C6 |
C3 |
H9 |
170.844 |
|
C6 |
C3 |
H15 |
82.777 |
| H7 |
C1 |
H13 |
106.323 |
|
H8 |
C2 |
H14 |
106.423 |
| H9 |
C3 |
H15 |
106.323 |
|
H10 |
C4 |
H16 |
106.323 |
| H11 |
C5 |
H17 |
106.423 |
|
H12 |
C6 |
H18 |
106.323 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
