return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O3 (1,3,5-Trioxane)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-342.681217
Energy at 298.15K 
HF Energy-341.675427
Nuclear repulsion energy270.362705
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3188 3059 10.56      
2 A1 2989 2867 192.84      
3 A1 1527 1465 3.13      
4 A1 1265 1213 27.22      
5 A1 1008 967 83.32      
6 A1 774 742 1.11      
7 A1 477 458 19.95      
8 A2 1409 1351 0.00      
9 A2 1263 1212 0.00      
10 A2 1040 998 0.00      
11 E 3184 3055 37.86      
12 E 2970 2849 23.64      
13 E 1510 1449 1.34      
13 E 1510 1449 1.34      
14 E 1464 1404 46.68      
15 E 1345 1291 2.47      
15 E 1345 1291 2.47      
16 E 1224 1174 254.11      
17 E 1097 1053 34.90      
17 E 1097 1053 34.91      
18 E 997 957 28.62      
18 E 997 957 28.62      
19 E 539 517 7.96      
19 E 539 517 7.96      
20 E 303 291 0.21      
20 E 303 291 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 22102.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 21204.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.17578 0.17578 0.09791

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.324 0.182
C2 -1.147 -0.662 0.182
C3 1.147 -0.662 0.182
O4 -1.166 0.673 -0.269
O5 1.166 0.673 -0.269
O6 0.000 -1.346 -0.269
H7 0.000 2.339 -0.239
H8 0.000 1.363 1.297
H9 -2.025 -1.169 -0.239
H10 -1.180 -0.681 1.297
H11 2.025 -1.169 -0.239
H12 1.180 -0.681 1.297

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 O6 H7 H8 H9 H10 H11 H12
C12.29362.29361.40921.40922.70781.09841.11533.23982.58023.23982.5802
C22.29362.29361.40922.70781.40923.23982.58021.09841.11533.23982.5802
C32.29362.29362.70781.40921.40923.23982.58023.23982.58021.09841.1153
O41.40921.40922.70782.33102.33102.03322.07012.03322.07013.68453.1284
O51.40922.70781.40922.33102.33102.03322.07013.68453.12842.03322.0701
O62.70781.40921.40922.33102.33103.68453.12842.03322.07012.03322.0701
H71.09843.23983.23982.03322.03323.68451.81974.05053.58774.05053.5877
H81.11532.58022.58022.07012.07013.12841.81973.58772.36023.58772.3602
H93.23981.09843.23982.03323.68452.03324.05053.58771.81974.05053.5877
H102.58021.11532.58022.07013.12842.07013.58772.36021.81973.58772.3602
H113.23983.23981.09843.68452.03322.03324.05053.58774.05053.58771.8197
H122.58022.58021.11533.12842.07012.07013.58772.36023.58772.36021.8197

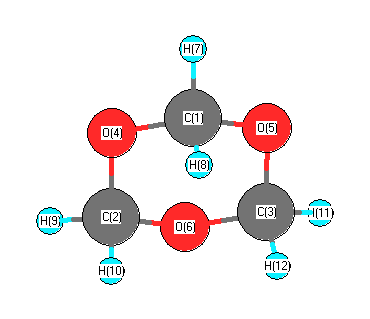
picture of 1,3,5-Trioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O4 C2 108.932 C1 O5 C3 108.932
C2 O6 C3 108.932 O4 C1 O5 111.595
O4 C1 H7 107.706 O4 C1 H8 109.617
O4 C2 O6 111.595 O4 C2 H9 107.706
O4 C2 H10 109.617 O5 C1 H7 107.706
O5 C1 H8 109.617 O5 C3 O6 111.595
O5 C3 H11 107.706 O5 C3 H12 109.617
O6 C2 H9 107.706 O6 C2 H10 109.617
O6 C3 H11 107.706 O6 C3 H12 109.617
H7 C1 H8 110.571 H9 C2 H10 110.571
H11 C3 H12 110.571
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability