Jump to
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.755684 |
| Energy at 298.15K | -251.769868 |
| Nuclear repulsion energy | 256.255917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3369 |
3374 |
2.82 |
|
|
|
| 2 |
A' |
2989 |
2994 |
86.67 |
|
|
|
| 3 |
A' |
2977 |
2982 |
13.90 |
|
|
|
| 4 |
A' |
2972 |
2977 |
68.06 |
|
|
|
| 5 |
A' |
2942 |
2947 |
16.46 |
|
|
|
| 6 |
A' |
2925 |
2930 |
29.19 |
|
|
|
| 7 |
A' |
2793 |
2798 |
173.53 |
|
|
|
| 8 |
A' |
1437 |
1439 |
0.82 |
|
|
|
| 9 |
A' |
1424 |
1426 |
5.45 |
|
|
|
| 10 |
A' |
1414 |
1416 |
5.49 |
|
|
|
| 11 |
A' |
1371 |
1373 |
5.50 |
|
|
|
| 12 |
A' |
1332 |
1334 |
0.82 |
|
|
|
| 13 |
A' |
1265 |
1267 |
0.39 |
|
|
|
| 14 |
A' |
1243 |
1245 |
3.42 |
|
|
|
| 15 |
A' |
1123 |
1125 |
4.13 |
|
|
|
| 16 |
A' |
1037 |
1039 |
6.78 |
|
|
|
| 17 |
A' |
1016 |
1018 |
2.87 |
|
|
|
| 18 |
A' |
890 |
891 |
3.29 |
|
|
|
| 19 |
A' |
841 |
843 |
9.61 |
|
|
|
| 20 |
A' |
798 |
799 |
1.06 |
|
|
|
| 21 |
A' |
748 |
749 |
75.10 |
|
|
|
| 22 |
A' |
533 |
534 |
15.29 |
|
|
|
| 23 |
A' |
425 |
426 |
5.83 |
|
|
|
| 24 |
A' |
391 |
392 |
1.26 |
|
|
|
| 25 |
A' |
243 |
243 |
1.04 |
|
|
|
| 26 |
A" |
2984 |
2989 |
25.44 |
|
|
|
| 27 |
A" |
2973 |
2978 |
90.76 |
|
|
|
| 28 |
A" |
2942 |
2947 |
24.14 |
|
|
|
| 29 |
A" |
2784 |
2789 |
33.71 |
|
|
|
| 30 |
A" |
1436 |
1438 |
0.15 |
|
|
|
| 31 |
A" |
1421 |
1423 |
0.04 |
|
|
|
| 32 |
A" |
1404 |
1406 |
1.26 |
|
|
|
| 33 |
A" |
1329 |
1331 |
0.58 |
|
|
|
| 34 |
A" |
1314 |
1316 |
13.99 |
|
|
|
| 35 |
A" |
1294 |
1296 |
19.42 |
|
|
|
| 36 |
A" |
1248 |
1250 |
3.54 |
|
|
|
| 37 |
A" |
1145 |
1147 |
4.39 |
|
|
|
| 38 |
A" |
1125 |
1126 |
4.74 |
|
|
|
| 39 |
A" |
1084 |
1086 |
18.87 |
|
|
|
| 40 |
A" |
1034 |
1036 |
2.60 |
|
|
|
| 41 |
A" |
950 |
951 |
0.45 |
|
|
|
| 42 |
A" |
855 |
856 |
0.06 |
|
|
|
| 43 |
A" |
794 |
796 |
0.93 |
|
|
|
| 44 |
A" |
440 |
440 |
0.81 |
|
|
|
| 45 |
A" |
239 |
239 |
0.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33646.0 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 33699.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.637 |
1.344 |
0.000 |
| H2 |
0.561 |
2.452 |
0.000 |
| H3 |
1.724 |
1.102 |
0.000 |
| C4 |
-0.013 |
0.755 |
1.275 |
| C5 |
-0.013 |
0.755 |
-1.275 |
| C6 |
-0.013 |
-0.788 |
-1.223 |
| C7 |
-0.013 |
-0.788 |
1.223 |
| N8 |
-0.710 |
-1.238 |
0.000 |
| H9 |
-0.766 |
-2.266 |
0.000 |
| H10 |
0.523 |
1.102 |
2.183 |
| H11 |
0.523 |
1.102 |
-2.183 |
| H12 |
-1.064 |
1.106 |
1.354 |
| H13 |
-1.064 |
1.106 |
-1.354 |
| H14 |
1.051 |
-1.150 |
-1.275 |
| H15 |
1.051 |
-1.150 |
1.275 |
| H16 |
-0.544 |
-1.206 |
-2.103 |
| H17 |
-0.544 |
-1.206 |
2.103 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1104 | 1.1139 | 1.5474 | 1.5474 | 2.5424 | 2.5424 | 2.9126 | 3.8733 | 2.1996 | 2.1996 | 2.1875 | 2.1875 | 2.8320 | 2.8320 | 3.5100 | 3.5100 |
H2 | 1.1104 | | 1.7820 | 2.1989 | 2.1989 | 3.5104 | 3.5104 | 3.9032 | 4.9013 | 2.5672 | 2.5672 | 2.5077 | 2.5077 | 3.8527 | 3.8527 | 4.3618 | 4.3618 | H3 | 1.1139 | 1.7820 | | 2.1823 | 2.1823 | 2.8432 | 2.8432 | 3.3764 | 4.1886 | 2.4919 | 2.4919 | 3.1001 | 3.1001 | 2.6743 | 2.6743 | 3.8590 | 3.8590 | C4 | 1.5474 | 2.1989 | 2.1823 | | 2.5492 | 2.9354 | 1.5436 | 2.4663 | 3.3641 | 1.1106 | 3.5163 | 1.1116 | 2.8528 | 3.3558 | 2.1821 | 3.9414 | 2.1938 | C5 | 1.5474 | 2.1989 | 2.1823 | 2.5492 | | 1.5436 | 2.9354 | 2.4663 | 3.3641 | 3.5163 | 1.1106 | 2.8528 | 1.1116 | 2.1821 | 3.3558 | 2.1938 | 3.9414 | C6 | 2.5424 | 3.5104 | 2.8432 | 2.9354 | 1.5436 | | 2.4453 | 1.4775 | 2.0608 | 3.9319 | 2.1869 | 3.3663 | 2.1703 | 1.1250 | 2.7387 | 1.1097 | 3.3936 | C7 | 2.5424 | 3.5104 | 2.8432 | 1.5436 | 2.9354 | 2.4453 | | 1.4775 | 2.0608 | 2.1869 | 3.9319 | 2.1703 | 3.3663 | 2.7387 | 1.1250 | 3.3936 | 1.1097 | N8 | 2.9126 | 3.9032 | 3.3764 | 2.4663 | 2.4663 | 1.4775 | 1.4775 | | 1.0291 | 3.4299 | 3.4299 | 2.7304 | 2.7304 | 2.1753 | 2.1753 | 2.1098 | 2.1098 | H9 | 3.8733 | 4.9013 | 4.1886 | 3.3641 | 3.3641 | 2.0608 | 2.0608 | 1.0291 | | 4.2158 | 4.2158 | 3.6459 | 3.6459 | 2.4841 | 2.4841 | 2.3655 | 2.3655 | H10 | 2.1996 | 2.5672 | 2.4919 | 1.1106 | 3.5163 | 3.9319 | 2.1869 | 3.4299 | 4.2158 | | 4.3665 | 1.7911 | 3.8771 | 4.1607 | 2.4856 | 4.9837 | 2.5440 | H11 | 2.1996 | 2.5672 | 2.4919 | 3.5163 | 1.1106 | 2.1869 | 3.9319 | 3.4299 | 4.2158 | 4.3665 | | 3.8771 | 1.7911 | 2.4856 | 4.1607 | 2.5440 | 4.9837 | H12 | 2.1875 | 2.5077 | 3.1001 | 1.1116 | 2.8528 | 3.3663 | 2.1703 | 2.7304 | 3.6459 | 1.7911 | 3.8771 | | 2.7079 | 4.0592 | 3.0940 | 4.1913 | 2.4855 | H13 | 2.1875 | 2.5077 | 3.1001 | 2.8528 | 1.1116 | 2.1703 | 3.3663 | 2.7304 | 3.6459 | 3.8771 | 1.7911 | 2.7079 | | 3.0940 | 4.0592 | 2.4855 | 4.1913 | H14 | 2.8320 | 3.8527 | 2.6743 | 3.3558 | 2.1821 | 1.1250 | 2.7387 | 2.1753 | 2.4841 | 4.1607 | 2.4856 | 4.0592 | 3.0940 | | 2.5497 | 1.7975 | 3.7356 | H15 | 2.8320 | 3.8527 | 2.6743 | 2.1821 | 3.3558 | 2.7387 | 1.1250 | 2.1753 | 2.4841 | 2.4856 | 4.1607 | 3.0940 | 4.0592 | 2.5497 | | 3.7356 | 1.7975 | H16 | 3.5100 | 4.3618 | 3.8590 | 3.9414 | 2.1938 | 1.1097 | 3.3936 | 2.1098 | 2.3655 | 4.9837 | 2.5440 | 4.1913 | 2.4855 | 1.7975 | 3.7356 | | 4.2060 | H17 | 3.5100 | 4.3618 | 3.8590 | 2.1938 | 3.9414 | 3.3936 | 1.1097 | 2.1098 | 2.3655 | 2.5440 | 4.9837 | 2.4855 | 4.1913 | 3.7356 | 1.7975 | 4.2060 | |
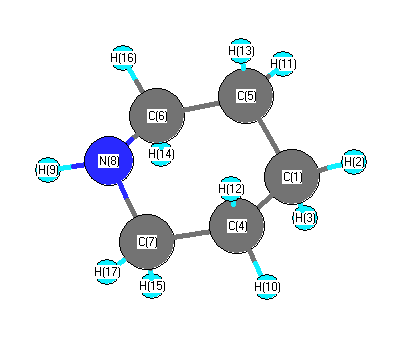 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.671 |
|
C1 |
C4 |
H10 |
110.616 |
| C1 |
C4 |
H12 |
109.623 |
|
C1 |
C5 |
C6 |
110.671 |
| C1 |
C5 |
H11 |
110.616 |
|
C1 |
C5 |
H13 |
109.623 |
| H2 |
C1 |
H3 |
106.476 |
|
H2 |
C1 |
C4 |
110.574 |
| H2 |
C1 |
C5 |
110.574 |
|
H3 |
C1 |
C4 |
109.090 |
| H3 |
C1 |
C5 |
109.090 |
|
C4 |
C1 |
C5 |
110.914 |
| C4 |
C7 |
N8 |
109.422 |
|
C4 |
C7 |
H15 |
108.693 |
| C4 |
C7 |
H17 |
110.477 |
|
C5 |
C6 |
N8 |
109.422 |
| C5 |
C6 |
H14 |
108.693 |
|
C5 |
C6 |
H16 |
110.477 |
| C6 |
C5 |
H11 |
109.883 |
|
C6 |
C5 |
H13 |
108.552 |
| C6 |
N8 |
C7 |
111.683 |
|
C6 |
N8 |
H9 |
109.286 |
| C7 |
C4 |
H10 |
109.883 |
|
C7 |
C4 |
H12 |
108.552 |
| C7 |
N8 |
H9 |
109.286 |
|
N8 |
C6 |
H14 |
112.705 |
| N8 |
C6 |
H16 |
108.428 |
|
N8 |
C7 |
H15 |
112.705 |
| N8 |
C7 |
H17 |
108.428 |
|
H10 |
C4 |
H12 |
107.415 |
| H11 |
C5 |
H13 |
107.415 |
|
H14 |
C6 |
H16 |
107.096 |
| H15 |
C7 |
H17 |
107.096 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.024 |
|
|
|
| 2 |
H |
-0.020 |
|
|
|
| 3 |
H |
-0.015 |
|
|
|
| 4 |
C |
0.049 |
|
|
|
| 5 |
C |
0.049 |
|
|
|
| 6 |
C |
0.081 |
|
|
|
| 7 |
C |
0.081 |
|
|
|
| 8 |
N |
-0.150 |
|
|
|
| 9 |
H |
0.058 |
|
|
|
| 10 |
H |
-0.023 |
|
|
|
| 11 |
H |
-0.023 |
|
|
|
| 12 |
H |
-0.008 |
|
|
|
| 13 |
H |
-0.008 |
|
|
|
| 14 |
H |
-0.029 |
|
|
|
| 15 |
H |
-0.029 |
|
|
|
| 16 |
H |
-0.019 |
|
|
|
| 17 |
H |
-0.019 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.485 |
-0.514 |
0.000 |
0.707 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.525 |
0.144 |
0.000 |
| y |
0.144 |
-37.113 |
0.000 |
| z |
0.000 |
0.000 |
-38.830 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.554 |
0.144 |
0.000 |
| y |
0.144 |
3.064 |
0.000 |
| z |
0.000 |
0.000 |
0.490 |
|
| Polar |
|---|
| 3z2-r2 | 0.979 |
|---|
| x2-y2 | -4.412 |
|---|
| xy | 0.144 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.655 |
0.307 |
0.000 |
| y |
0.307 |
9.801 |
0.000 |
| z |
0.000 |
0.000 |
10.308 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.754983 |
| Energy at 298.15K | -251.769101 |
| Nuclear repulsion energy | 255.627120 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3324 |
3329 |
5.37 |
|
|
|
| 2 |
A' |
2979 |
2984 |
70.56 |
|
|
|
| 3 |
A' |
2973 |
2978 |
53.74 |
|
|
|
| 4 |
A' |
2965 |
2970 |
69.15 |
|
|
|
| 5 |
A' |
2924 |
2929 |
45.52 |
|
|
|
| 6 |
A' |
2919 |
2924 |
58.97 |
|
|
|
| 7 |
A' |
2918 |
2923 |
0.98 |
|
|
|
| 8 |
A' |
1432 |
1435 |
1.28 |
|
|
|
| 9 |
A' |
1414 |
1417 |
5.23 |
|
|
|
| 10 |
A' |
1414 |
1416 |
2.95 |
|
|
|
| 11 |
A' |
1345 |
1348 |
7.79 |
|
|
|
| 12 |
A' |
1329 |
1331 |
0.49 |
|
|
|
| 13 |
A' |
1291 |
1293 |
0.53 |
|
|
|
| 14 |
A' |
1235 |
1237 |
7.83 |
|
|
|
| 15 |
A' |
1149 |
1151 |
3.42 |
|
|
|
| 16 |
A' |
1014 |
1016 |
1.00 |
|
|
|
| 17 |
A' |
980 |
982 |
6.01 |
|
|
|
| 18 |
A' |
899 |
900 |
18.73 |
|
|
|
| 19 |
A' |
832 |
833 |
23.69 |
|
|
|
| 20 |
A' |
792 |
794 |
5.43 |
|
|
|
| 21 |
A' |
746 |
747 |
80.88 |
|
|
|
| 22 |
A' |
537 |
538 |
1.38 |
|
|
|
| 23 |
A' |
425 |
426 |
0.95 |
|
|
|
| 24 |
A' |
379 |
379 |
4.49 |
|
|
|
| 25 |
A' |
235 |
235 |
3.65 |
|
|
|
| 26 |
A" |
2977 |
2982 |
48.31 |
|
|
|
| 27 |
A" |
2962 |
2967 |
84.06 |
|
|
|
| 28 |
A" |
2919 |
2924 |
3.29 |
|
|
|
| 29 |
A" |
2913 |
2918 |
40.95 |
|
|
|
| 30 |
A" |
1441 |
1443 |
4.77 |
|
|
|
| 31 |
A" |
1410 |
1412 |
2.34 |
|
|
|
| 32 |
A" |
1402 |
1404 |
0.45 |
|
|
|
| 33 |
A" |
1330 |
1332 |
5.92 |
|
|
|
| 34 |
A" |
1318 |
1320 |
2.89 |
|
|
|
| 35 |
A" |
1299 |
1301 |
0.80 |
|
|
|
| 36 |
A" |
1255 |
1257 |
1.21 |
|
|
|
| 37 |
A" |
1167 |
1169 |
9.71 |
|
|
|
| 38 |
A" |
1086 |
1088 |
1.79 |
|
|
|
| 39 |
A" |
1065 |
1067 |
20.22 |
|
|
|
| 40 |
A" |
1033 |
1035 |
4.14 |
|
|
|
| 41 |
A" |
918 |
920 |
2.97 |
|
|
|
| 42 |
A" |
843 |
845 |
0.02 |
|
|
|
| 43 |
A" |
779 |
780 |
0.30 |
|
|
|
| 44 |
A" |
445 |
446 |
0.62 |
|
|
|
| 45 |
A" |
226 |
227 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33618.8 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 33672.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.643 |
1.341 |
0.000 |
| H2 |
0.573 |
2.449 |
0.000 |
| H3 |
1.729 |
1.095 |
0.000 |
| C4 |
-0.008 |
0.753 |
1.275 |
| C5 |
-0.008 |
0.753 |
-1.275 |
| C6 |
-0.008 |
-0.797 |
-1.226 |
| C7 |
-0.008 |
-0.797 |
1.226 |
| N8 |
-0.609 |
-1.371 |
0.000 |
| H9 |
-1.617 |
-1.146 |
0.000 |
| H10 |
0.520 |
1.109 |
2.186 |
| H11 |
0.520 |
1.109 |
-2.186 |
| H12 |
-1.060 |
1.113 |
1.356 |
| H13 |
-1.060 |
1.113 |
-1.356 |
| H14 |
1.042 |
-1.164 |
-1.284 |
| H15 |
1.042 |
-1.164 |
1.284 |
| H16 |
-0.540 |
-1.218 |
-2.104 |
| H17 |
-0.540 |
-1.218 |
2.104 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1104 | 1.1141 | 1.5477 | 1.5477 | 2.5490 | 2.5490 | 2.9874 | 3.3605 | 2.2018 | 2.2018 | 2.1885 | 2.1885 | 2.8432 | 2.8432 | 3.5183 | 3.5183 |
H2 | 1.1104 | | 1.7808 | 2.2005 | 2.2005 | 3.5183 | 3.5183 | 3.9996 | 4.2103 | 2.5650 | 2.5650 | 2.5085 | 2.5085 | 3.8633 | 3.8633 | 4.3727 | 4.3727 | H3 | 1.1141 | 1.7808 | | 2.1821 | 2.1821 | 2.8459 | 2.8459 | 3.3981 | 4.0271 | 2.4981 | 2.4981 | 3.1012 | 3.1012 | 2.6874 | 2.6874 | 3.8637 | 3.8637 | C4 | 1.5477 | 2.2005 | 2.1821 | | 2.5500 | 2.9421 | 1.5505 | 2.5494 | 2.7964 | 1.1120 | 3.5193 | 1.1142 | 2.8563 | 3.3653 | 2.1860 | 3.9481 | 2.2038 | C5 | 1.5477 | 2.2005 | 2.1821 | 2.5500 | | 1.5505 | 2.9421 | 2.5494 | 2.7964 | 3.5193 | 1.1120 | 2.8563 | 1.1142 | 2.1860 | 3.3653 | 2.2038 | 3.9481 | C6 | 2.5490 | 3.5183 | 2.8459 | 2.9421 | 1.5505 | | 2.4517 | 1.4810 | 2.0523 | 3.9436 | 2.1984 | 3.3793 | 2.1839 | 1.1142 | 2.7451 | 1.1100 | 3.3984 | C7 | 2.5490 | 3.5183 | 2.8459 | 1.5505 | 2.9421 | 2.4517 | | 1.4810 | 2.0523 | 2.1984 | 3.9436 | 2.1839 | 3.3793 | 2.7451 | 1.1142 | 3.3984 | 1.1100 | N8 | 2.9874 | 3.9996 | 3.3981 | 2.5494 | 2.5494 | 1.4810 | 1.4810 | | 1.0330 | 3.4937 | 3.4937 | 2.8663 | 2.8663 | 2.1012 | 2.1012 | 2.1108 | 2.1108 | H9 | 3.3605 | 4.2103 | 4.0271 | 2.7964 | 2.7964 | 2.0523 | 2.0523 | 1.0330 | | 3.7991 | 3.7991 | 2.6934 | 2.6934 | 2.9525 | 2.9525 | 2.3646 | 2.3646 | H10 | 2.2018 | 2.5650 | 2.4981 | 1.1120 | 3.5193 | 3.9436 | 2.1984 | 3.4937 | 3.7991 | | 4.3723 | 1.7849 | 3.8788 | 4.1806 | 2.5006 | 4.9947 | 2.5589 | H11 | 2.2018 | 2.5650 | 2.4981 | 3.5193 | 1.1120 | 2.1984 | 3.9436 | 3.4937 | 3.7991 | 4.3723 | | 3.8788 | 1.7849 | 2.5006 | 4.1806 | 2.5589 | 4.9947 | H12 | 2.1885 | 2.5085 | 3.1012 | 1.1142 | 2.8563 | 3.3793 | 2.1839 | 2.8663 | 2.6934 | 1.7849 | 3.8788 | | 2.7124 | 4.0708 | 3.0996 | 4.2047 | 2.5030 | H13 | 2.1885 | 2.5085 | 3.1012 | 2.8563 | 1.1142 | 2.1839 | 3.3793 | 2.8663 | 2.6934 | 3.8788 | 1.7849 | 2.7124 | | 3.0996 | 4.0708 | 2.5030 | 4.2047 | H14 | 2.8432 | 3.8633 | 2.6874 | 3.3653 | 2.1860 | 1.1142 | 2.7451 | 2.1012 | 2.9525 | 4.1806 | 2.5006 | 4.0708 | 3.0996 | | 2.5673 | 1.7831 | 3.7394 | H15 | 2.8432 | 3.8633 | 2.6874 | 2.1860 | 3.3653 | 2.7451 | 1.1142 | 2.1012 | 2.9525 | 2.5006 | 4.1806 | 3.0996 | 4.0708 | 2.5673 | | 3.7394 | 1.7831 | H16 | 3.5183 | 4.3727 | 3.8637 | 3.9481 | 2.2038 | 1.1100 | 3.3984 | 2.1108 | 2.3646 | 4.9947 | 2.5589 | 4.2047 | 2.5030 | 1.7831 | 3.7394 | | 4.2082 | H17 | 3.5183 | 4.3727 | 3.8637 | 2.2038 | 3.9481 | 3.3984 | 1.1100 | 2.1108 | 2.3646 | 2.5589 | 4.9947 | 2.5030 | 4.2047 | 3.7394 | 1.7831 | 4.2082 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.716 |
|
C1 |
C4 |
H10 |
110.690 |
| C1 |
C4 |
H12 |
109.520 |
|
C1 |
C5 |
C6 |
110.716 |
| C1 |
C5 |
H11 |
110.690 |
|
C1 |
C5 |
H13 |
109.520 |
| H2 |
C1 |
H3 |
106.362 |
|
H2 |
C1 |
C4 |
110.677 |
| H2 |
C1 |
C5 |
110.677 |
|
H3 |
C1 |
C4 |
109.032 |
| H3 |
C1 |
C5 |
109.032 |
|
C4 |
C1 |
C5 |
110.930 |
| C4 |
C7 |
N8 |
114.466 |
|
C4 |
C7 |
H15 |
109.145 |
| C4 |
C7 |
H17 |
110.774 |
|
C5 |
C6 |
N8 |
114.466 |
| C5 |
C6 |
H14 |
109.145 |
|
C5 |
C6 |
H16 |
110.774 |
| C6 |
C5 |
H11 |
110.232 |
|
C6 |
C5 |
H13 |
108.982 |
| C6 |
N8 |
C7 |
111.727 |
|
C6 |
N8 |
H9 |
108.118 |
| C7 |
C4 |
H10 |
110.232 |
|
C7 |
C4 |
H12 |
108.982 |
| C7 |
N8 |
H9 |
108.118 |
|
N8 |
C6 |
H14 |
107.280 |
| N8 |
C6 |
H16 |
108.254 |
|
N8 |
C7 |
H15 |
107.280 |
| N8 |
C7 |
H17 |
108.254 |
|
H10 |
C4 |
H12 |
106.596 |
| H11 |
C5 |
H13 |
106.596 |
|
H14 |
C6 |
H16 |
106.578 |
| H15 |
C7 |
H17 |
106.578 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.034 |
|
|
|
| 2 |
H |
-0.021 |
|
|
|
| 3 |
H |
-0.012 |
|
|
|
| 4 |
C |
0.031 |
|
|
|
| 5 |
C |
0.031 |
|
|
|
| 6 |
C |
0.078 |
|
|
|
| 7 |
C |
0.078 |
|
|
|
| 8 |
N |
-0.152 |
|
|
|
| 9 |
H |
0.057 |
|
|
|
| 10 |
H |
-0.023 |
|
|
|
| 11 |
H |
-0.023 |
|
|
|
| 12 |
H |
-0.018 |
|
|
|
| 13 |
H |
-0.018 |
|
|
|
| 14 |
H |
-0.003 |
|
|
|
| 15 |
H |
-0.003 |
|
|
|
| 16 |
H |
-0.017 |
|
|
|
| 17 |
H |
-0.017 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.300 |
0.902 |
0.000 |
0.951 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.693 |
-0.148 |
0.000 |
| y |
-0.148 |
-43.399 |
0.000 |
| z |
0.000 |
0.000 |
-38.834 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.424 |
-0.148 |
0.000 |
| y |
-0.148 |
-5.135 |
0.000 |
| z |
0.000 |
0.000 |
1.711 |
|
| Polar |
|---|
| 3z2-r2 | 3.423 |
|---|
| x2-y2 | 5.706 |
|---|
| xy | -0.148 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.881 |
0.297 |
0.000 |
| y |
0.297 |
9.285 |
0.000 |
| z |
0.000 |
0.000 |
10.332 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
