Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.830581 |
| Energy at 298.15K | -251.844946 |
| Nuclear repulsion energy | 259.053261 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3526 |
3402 |
0.24 |
|
|
|
| 2 |
A' |
3094 |
2986 |
65.31 |
|
|
|
| 3 |
A' |
3080 |
2972 |
26.68 |
|
|
|
| 4 |
A' |
3075 |
2967 |
43.67 |
|
|
|
| 5 |
A' |
3038 |
2931 |
18.12 |
|
|
|
| 6 |
A' |
3021 |
2915 |
30.61 |
|
|
|
| 7 |
A' |
2901 |
2799 |
157.70 |
|
|
|
| 8 |
A' |
1478 |
1426 |
1.99 |
|
|
|
| 9 |
A' |
1463 |
1412 |
8.37 |
|
|
|
| 10 |
A' |
1451 |
1400 |
7.96 |
|
|
|
| 11 |
A' |
1417 |
1367 |
2.56 |
|
|
|
| 12 |
A' |
1372 |
1324 |
0.29 |
|
|
|
| 13 |
A' |
1308 |
1262 |
0.80 |
|
|
|
| 14 |
A' |
1280 |
1235 |
3.62 |
|
|
|
| 15 |
A' |
1157 |
1117 |
5.16 |
|
|
|
| 16 |
A' |
1072 |
1034 |
2.28 |
|
|
|
| 17 |
A' |
1056 |
1019 |
10.59 |
|
|
|
| 18 |
A' |
924 |
892 |
0.97 |
|
|
|
| 19 |
A' |
873 |
842 |
18.48 |
|
|
|
| 20 |
A' |
839 |
809 |
0.43 |
|
|
|
| 21 |
A' |
782 |
754 |
73.05 |
|
|
|
| 22 |
A' |
541 |
522 |
17.65 |
|
|
|
| 23 |
A' |
435 |
420 |
5.97 |
|
|
|
| 24 |
A' |
397 |
383 |
1.29 |
|
|
|
| 25 |
A' |
248 |
239 |
1.18 |
|
|
|
| 26 |
A" |
3090 |
2982 |
24.32 |
|
|
|
| 27 |
A" |
3074 |
2967 |
75.38 |
|
|
|
| 28 |
A" |
3038 |
2932 |
23.19 |
|
|
|
| 29 |
A" |
2895 |
2794 |
32.31 |
|
|
|
| 30 |
A" |
1483 |
1431 |
1.37 |
|
|
|
| 31 |
A" |
1461 |
1410 |
0.32 |
|
|
|
| 32 |
A" |
1440 |
1390 |
3.44 |
|
|
|
| 33 |
A" |
1376 |
1327 |
0.12 |
|
|
|
| 34 |
A" |
1355 |
1307 |
26.74 |
|
|
|
| 35 |
A" |
1332 |
1286 |
8.62 |
|
|
|
| 36 |
A" |
1286 |
1241 |
3.73 |
|
|
|
| 37 |
A" |
1194 |
1153 |
11.51 |
|
|
|
| 38 |
A" |
1161 |
1120 |
9.46 |
|
|
|
| 39 |
A" |
1155 |
1114 |
1.59 |
|
|
|
| 40 |
A" |
1077 |
1039 |
2.48 |
|
|
|
| 41 |
A" |
974 |
940 |
0.54 |
|
|
|
| 42 |
A" |
898 |
866 |
0.31 |
|
|
|
| 43 |
A" |
816 |
787 |
0.45 |
|
|
|
| 44 |
A" |
450 |
434 |
0.78 |
|
|
|
| 45 |
A" |
246 |
237 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34812.8 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 33594.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.631 |
1.329 |
0.000 |
| H2 |
0.561 |
2.430 |
0.000 |
| H3 |
1.710 |
1.086 |
0.000 |
| C4 |
-0.014 |
0.747 |
1.259 |
| C5 |
-0.014 |
0.747 |
-1.259 |
| C6 |
-0.014 |
-0.779 |
-1.207 |
| C7 |
-0.014 |
-0.779 |
1.207 |
| N8 |
-0.697 |
-1.226 |
0.000 |
| H9 |
-0.759 |
-2.243 |
0.000 |
| H10 |
0.513 |
1.094 |
2.164 |
| H11 |
0.513 |
1.094 |
-2.164 |
| H12 |
-1.059 |
1.095 |
1.335 |
| H13 |
-1.059 |
1.095 |
-1.335 |
| H14 |
1.041 |
-1.139 |
-1.262 |
| H15 |
1.041 |
-1.139 |
1.262 |
| H16 |
-0.540 |
-1.191 |
-2.083 |
| H17 |
-0.540 |
-1.191 |
2.083 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1028 | 1.1064 | 1.5298 | 1.5298 | 2.5133 | 2.5133 | 2.8801 | 3.8336 | 2.1797 | 2.1797 | 2.1665 | 2.1665 | 2.8025 | 2.8025 | 3.4734 | 3.4734 |
H2 | 1.1028 | | 1.7685 | 2.1789 | 2.1789 | 3.4758 | 3.4758 | 3.8665 | 4.8560 | 2.5435 | 2.5435 | 2.4877 | 2.4877 | 3.8158 | 3.8158 | 4.3202 | 4.3202 | H3 | 1.1064 | 1.7685 | | 2.1618 | 2.1618 | 2.8119 | 2.8119 | 3.3379 | 4.1451 | 2.4729 | 2.4729 | 3.0745 | 3.0745 | 2.6440 | 2.6440 | 3.8198 | 3.8198 | C4 | 1.5298 | 2.1789 | 2.1618 | | 2.5182 | 2.9001 | 1.5264 | 2.4385 | 3.3291 | 1.1028 | 3.4805 | 1.1041 | 2.8182 | 3.3208 | 2.1611 | 3.8994 | 2.1709 | C5 | 1.5298 | 2.1789 | 2.1618 | 2.5182 | | 1.5264 | 2.9001 | 2.4385 | 3.3291 | 3.4805 | 1.1028 | 2.8182 | 1.1041 | 2.1611 | 3.3208 | 2.1709 | 3.8994 | C6 | 2.5133 | 3.4758 | 2.8119 | 2.9001 | 1.5264 | | 2.4146 | 1.4576 | 2.0392 | 3.8918 | 2.1673 | 3.3263 | 2.1489 | 1.1165 | 2.7097 | 1.1022 | 3.3580 | C7 | 2.5133 | 3.4758 | 2.8119 | 1.5264 | 2.9001 | 2.4146 | | 1.4576 | 2.0392 | 2.1673 | 3.8918 | 2.1489 | 3.3263 | 2.7097 | 1.1165 | 3.3580 | 1.1022 | N8 | 2.8801 | 3.8665 | 3.3379 | 2.4385 | 2.4385 | 1.4576 | 1.4576 | | 1.0189 | 3.3953 | 3.3953 | 2.7020 | 2.7020 | 2.1501 | 2.1501 | 2.0896 | 2.0896 | H9 | 3.8336 | 4.8560 | 4.1451 | 3.3291 | 3.3291 | 2.0392 | 2.0392 | 1.0189 | | 4.1756 | 4.1756 | 3.6076 | 3.6076 | 2.4606 | 2.4606 | 2.3443 | 2.3443 | H10 | 2.1797 | 2.5435 | 2.4729 | 1.1028 | 3.4805 | 3.8918 | 2.1673 | 3.3953 | 4.1756 | | 4.3274 | 1.7773 | 3.8357 | 4.1232 | 2.4649 | 4.9363 | 2.5171 | H11 | 2.1797 | 2.5435 | 2.4729 | 3.4805 | 1.1028 | 2.1673 | 3.8918 | 3.3953 | 4.1756 | 4.3274 | | 3.8357 | 1.7773 | 2.4649 | 4.1232 | 2.5171 | 4.9363 | H12 | 2.1665 | 2.4877 | 3.0745 | 1.1041 | 2.8182 | 3.3263 | 2.1489 | 2.7020 | 3.6076 | 1.7773 | 3.8357 | | 2.6699 | 4.0183 | 3.0670 | 4.1447 | 2.4605 | H13 | 2.1665 | 2.4877 | 3.0745 | 2.8182 | 1.1041 | 2.1489 | 3.3263 | 2.7020 | 3.6076 | 3.8357 | 1.7773 | 2.6699 | | 3.0670 | 4.0183 | 2.4605 | 4.1447 | H14 | 2.8025 | 3.8158 | 2.6440 | 3.3208 | 2.1611 | 1.1165 | 2.7097 | 2.1501 | 2.4606 | 4.1232 | 2.4649 | 4.0183 | 3.0670 | | 2.5246 | 1.7829 | 3.7011 | H15 | 2.8025 | 3.8158 | 2.6440 | 2.1611 | 3.3208 | 2.7097 | 1.1165 | 2.1501 | 2.4606 | 2.4649 | 4.1232 | 3.0670 | 4.0183 | 2.5246 | | 3.7011 | 1.7829 | H16 | 3.4734 | 4.3202 | 3.8198 | 3.8994 | 2.1709 | 1.1022 | 3.3580 | 2.0896 | 2.3443 | 4.9363 | 2.5171 | 4.1447 | 2.4605 | 1.7829 | 3.7011 | | 4.1668 | H17 | 3.4734 | 4.3202 | 3.8198 | 2.1709 | 3.8994 | 3.3580 | 1.1022 | 2.0896 | 2.3443 | 2.5171 | 4.9363 | 2.4605 | 4.1447 | 3.7011 | 1.7829 | 4.1668 | |
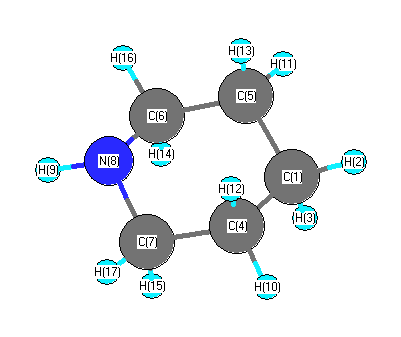 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.639 |
|
C1 |
C4 |
H10 |
110.735 |
| C1 |
C4 |
H12 |
109.619 |
|
C1 |
C5 |
C6 |
110.639 |
| C1 |
C5 |
H11 |
110.735 |
|
C1 |
C5 |
H13 |
109.619 |
| H2 |
C1 |
H3 |
106.360 |
|
H2 |
C1 |
C4 |
110.669 |
| H2 |
C1 |
C5 |
110.669 |
|
H3 |
C1 |
C4 |
109.120 |
| H3 |
C1 |
C5 |
109.120 |
|
C4 |
C1 |
C5 |
110.781 |
| C4 |
C7 |
N8 |
109.583 |
|
C4 |
C7 |
H15 |
108.719 |
| C4 |
C7 |
H17 |
110.308 |
|
C5 |
C6 |
N8 |
109.583 |
| C5 |
C6 |
H14 |
108.719 |
|
C5 |
C6 |
H16 |
110.308 |
| C6 |
C5 |
H11 |
109.995 |
|
C6 |
C5 |
H13 |
108.483 |
| C6 |
N8 |
C7 |
111.845 |
|
C6 |
N8 |
H9 |
109.587 |
| C7 |
C4 |
H10 |
109.995 |
|
C7 |
C4 |
H12 |
108.483 |
| C7 |
N8 |
H9 |
109.587 |
|
N8 |
C6 |
H14 |
112.613 |
| N8 |
C6 |
H16 |
108.638 |
|
N8 |
C7 |
H15 |
112.613 |
| N8 |
C7 |
H17 |
108.638 |
|
H10 |
C4 |
H12 |
107.282 |
| H11 |
C5 |
H13 |
107.282 |
|
H14 |
C6 |
H16 |
106.947 |
| H15 |
C7 |
H17 |
106.947 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.067 |
|
|
|
| 2 |
H |
0.027 |
|
|
|
| 3 |
H |
0.027 |
|
|
|
| 4 |
C |
-0.041 |
|
|
|
| 5 |
C |
-0.041 |
|
|
|
| 6 |
C |
0.023 |
|
|
|
| 7 |
C |
0.023 |
|
|
|
| 8 |
N |
-0.226 |
|
|
|
| 9 |
H |
0.087 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
H |
0.036 |
|
|
|
| 13 |
H |
0.036 |
|
|
|
| 14 |
H |
0.009 |
|
|
|
| 15 |
H |
0.009 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
| 17 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.563 |
-0.511 |
0.000 |
0.760 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.827 |
0.066 |
0.000 |
| y |
0.066 |
-36.348 |
0.000 |
| z |
0.000 |
0.000 |
-37.960 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.673 |
0.066 |
0.000 |
| y |
0.066 |
3.045 |
0.000 |
| z |
0.000 |
0.000 |
0.627 |
|
| Polar |
|---|
| 3z2-r2 | 1.255 |
|---|
| x2-y2 | -4.479 |
|---|
| xy | 0.066 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.300 |
0.269 |
0.000 |
| y |
0.269 |
9.339 |
0.000 |
| z |
0.000 |
0.000 |
9.791 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.829786 |
| Energy at 298.15K | -251.844094 |
| Nuclear repulsion energy | 258.520881 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3485 |
3363 |
0.82 |
|
|
|
| 2 |
A' |
3084 |
2976 |
73.17 |
|
|
|
| 3 |
A' |
3078 |
2971 |
34.90 |
|
|
|
| 4 |
A' |
3072 |
2965 |
43.18 |
|
|
|
| 5 |
A' |
3021 |
2916 |
38.00 |
|
|
|
| 6 |
A' |
3017 |
2912 |
71.76 |
|
|
|
| 7 |
A' |
3016 |
2910 |
0.58 |
|
|
|
| 8 |
A' |
1471 |
1420 |
2.15 |
|
|
|
| 9 |
A' |
1454 |
1403 |
9.75 |
|
|
|
| 10 |
A' |
1451 |
1400 |
4.63 |
|
|
|
| 11 |
A' |
1385 |
1337 |
5.74 |
|
|
|
| 12 |
A' |
1372 |
1324 |
1.13 |
|
|
|
| 13 |
A' |
1332 |
1285 |
0.04 |
|
|
|
| 14 |
A' |
1274 |
1229 |
8.65 |
|
|
|
| 15 |
A' |
1181 |
1140 |
2.95 |
|
|
|
| 16 |
A' |
1056 |
1019 |
1.09 |
|
|
|
| 17 |
A' |
1020 |
984 |
9.53 |
|
|
|
| 18 |
A' |
928 |
896 |
5.28 |
|
|
|
| 19 |
A' |
868 |
837 |
44.42 |
|
|
|
| 20 |
A' |
829 |
800 |
0.23 |
|
|
|
| 21 |
A' |
776 |
749 |
86.19 |
|
|
|
| 22 |
A' |
546 |
527 |
1.53 |
|
|
|
| 23 |
A' |
436 |
421 |
1.03 |
|
|
|
| 24 |
A' |
387 |
373 |
5.02 |
|
|
|
| 25 |
A' |
240 |
232 |
4.18 |
|
|
|
| 26 |
A" |
3081 |
2973 |
31.65 |
|
|
|
| 27 |
A" |
3071 |
2964 |
74.82 |
|
|
|
| 28 |
A" |
3017 |
2911 |
2.10 |
|
|
|
| 29 |
A" |
3012 |
2907 |
45.01 |
|
|
|
| 30 |
A" |
1486 |
1434 |
8.15 |
|
|
|
| 31 |
A" |
1449 |
1398 |
5.15 |
|
|
|
| 32 |
A" |
1441 |
1391 |
1.31 |
|
|
|
| 33 |
A" |
1373 |
1325 |
1.42 |
|
|
|
| 34 |
A" |
1360 |
1313 |
4.00 |
|
|
|
| 35 |
A" |
1330 |
1283 |
1.44 |
|
|
|
| 36 |
A" |
1290 |
1245 |
0.90 |
|
|
|
| 37 |
A" |
1220 |
1178 |
16.25 |
|
|
|
| 38 |
A" |
1148 |
1107 |
15.21 |
|
|
|
| 39 |
A" |
1114 |
1075 |
0.83 |
|
|
|
| 40 |
A" |
1075 |
1037 |
2.72 |
|
|
|
| 41 |
A" |
942 |
909 |
3.44 |
|
|
|
| 42 |
A" |
888 |
857 |
0.20 |
|
|
|
| 43 |
A" |
802 |
774 |
0.10 |
|
|
|
| 44 |
A" |
456 |
440 |
0.70 |
|
|
|
| 45 |
A" |
234 |
226 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34783.6 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 33566.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.636 |
1.325 |
0.000 |
| H2 |
0.570 |
2.426 |
0.000 |
| H3 |
1.715 |
1.080 |
0.000 |
| C4 |
-0.008 |
0.743 |
1.260 |
| C5 |
-0.008 |
0.743 |
-1.260 |
| C6 |
-0.008 |
-0.789 |
-1.209 |
| C7 |
-0.008 |
-0.789 |
1.209 |
| N8 |
-0.605 |
-1.349 |
0.000 |
| H9 |
-1.604 |
-1.133 |
0.000 |
| H10 |
0.514 |
1.096 |
2.166 |
| H11 |
0.514 |
1.096 |
-2.166 |
| H12 |
-1.052 |
1.101 |
1.340 |
| H13 |
-1.052 |
1.101 |
-1.340 |
| H14 |
1.035 |
-1.154 |
-1.265 |
| H15 |
1.035 |
-1.154 |
1.265 |
| H16 |
-0.530 |
-1.210 |
-2.084 |
| H17 |
-0.530 |
-1.210 |
2.084 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1027 | 1.1064 | 1.5305 | 1.5305 | 2.5192 | 2.5192 | 2.9484 | 3.3258 | 2.1814 | 2.1814 | 2.1674 | 2.1674 | 2.8119 | 2.8119 | 3.4829 | 3.4829 |
H2 | 1.1027 | | 1.7676 | 2.1804 | 2.1804 | 3.4828 | 3.4828 | 3.9536 | 4.1701 | 2.5421 | 2.5421 | 2.4861 | 2.4861 | 3.8254 | 3.8254 | 4.3326 | 4.3326 | H3 | 1.1064 | 1.7676 | | 2.1615 | 2.1615 | 2.8150 | 2.8150 | 3.3589 | 3.9891 | 2.4766 | 2.4766 | 3.0749 | 3.0749 | 2.6560 | 2.6560 | 3.8246 | 3.8246 | C4 | 1.5305 | 2.1804 | 2.1615 | | 2.5203 | 2.9059 | 1.5325 | 2.5142 | 2.7664 | 1.1038 | 3.4837 | 1.1066 | 2.8249 | 3.3264 | 2.1650 | 3.9078 | 2.1828 | C5 | 1.5305 | 2.1804 | 2.1615 | 2.5203 | | 1.5325 | 2.9059 | 2.5142 | 2.7664 | 3.4837 | 1.1038 | 2.8249 | 1.1066 | 2.1650 | 3.3264 | 2.1828 | 3.9078 | C6 | 2.5192 | 3.4828 | 2.8150 | 2.9059 | 1.5325 | | 2.4186 | 1.4603 | 2.0315 | 3.9011 | 2.1775 | 3.3410 | 2.1632 | 1.1066 | 2.7102 | 1.1024 | 3.3612 | C7 | 2.5192 | 3.4828 | 2.8150 | 1.5325 | 2.9059 | 2.4186 | | 1.4603 | 2.0315 | 2.1775 | 3.9011 | 2.1632 | 3.3410 | 2.7102 | 1.1066 | 3.3612 | 1.1024 | N8 | 2.9484 | 3.9536 | 3.3589 | 2.5142 | 2.5142 | 1.4603 | 1.4603 | | 1.0223 | 3.4532 | 3.4532 | 2.8288 | 2.8288 | 2.0801 | 2.0801 | 2.0904 | 2.0904 | H9 | 3.3258 | 4.1701 | 3.9891 | 2.7664 | 2.7664 | 2.0315 | 2.0315 | 1.0223 | | 3.7614 | 3.7614 | 2.6632 | 2.6632 | 2.9264 | 2.9264 | 2.3460 | 2.3460 | H10 | 2.1814 | 2.5421 | 2.4766 | 1.1038 | 3.4837 | 3.9011 | 2.1775 | 3.4532 | 3.7614 | | 4.3319 | 1.7708 | 3.8402 | 4.1363 | 2.4792 | 4.9471 | 2.5328 | H11 | 2.1814 | 2.5421 | 2.4766 | 3.4837 | 1.1038 | 2.1775 | 3.9011 | 3.4532 | 3.7614 | 4.3319 | | 3.8402 | 1.7708 | 2.4792 | 4.1363 | 2.5328 | 4.9471 | H12 | 2.1674 | 2.4861 | 3.0749 | 1.1066 | 2.8249 | 3.3410 | 2.1632 | 2.8288 | 2.6632 | 1.7708 | 3.8402 | | 2.6805 | 4.0288 | 3.0738 | 4.1643 | 2.4834 | H13 | 2.1674 | 2.4861 | 3.0749 | 2.8249 | 1.1066 | 2.1632 | 3.3410 | 2.8288 | 2.6632 | 3.8402 | 1.7708 | 2.6805 | | 3.0738 | 4.0288 | 2.4834 | 4.1643 | H14 | 2.8119 | 3.8254 | 2.6560 | 3.3264 | 2.1650 | 1.1066 | 2.7102 | 2.0801 | 2.9264 | 4.1363 | 2.4792 | 4.0288 | 3.0738 | | 2.5306 | 1.7671 | 3.6976 | H15 | 2.8119 | 3.8254 | 2.6560 | 2.1650 | 3.3264 | 2.7102 | 1.1066 | 2.0801 | 2.9264 | 2.4792 | 4.1363 | 3.0738 | 4.0288 | 2.5306 | | 3.6976 | 1.7671 | H16 | 3.4829 | 4.3326 | 3.8246 | 3.9078 | 2.1828 | 1.1024 | 3.3612 | 2.0904 | 2.3460 | 4.9471 | 2.5328 | 4.1643 | 2.4834 | 1.7671 | 3.6976 | | 4.1687 | H17 | 3.4829 | 4.3326 | 3.8246 | 2.1828 | 3.9078 | 3.3612 | 1.1024 | 2.0904 | 2.3460 | 2.5328 | 4.9471 | 2.4834 | 4.1643 | 3.6976 | 1.7671 | 4.1687 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.661 |
|
C1 |
C4 |
H10 |
110.760 |
| C1 |
C4 |
H12 |
109.496 |
|
C1 |
C5 |
C6 |
110.661 |
| C1 |
C5 |
H11 |
110.760 |
|
C1 |
C5 |
H13 |
109.496 |
| H2 |
C1 |
H3 |
106.281 |
|
H2 |
C1 |
C4 |
110.745 |
| H2 |
C1 |
C5 |
110.745 |
|
H3 |
C1 |
C4 |
109.048 |
| H3 |
C1 |
C5 |
109.048 |
|
C4 |
C1 |
C5 |
110.843 |
| C4 |
C7 |
N8 |
114.277 |
|
C4 |
C7 |
H15 |
109.172 |
| C4 |
C7 |
H17 |
110.813 |
|
C5 |
C6 |
N8 |
114.277 |
| C5 |
C6 |
H14 |
109.172 |
|
C5 |
C6 |
H16 |
110.813 |
| C6 |
C5 |
H11 |
110.312 |
|
C6 |
C5 |
H13 |
109.033 |
| C6 |
N8 |
C7 |
111.814 |
|
C6 |
N8 |
H9 |
108.541 |
| C7 |
C4 |
H10 |
110.312 |
|
C7 |
C4 |
H12 |
109.033 |
| C7 |
N8 |
H9 |
108.541 |
|
N8 |
C6 |
H14 |
107.465 |
| N8 |
C6 |
H16 |
108.506 |
|
N8 |
C7 |
H15 |
107.465 |
| N8 |
C7 |
H17 |
108.506 |
|
H10 |
C4 |
H12 |
106.473 |
| H11 |
C5 |
H13 |
106.473 |
|
H14 |
C6 |
H16 |
106.252 |
| H15 |
C7 |
H17 |
106.252 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.056 |
|
|
|
| 2 |
H |
0.026 |
|
|
|
| 3 |
H |
0.030 |
|
|
|
| 4 |
C |
-0.062 |
|
|
|
| 5 |
C |
-0.062 |
|
|
|
| 6 |
C |
0.022 |
|
|
|
| 7 |
C |
0.022 |
|
|
|
| 8 |
N |
-0.227 |
|
|
|
| 9 |
H |
0.085 |
|
|
|
| 10 |
H |
0.024 |
|
|
|
| 11 |
H |
0.024 |
|
|
|
| 12 |
H |
0.024 |
|
|
|
| 13 |
H |
0.024 |
|
|
|
| 14 |
H |
0.036 |
|
|
|
| 15 |
H |
0.036 |
|
|
|
| 16 |
H |
0.028 |
|
|
|
| 17 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.294 |
0.989 |
0.000 |
1.032 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.860 |
-0.201 |
0.000 |
| y |
-0.201 |
-42.892 |
0.000 |
| z |
0.000 |
0.000 |
-37.949 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.561 |
-0.201 |
0.000 |
| y |
-0.201 |
-5.488 |
0.000 |
| z |
0.000 |
0.000 |
1.927 |
|
| Polar |
|---|
| 3z2-r2 | 3.853 |
|---|
| x2-y2 | 6.032 |
|---|
| xy | -0.201 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.517 |
0.247 |
0.000 |
| y |
0.247 |
8.865 |
0.000 |
| z |
0.000 |
0.000 |
9.805 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
