Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -311.868215 |
| Energy at 298.15K | -311.883528 |
| HF Energy | -311.868215 |
| Nuclear repulsion energy | 312.789842 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3056 |
3038 |
34.83 |
|
|
|
| 2 |
A1 |
2974 |
2957 |
51.84 |
|
|
|
| 3 |
A1 |
2967 |
2950 |
15.31 |
|
|
|
| 4 |
A1 |
2864 |
2847 |
100.54 |
|
|
|
| 5 |
A1 |
1459 |
1451 |
1.26 |
|
|
|
| 6 |
A1 |
1433 |
1425 |
5.59 |
|
|
|
| 7 |
A1 |
1418 |
1410 |
0.07 |
|
|
|
| 8 |
A1 |
1390 |
1382 |
0.88 |
|
|
|
| 9 |
A1 |
1347 |
1339 |
0.72 |
|
|
|
| 10 |
A1 |
1276 |
1269 |
2.23 |
|
|
|
| 11 |
A1 |
1138 |
1132 |
1.69 |
|
|
|
| 12 |
A1 |
1050 |
1044 |
5.72 |
|
|
|
| 13 |
A1 |
987 |
981 |
18.20 |
|
|
|
| 14 |
A1 |
897 |
892 |
9.11 |
|
|
|
| 15 |
A1 |
421 |
418 |
0.33 |
|
|
|
| 16 |
A1 |
312 |
310 |
0.17 |
|
|
|
| 17 |
A1 |
100 |
99 |
0.12 |
|
|
|
| 18 |
A2 |
3043 |
3026 |
0.00 |
|
|
|
| 19 |
A2 |
3013 |
2996 |
0.00 |
|
|
|
| 20 |
A2 |
2887 |
2870 |
0.00 |
|
|
|
| 21 |
A2 |
1421 |
1413 |
0.00 |
|
|
|
| 22 |
A2 |
1258 |
1251 |
0.00 |
|
|
|
| 23 |
A2 |
1207 |
1200 |
0.00 |
|
|
|
| 24 |
A2 |
1124 |
1117 |
0.00 |
|
|
|
| 25 |
A2 |
862 |
857 |
0.00 |
|
|
|
| 26 |
A2 |
748 |
743 |
0.00 |
|
|
|
| 27 |
A2 |
233 |
232 |
0.00 |
|
|
|
| 28 |
A2 |
135 |
134 |
0.00 |
|
|
|
| 29 |
A2 |
75 |
75 |
0.00 |
|
|
|
| 30 |
B1 |
3044 |
3026 |
95.18 |
|
|
|
| 31 |
B1 |
3014 |
2996 |
14.44 |
|
|
|
| 32 |
B1 |
2881 |
2864 |
131.46 |
|
|
|
| 33 |
B1 |
1421 |
1413 |
13.32 |
|
|
|
| 34 |
B1 |
1258 |
1251 |
0.63 |
|
|
|
| 35 |
B1 |
1212 |
1205 |
3.47 |
|
|
|
| 36 |
B1 |
1148 |
1141 |
3.78 |
|
|
|
| 37 |
B1 |
873 |
868 |
3.69 |
|
|
|
| 38 |
B1 |
749 |
744 |
5.12 |
|
|
|
| 39 |
B1 |
234 |
233 |
0.14 |
|
|
|
| 40 |
B1 |
153 |
152 |
3.93 |
|
|
|
| 41 |
B1 |
55 |
54 |
0.20 |
|
|
|
| 42 |
B2 |
3056 |
3038 |
18.30 |
|
|
|
| 43 |
B2 |
2974 |
2957 |
2.61 |
|
|
|
| 44 |
B2 |
2966 |
2949 |
39.87 |
|
|
|
| 45 |
B2 |
2851 |
2834 |
18.35 |
|
|
|
| 46 |
B2 |
1440 |
1432 |
9.61 |
|
|
|
| 47 |
B2 |
1429 |
1421 |
0.01 |
|
|
|
| 48 |
B2 |
1417 |
1408 |
2.23 |
|
|
|
| 49 |
B2 |
1347 |
1339 |
0.76 |
|
|
|
| 50 |
B2 |
1341 |
1333 |
32.54 |
|
|
|
| 51 |
B2 |
1261 |
1254 |
19.72 |
|
|
|
| 52 |
B2 |
1130 |
1123 |
250.31 |
|
|
|
| 53 |
B2 |
1091 |
1085 |
1.31 |
|
|
|
| 54 |
B2 |
1043 |
1037 |
0.22 |
|
|
|
| 55 |
B2 |
878 |
873 |
1.26 |
|
|
|
| 56 |
B2 |
501 |
498 |
4.89 |
|
|
|
| 57 |
B2 |
241 |
240 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41050.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 40812.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.394 |
| C2 |
0.000 |
1.181 |
-0.394 |
| C3 |
0.000 |
-1.181 |
-0.394 |
| C4 |
0.000 |
2.391 |
0.533 |
| C5 |
0.000 |
-2.391 |
0.533 |
| C6 |
0.000 |
3.717 |
-0.230 |
| C7 |
0.000 |
-3.717 |
-0.230 |
| H8 |
0.898 |
1.205 |
-1.065 |
| H9 |
-0.898 |
1.205 |
-1.065 |
| H10 |
-0.898 |
-1.205 |
-1.065 |
| H11 |
0.898 |
-1.205 |
-1.065 |
| H12 |
0.889 |
2.323 |
1.196 |
| H13 |
-0.889 |
2.323 |
1.196 |
| H14 |
-0.889 |
-2.323 |
1.196 |
| H15 |
0.889 |
-2.323 |
1.196 |
| H16 |
0.000 |
4.580 |
0.464 |
| H17 |
-0.895 |
3.813 |
-0.880 |
| H18 |
0.895 |
3.813 |
-0.880 |
| H19 |
0.000 |
-4.580 |
0.464 |
| H20 |
0.895 |
-3.813 |
-0.880 |
| H21 |
-0.895 |
-3.813 |
-0.880 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4199 | 1.4199 | 2.3947 | 2.3947 | 3.7686 | 3.7686 | 2.0944 | 2.0944 | 2.0944 | 2.0944 | 2.6139 | 2.6139 | 2.6139 | 2.6139 | 4.5810 | 4.1186 | 4.1186 | 4.5810 | 4.1186 | 4.1186 |
C2 | 1.4199 | | 2.3627 | 1.5238 | 3.6903 | 2.5407 | 4.9008 | 1.1212 | 1.1212 | 2.6365 | 2.6365 | 2.1502 | 2.1502 | 3.9500 | 3.9500 | 3.5058 | 2.8218 | 2.8218 | 5.8254 | 5.0971 | 5.0971 | C3 | 1.4199 | 2.3627 | | 3.6903 | 1.5238 | 4.9008 | 2.5407 | 2.6365 | 2.6365 | 1.1212 | 1.1212 | 3.9500 | 3.9500 | 2.1502 | 2.1502 | 5.8254 | 5.0971 | 5.0971 | 3.5058 | 2.8218 | 2.8218 | C4 | 2.3947 | 1.5238 | 3.6903 | | 4.7812 | 1.5299 | 6.1548 | 2.1832 | 2.1832 | 4.0360 | 4.0360 | 1.1111 | 1.1111 | 4.8427 | 4.8427 | 2.1909 | 2.1958 | 2.1958 | 6.9714 | 6.4252 | 6.4252 | C5 | 2.3947 | 3.6903 | 1.5238 | 4.7812 | | 6.1548 | 1.5299 | 4.0360 | 4.0360 | 2.1832 | 2.1832 | 4.8427 | 4.8427 | 1.1111 | 1.1111 | 6.9714 | 6.4252 | 6.4252 | 2.1909 | 2.1958 | 2.1958 | C6 | 3.7686 | 2.5407 | 4.9008 | 1.5299 | 6.1548 | | 7.4335 | 2.7951 | 2.7951 | 5.0723 | 5.0723 | 2.1829 | 2.1829 | 6.2695 | 6.2695 | 1.1080 | 1.1104 | 1.1104 | 8.3262 | 7.6106 | 7.6106 | C7 | 3.7686 | 4.9008 | 2.5407 | 6.1548 | 1.5299 | 7.4335 | | 5.0723 | 5.0723 | 2.7951 | 2.7951 | 6.2695 | 6.2695 | 2.1829 | 2.1829 | 8.3262 | 7.6106 | 7.6106 | 1.1080 | 1.1104 | 1.1104 | H8 | 2.0944 | 1.1212 | 2.6365 | 2.1832 | 4.0360 | 2.7951 | 5.0723 | | 1.7955 | 3.0054 | 2.4100 | 2.5228 | 3.0915 | 4.5559 | 4.1909 | 3.8130 | 3.1700 | 2.6146 | 6.0512 | 5.0215 | 5.3318 | H9 | 2.0944 | 1.1212 | 2.6365 | 2.1832 | 4.0360 | 2.7951 | 5.0723 | 1.7955 | | 2.4100 | 3.0054 | 3.0915 | 2.5228 | 4.1909 | 4.5559 | 3.8130 | 2.6146 | 3.1700 | 6.0512 | 5.3318 | 5.0215 | H10 | 2.0944 | 2.6365 | 1.1212 | 4.0360 | 2.1832 | 5.0723 | 2.7951 | 3.0054 | 2.4100 | | 1.7955 | 4.5559 | 4.1909 | 2.5228 | 3.0915 | 6.0512 | 5.0215 | 5.3318 | 3.8130 | 3.1700 | 2.6146 | H11 | 2.0944 | 2.6365 | 1.1212 | 4.0360 | 2.1832 | 5.0723 | 2.7951 | 2.4100 | 3.0054 | 1.7955 | | 4.1909 | 4.5559 | 3.0915 | 2.5228 | 6.0512 | 5.3318 | 5.0215 | 3.8130 | 2.6146 | 3.1700 | H12 | 2.6139 | 2.1502 | 3.9500 | 1.1111 | 4.8427 | 2.1829 | 6.2695 | 2.5228 | 3.0915 | 4.5559 | 4.1909 | | 1.7782 | 4.9754 | 4.6468 | 2.5338 | 3.1165 | 2.5555 | 6.9992 | 6.4782 | 6.7193 | H13 | 2.6139 | 2.1502 | 3.9500 | 1.1111 | 4.8427 | 2.1829 | 6.2695 | 3.0915 | 2.5228 | 4.1909 | 4.5559 | 1.7782 | | 4.6468 | 4.9754 | 2.5338 | 2.5555 | 3.1165 | 6.9992 | 6.7193 | 6.4782 | H14 | 2.6139 | 3.9500 | 2.1502 | 4.8427 | 1.1111 | 6.2695 | 2.1829 | 4.5559 | 4.1909 | 2.5228 | 3.0915 | 4.9754 | 4.6468 | | 1.7782 | 6.9992 | 6.4782 | 6.7193 | 2.5338 | 3.1165 | 2.5555 | H15 | 2.6139 | 3.9500 | 2.1502 | 4.8427 | 1.1111 | 6.2695 | 2.1829 | 4.1909 | 4.5559 | 3.0915 | 2.5228 | 4.6468 | 4.9754 | 1.7782 | | 6.9992 | 6.7193 | 6.4782 | 2.5338 | 2.5555 | 3.1165 | H16 | 4.5810 | 3.5058 | 5.8254 | 2.1909 | 6.9714 | 1.1080 | 8.3262 | 3.8130 | 3.8130 | 6.0512 | 6.0512 | 2.5338 | 2.5338 | 6.9992 | 6.9992 | | 1.7881 | 1.7881 | 9.1609 | 8.5475 | 8.5475 | H17 | 4.1186 | 2.8218 | 5.0971 | 2.1958 | 6.4252 | 1.1104 | 7.6106 | 3.1700 | 2.6146 | 5.0215 | 5.3318 | 3.1165 | 2.5555 | 6.4782 | 6.7193 | 1.7881 | | 1.7893 | 8.5475 | 7.8332 | 7.6261 | H18 | 4.1186 | 2.8218 | 5.0971 | 2.1958 | 6.4252 | 1.1104 | 7.6106 | 2.6146 | 3.1700 | 5.3318 | 5.0215 | 2.5555 | 3.1165 | 6.7193 | 6.4782 | 1.7881 | 1.7893 | | 8.5475 | 7.6261 | 7.8332 | H19 | 4.5810 | 5.8254 | 3.5058 | 6.9714 | 2.1909 | 8.3262 | 1.1080 | 6.0512 | 6.0512 | 3.8130 | 3.8130 | 6.9992 | 6.9992 | 2.5338 | 2.5338 | 9.1609 | 8.5475 | 8.5475 | | 1.7881 | 1.7881 | H20 | 4.1186 | 5.0971 | 2.8218 | 6.4252 | 2.1958 | 7.6106 | 1.1104 | 5.0215 | 5.3318 | 3.1700 | 2.6146 | 6.4782 | 6.7193 | 3.1165 | 2.5555 | 8.5475 | 7.8332 | 7.6261 | 1.7881 | | 1.7893 | H21 | 4.1186 | 5.0971 | 2.8218 | 6.4252 | 2.1958 | 7.6106 | 1.1104 | 5.3318 | 5.0215 | 2.6146 | 3.1700 | 6.7193 | 6.4782 | 2.5555 | 3.1165 | 8.5475 | 7.6261 | 7.8332 | 1.7881 | 1.7893 | |
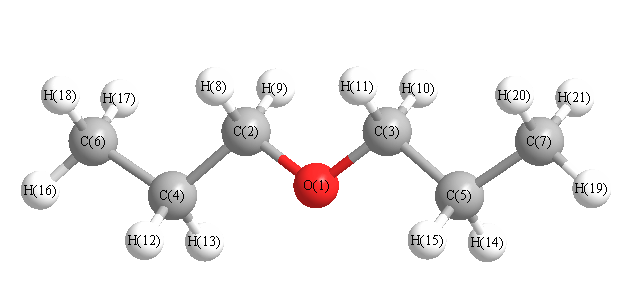 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.827 |
|
O1 |
C2 |
H8 |
110.467 |
| O1 |
C2 |
H9 |
110.467 |
|
O1 |
C3 |
C5 |
108.827 |
| O1 |
C3 |
H10 |
110.467 |
|
O1 |
C3 |
H11 |
110.467 |
| C2 |
O1 |
C3 |
112.613 |
|
C2 |
C4 |
C6 |
112.612 |
| C2 |
C4 |
H12 |
108.366 |
|
C2 |
C4 |
H13 |
108.366 |
| C3 |
C5 |
C7 |
112.612 |
|
C3 |
C5 |
H14 |
108.366 |
| C3 |
C5 |
H15 |
108.366 |
|
C4 |
C2 |
H8 |
110.335 |
| C4 |
C2 |
H9 |
110.335 |
|
C4 |
C6 |
H16 |
111.306 |
| C4 |
C6 |
H17 |
111.551 |
|
C4 |
C6 |
H18 |
111.551 |
| C5 |
C3 |
H10 |
110.335 |
|
C5 |
C3 |
H11 |
110.335 |
| C5 |
C7 |
H19 |
111.306 |
|
C5 |
C7 |
H20 |
111.551 |
| C5 |
C7 |
H21 |
111.551 |
|
C6 |
C4 |
H12 |
110.484 |
| C6 |
C4 |
H13 |
110.484 |
|
C7 |
C5 |
H14 |
110.484 |
| C7 |
C5 |
H15 |
110.484 |
|
H8 |
C2 |
H9 |
106.400 |
| H10 |
C3 |
H11 |
106.400 |
|
H12 |
C4 |
H13 |
106.297 |
| H14 |
C5 |
H15 |
106.297 |
|
H16 |
C6 |
H17 |
107.423 |
| H16 |
C6 |
H18 |
107.423 |
|
H17 |
C6 |
H18 |
107.356 |
| H19 |
C7 |
H20 |
107.423 |
|
H19 |
C7 |
H21 |
107.423 |
| H20 |
C7 |
H21 |
107.356 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.285 |
|
|
|
| 2 |
C |
0.116 |
|
|
|
| 3 |
C |
0.116 |
|
|
|
| 4 |
C |
-0.072 |
|
|
|
| 5 |
C |
-0.072 |
|
|
|
| 6 |
C |
-0.070 |
|
|
|
| 7 |
C |
-0.070 |
|
|
|
| 8 |
H |
0.006 |
|
|
|
| 9 |
H |
0.006 |
|
|
|
| 10 |
H |
0.006 |
|
|
|
| 11 |
H |
0.006 |
|
|
|
| 12 |
H |
0.029 |
|
|
|
| 13 |
H |
0.029 |
|
|
|
| 14 |
H |
0.029 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
| 16 |
H |
0.032 |
|
|
|
| 17 |
H |
0.033 |
|
|
|
| 18 |
H |
0.033 |
|
|
|
| 19 |
H |
0.032 |
|
|
|
| 20 |
H |
0.033 |
|
|
|
| 21 |
H |
0.033 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.739 |
0.739 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.348 |
0.000 |
0.000 |
| y |
0.000 |
-43.897 |
0.000 |
| z |
0.000 |
0.000 |
-47.317 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.741 |
0.000 |
0.000 |
| y |
0.000 |
2.936 |
0.000 |
| z |
0.000 |
0.000 |
-2.195 |
|
| Polar |
|---|
| 3z2-r2 | -4.389 |
|---|
| x2-y2 | -2.451 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.823 |
0.000 |
0.000 |
| y |
0.000 |
14.507 |
0.000 |
| z |
0.000 |
0.000 |
10.297 |
<r2> (average value of r
2) Å
2
| <r2> |
437.782 |
| (<r2>)1/2 |
20.923 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -311.869673 |
| Energy at 298.15K | -311.885201 |
| HF Energy | -311.869673 |
| Nuclear repulsion energy | 323.254319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3069 |
3051 |
1.90 |
|
|
|
| 2 |
A |
3049 |
3031 |
50.66 |
|
|
|
| 3 |
A |
3011 |
2993 |
14.15 |
|
|
|
| 4 |
A |
2971 |
2954 |
3.81 |
|
|
|
| 5 |
A |
2970 |
2953 |
4.85 |
|
|
|
| 6 |
A |
2900 |
2883 |
11.00 |
|
|
|
| 7 |
A |
2869 |
2852 |
100.20 |
|
|
|
| 8 |
A |
1452 |
1443 |
2.94 |
|
|
|
| 9 |
A |
1434 |
1426 |
3.53 |
|
|
|
| 10 |
A |
1415 |
1407 |
6.67 |
|
|
|
| 11 |
A |
1399 |
1391 |
1.95 |
|
|
|
| 12 |
A |
1382 |
1374 |
2.51 |
|
|
|
| 13 |
A |
1345 |
1337 |
2.74 |
|
|
|
| 14 |
A |
1313 |
1305 |
0.82 |
|
|
|
| 15 |
A |
1263 |
1255 |
2.57 |
|
|
|
| 16 |
A |
1209 |
1202 |
0.36 |
|
|
|
| 17 |
A |
1131 |
1125 |
5.59 |
|
|
|
| 18 |
A |
1099 |
1093 |
7.11 |
|
|
|
| 19 |
A |
1065 |
1059 |
9.12 |
|
|
|
| 20 |
A |
925 |
919 |
6.54 |
|
|
|
| 21 |
A |
893 |
888 |
6.72 |
|
|
|
| 22 |
A |
884 |
879 |
1.34 |
|
|
|
| 23 |
A |
730 |
725 |
0.34 |
|
|
|
| 24 |
A |
469 |
466 |
0.18 |
|
|
|
| 25 |
A |
327 |
325 |
0.32 |
|
|
|
| 26 |
A |
264 |
263 |
0.67 |
|
|
|
| 27 |
A |
166 |
165 |
0.02 |
|
|
|
| 28 |
A |
111 |
111 |
0.00 |
|
|
|
| 29 |
A |
44 |
44 |
0.00 |
|
|
|
| 30 |
B |
3069 |
3051 |
42.16 |
|
|
|
| 31 |
B |
3049 |
3031 |
18.72 |
|
|
|
| 32 |
B |
3011 |
2993 |
24.22 |
|
|
|
| 33 |
B |
2971 |
2954 |
40.19 |
|
|
|
| 34 |
B |
2970 |
2953 |
75.25 |
|
|
|
| 35 |
B |
2894 |
2877 |
118.33 |
|
|
|
| 36 |
B |
2856 |
2840 |
21.59 |
|
|
|
| 37 |
B |
1433 |
1424 |
4.02 |
|
|
|
| 38 |
B |
1428 |
1420 |
8.49 |
|
|
|
| 39 |
B |
1414 |
1406 |
7.44 |
|
|
|
| 40 |
B |
1399 |
1391 |
3.82 |
|
|
|
| 41 |
B |
1345 |
1337 |
11.19 |
|
|
|
| 42 |
B |
1332 |
1324 |
26.72 |
|
|
|
| 43 |
B |
1305 |
1297 |
29.62 |
|
|
|
| 44 |
B |
1253 |
1245 |
2.17 |
|
|
|
| 45 |
B |
1215 |
1208 |
5.81 |
|
|
|
| 46 |
B |
1137 |
1130 |
59.21 |
|
|
|
| 47 |
B |
1123 |
1116 |
126.09 |
|
|
|
| 48 |
B |
1077 |
1071 |
9.20 |
|
|
|
| 49 |
B |
1000 |
994 |
49.20 |
|
|
|
| 50 |
B |
899 |
893 |
0.78 |
|
|
|
| 51 |
B |
871 |
866 |
0.44 |
|
|
|
| 52 |
B |
760 |
756 |
3.43 |
|
|
|
| 53 |
B |
485 |
482 |
1.03 |
|
|
|
| 54 |
B |
319 |
317 |
0.56 |
|
|
|
| 55 |
B |
218 |
217 |
2.61 |
|
|
|
| 56 |
B |
167 |
166 |
2.86 |
|
|
|
| 57 |
B |
73 |
73 |
1.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41112.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 40874.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.104 |
| C2 |
0.032 |
1.184 |
0.891 |
| C3 |
-0.032 |
-1.184 |
0.891 |
| C4 |
0.000 |
2.392 |
-0.039 |
| C5 |
0.000 |
-2.392 |
-0.039 |
| C6 |
-1.280 |
2.473 |
-0.875 |
| C7 |
1.280 |
-2.473 |
-0.875 |
| H8 |
-0.846 |
1.207 |
1.588 |
| H9 |
0.952 |
1.200 |
1.529 |
| H10 |
0.846 |
-1.207 |
1.588 |
| H11 |
-0.952 |
-1.200 |
1.529 |
| H12 |
0.888 |
2.341 |
-0.706 |
| H13 |
0.122 |
3.307 |
0.580 |
| H14 |
-0.888 |
-2.341 |
-0.706 |
| H15 |
-0.122 |
-3.307 |
0.580 |
| H16 |
-1.256 |
3.333 |
-1.574 |
| H17 |
-1.414 |
1.547 |
-1.467 |
| H18 |
-2.175 |
2.589 |
-0.228 |
| H19 |
1.256 |
-3.333 |
-1.574 |
| H20 |
1.414 |
-1.547 |
-1.467 |
| H21 |
2.175 |
-2.589 |
-0.228 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4221 | 1.4221 | 2.3959 | 2.3959 | 2.9517 | 2.9517 | 2.0918 | 2.0921 | 2.0918 | 2.0921 | 2.6312 | 3.3429 | 2.6312 | 3.3429 | 3.9369 | 2.6190 | 3.3980 | 3.9369 | 2.6190 | 3.3980 |
C2 | 1.4221 | | 2.3684 | 1.5250 | 3.6946 | 2.5499 | 4.2485 | 1.1212 | 1.1193 | 2.6197 | 2.6565 | 2.1499 | 2.1474 | 3.9773 | 4.5037 | 3.5149 | 2.7899 | 2.8458 | 5.2891 | 3.8637 | 4.4814 | C3 | 1.4221 | 2.3684 | | 3.6946 | 1.5250 | 4.2485 | 2.5499 | 2.6197 | 2.6565 | 1.1212 | 1.1193 | 3.9773 | 4.5037 | 2.1499 | 2.1474 | 5.2891 | 3.8637 | 4.4814 | 3.5149 | 2.7899 | 2.8458 | C4 | 2.3959 | 1.5250 | 3.6946 | | 4.7833 | 1.5309 | 5.0993 | 2.1836 | 2.1872 | 4.0388 | 4.0330 | 1.1117 | 1.1114 | 4.8609 | 5.7331 | 2.1954 | 2.1800 | 2.1923 | 6.0581 | 4.4215 | 5.4386 | C5 | 2.3959 | 3.6946 | 1.5250 | 4.7833 | | 5.0993 | 1.5309 | 4.0388 | 4.0330 | 2.1836 | 2.1872 | 4.8609 | 5.7331 | 1.1117 | 1.1114 | 6.0581 | 4.4215 | 5.4386 | 2.1954 | 2.1800 | 2.1923 | C6 | 2.9517 | 2.5499 | 4.2485 | 1.5309 | 5.0993 | | 5.5696 | 2.8032 | 3.5182 | 4.9120 | 4.4021 | 2.1790 | 2.1854 | 4.8327 | 6.0715 | 1.1084 | 1.1077 | 1.1107 | 6.3740 | 4.8753 | 6.1634 | C7 | 2.9517 | 4.2485 | 2.5499 | 5.0993 | 1.5309 | 5.5696 | | 4.9120 | 4.4021 | 2.8032 | 3.5182 | 4.8327 | 6.0715 | 2.1790 | 2.1854 | 6.3740 | 4.8753 | 6.1634 | 1.1084 | 1.1077 | 1.1107 | H8 | 2.0918 | 1.1212 | 2.6197 | 2.1836 | 4.0388 | 2.8032 | 4.9120 | | 1.7988 | 2.9476 | 2.4099 | 3.0913 | 2.5225 | 4.2246 | 4.6809 | 3.8324 | 3.1261 | 2.6410 | 5.9181 | 4.6930 | 5.1803 | H9 | 2.0921 | 1.1193 | 2.6565 | 2.1872 | 4.0330 | 3.5182 | 4.4021 | 1.7988 | | 2.4099 | 3.0633 | 2.5095 | 2.4548 | 4.5734 | 4.7290 | 4.3646 | 3.8330 | 3.8460 | 5.5016 | 4.0909 | 4.3524 | H10 | 2.0918 | 2.6197 | 1.1212 | 4.0388 | 2.1836 | 4.9120 | 2.8032 | 2.9476 | 2.4099 | | 1.7988 | 4.2246 | 4.6809 | 3.0913 | 2.5225 | 5.9181 | 4.6930 | 5.1803 | 3.8324 | 3.1261 | 2.6410 | H11 | 2.0921 | 2.6565 | 1.1193 | 4.0330 | 2.1872 | 4.4021 | 3.5182 | 2.4099 | 3.0633 | 1.7988 | | 4.5734 | 4.7290 | 2.5095 | 2.4548 | 5.5016 | 4.0909 | 4.3524 | 4.3646 | 3.8330 | 3.8460 | H12 | 2.6312 | 2.1499 | 3.9773 | 1.1117 | 4.8609 | 2.1790 | 4.8327 | 3.0913 | 2.5095 | 4.2246 | 4.5734 | | 1.7814 | 5.0072 | 5.8792 | 2.5173 | 2.5516 | 3.1106 | 5.7512 | 3.9961 | 5.1177 | H13 | 3.3429 | 2.1474 | 4.5037 | 1.1114 | 5.7331 | 2.1854 | 6.0715 | 2.5225 | 2.4548 | 4.6809 | 4.7290 | 1.7814 | | 5.8792 | 6.6176 | 2.5572 | 3.1058 | 2.5382 | 7.0715 | 5.4236 | 6.2954 | H14 | 2.6312 | 3.9773 | 2.1499 | 4.8609 | 1.1117 | 4.8327 | 2.1790 | 4.2246 | 4.5734 | 3.0913 | 2.5095 | 5.0072 | 5.8792 | | 1.7814 | 5.7512 | 3.9961 | 5.1177 | 2.5173 | 2.5516 | 3.1106 | H15 | 3.3429 | 4.5037 | 2.1474 | 5.7331 | 1.1114 | 6.0715 | 2.1854 | 4.6809 | 4.7290 | 2.5225 | 2.4548 | 5.8792 | 6.6176 | 1.7814 | | 7.0715 | 5.4236 | 6.2954 | 2.5572 | 3.1058 | 2.5382 | H16 | 3.9369 | 3.5149 | 5.2891 | 2.1954 | 6.0581 | 1.1084 | 6.3740 | 3.8324 | 4.3646 | 5.9181 | 5.5016 | 2.5173 | 2.5572 | 5.7512 | 7.0715 | | 1.7960 | 1.7918 | 7.1229 | 5.5632 | 6.9755 | H17 | 2.6190 | 2.7899 | 3.8637 | 2.1800 | 4.4215 | 1.1077 | 4.8753 | 3.1261 | 3.8330 | 4.6930 | 4.0909 | 2.5516 | 3.1058 | 3.9961 | 5.4236 | 1.7960 | | 1.7897 | 5.5632 | 4.1913 | 5.6149 | H18 | 3.3980 | 2.8458 | 4.4814 | 2.1923 | 5.4386 | 1.1107 | 6.1634 | 2.6410 | 3.8460 | 5.1803 | 4.3524 | 3.1106 | 2.5382 | 5.1177 | 6.2954 | 1.7918 | 1.7897 | | 6.9755 | 5.6149 | 6.7636 | H19 | 3.9369 | 5.2891 | 3.5149 | 6.0581 | 2.1954 | 6.3740 | 1.1084 | 5.9181 | 5.5016 | 3.8324 | 4.3646 | 5.7512 | 7.0715 | 2.5173 | 2.5572 | 7.1229 | 5.5632 | 6.9755 | | 1.7960 | 1.7918 | H20 | 2.6190 | 3.8637 | 2.7899 | 4.4215 | 2.1800 | 4.8753 | 1.1077 | 4.6930 | 4.0909 | 3.1261 | 3.8330 | 3.9961 | 5.4236 | 2.5516 | 3.1058 | 5.5632 | 4.1913 | 5.6149 | 1.7960 | | 1.7897 | H21 | 3.3980 | 4.4814 | 2.8458 | 5.4386 | 2.1923 | 6.1634 | 1.1107 | 5.1803 | 4.3524 | 2.6410 | 3.8460 | 5.1177 | 6.2954 | 3.1106 | 2.5382 | 6.9755 | 5.6149 | 6.7636 | 1.7918 | 1.7897 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.725 |
|
O1 |
C2 |
H8 |
110.100 |
| O1 |
C2 |
H9 |
110.243 |
|
O1 |
C3 |
C5 |
108.725 |
| O1 |
C3 |
H10 |
110.100 |
|
O1 |
C3 |
H11 |
110.243 |
| C2 |
O1 |
C3 |
112.753 |
|
C2 |
C4 |
C6 |
113.114 |
| C2 |
C4 |
H12 |
108.230 |
|
C2 |
C4 |
H13 |
108.054 |
| C3 |
C5 |
C7 |
113.114 |
|
C3 |
C5 |
H14 |
108.230 |
| C3 |
C5 |
H15 |
108.054 |
|
C4 |
C2 |
H8 |
110.287 |
| C4 |
C2 |
H9 |
110.675 |
|
C4 |
C6 |
H16 |
111.567 |
| C4 |
C6 |
H17 |
110.393 |
|
C4 |
C6 |
H18 |
111.189 |
| C5 |
C3 |
H10 |
110.287 |
|
C5 |
C3 |
H11 |
110.675 |
| C5 |
C7 |
H19 |
111.567 |
|
C5 |
C7 |
H20 |
110.393 |
| C5 |
C7 |
H21 |
111.189 |
|
C6 |
C4 |
H12 |
110.082 |
| C6 |
C4 |
H13 |
110.597 |
|
C7 |
C5 |
H14 |
110.082 |
| C7 |
C5 |
H15 |
110.597 |
|
H8 |
C2 |
H9 |
106.805 |
| H10 |
C3 |
H11 |
106.805 |
|
H12 |
C4 |
H13 |
106.512 |
| H14 |
C5 |
H15 |
106.512 |
|
H16 |
C6 |
H17 |
108.278 |
| H16 |
C6 |
H18 |
107.696 |
|
H17 |
C6 |
H18 |
107.565 |
| H19 |
C7 |
H20 |
108.278 |
|
H19 |
C7 |
H21 |
107.696 |
| H20 |
C7 |
H21 |
107.565 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.281 |
|
|
|
| 2 |
C |
0.110 |
|
|
|
| 3 |
C |
0.110 |
|
|
|
| 4 |
C |
-0.070 |
|
|
|
| 5 |
C |
-0.070 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
C |
-0.058 |
|
|
|
| 8 |
H |
0.005 |
|
|
|
| 9 |
H |
0.008 |
|
|
|
| 10 |
H |
0.005 |
|
|
|
| 11 |
H |
0.008 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.019 |
|
|
|
| 14 |
H |
0.031 |
|
|
|
| 15 |
H |
0.019 |
|
|
|
| 16 |
H |
0.026 |
|
|
|
| 17 |
H |
0.042 |
|
|
|
| 18 |
H |
0.027 |
|
|
|
| 19 |
H |
0.026 |
|
|
|
| 20 |
H |
0.042 |
|
|
|
| 21 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.821 |
0.821 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.153 |
0.331 |
0.000 |
| y |
0.331 |
-44.259 |
0.000 |
| z |
0.000 |
0.000 |
-46.199 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.923 |
0.331 |
0.000 |
| y |
0.331 |
2.417 |
0.000 |
| z |
0.000 |
0.000 |
-0.493 |
|
| Polar |
|---|
| 3z2-r2 | -0.987 |
|---|
| x2-y2 | -2.893 |
|---|
| xy | 0.331 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.558 |
-0.582 |
0.000 |
| y |
-0.582 |
13.007 |
0.000 |
| z |
0.000 |
0.000 |
10.705 |
<r2> (average value of r
2) Å
2
| <r2> |
347.241 |
| (<r2>)1/2 |
18.634 |
