Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -311.869686 |
| Energy at 298.15K | -311.885217 |
| HF Energy | -311.869686 |
| Nuclear repulsion energy | 323.214900 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3069 |
3052 |
2.13 |
|
|
|
| 2 |
A |
3049 |
3031 |
50.38 |
|
|
|
| 3 |
A |
3011 |
2993 |
14.42 |
|
|
|
| 4 |
A |
2971 |
2954 |
4.51 |
|
|
|
| 5 |
A |
2970 |
2953 |
3.91 |
|
|
|
| 6 |
A |
2900 |
2883 |
9.85 |
|
|
|
| 7 |
A |
2869 |
2852 |
101.33 |
|
|
|
| 8 |
A |
1452 |
1444 |
3.12 |
|
|
|
| 9 |
A |
1434 |
1425 |
3.41 |
|
|
|
| 10 |
A |
1415 |
1407 |
6.65 |
|
|
|
| 11 |
A |
1399 |
1391 |
2.05 |
|
|
|
| 12 |
A |
1381 |
1373 |
2.57 |
|
|
|
| 13 |
A |
1343 |
1336 |
2.63 |
|
|
|
| 14 |
A |
1312 |
1305 |
0.76 |
|
|
|
| 15 |
A |
1262 |
1255 |
2.58 |
|
|
|
| 16 |
A |
1210 |
1203 |
0.39 |
|
|
|
| 17 |
A |
1132 |
1125 |
5.59 |
|
|
|
| 18 |
A |
1099 |
1093 |
6.90 |
|
|
|
| 19 |
A |
1066 |
1060 |
9.31 |
|
|
|
| 20 |
A |
924 |
919 |
6.76 |
|
|
|
| 21 |
A |
892 |
887 |
6.33 |
|
|
|
| 22 |
A |
883 |
878 |
1.46 |
|
|
|
| 23 |
A |
729 |
725 |
0.34 |
|
|
|
| 24 |
A |
468 |
465 |
0.18 |
|
|
|
| 25 |
A |
326 |
324 |
0.33 |
|
|
|
| 26 |
A |
264 |
262 |
0.65 |
|
|
|
| 27 |
A |
164 |
163 |
0.02 |
|
|
|
| 28 |
A |
114 |
113 |
0.00 |
|
|
|
| 29 |
A |
48 |
48 |
0.00 |
|
|
|
| 30 |
B |
3069 |
3051 |
41.95 |
|
|
|
| 31 |
B |
3049 |
3031 |
18.87 |
|
|
|
| 32 |
B |
3011 |
2993 |
24.28 |
|
|
|
| 33 |
B |
2971 |
2954 |
41.55 |
|
|
|
| 34 |
B |
2970 |
2953 |
73.96 |
|
|
|
| 35 |
B |
2893 |
2876 |
119.09 |
|
|
|
| 36 |
B |
2856 |
2840 |
20.70 |
|
|
|
| 37 |
B |
1432 |
1424 |
4.23 |
|
|
|
| 38 |
B |
1429 |
1420 |
8.48 |
|
|
|
| 39 |
B |
1414 |
1406 |
7.46 |
|
|
|
| 40 |
B |
1399 |
1391 |
3.97 |
|
|
|
| 41 |
B |
1344 |
1336 |
11.89 |
|
|
|
| 42 |
B |
1331 |
1323 |
25.95 |
|
|
|
| 43 |
B |
1304 |
1297 |
29.32 |
|
|
|
| 44 |
B |
1252 |
1245 |
2.26 |
|
|
|
| 45 |
B |
1216 |
1208 |
5.74 |
|
|
|
| 46 |
B |
1137 |
1130 |
61.09 |
|
|
|
| 47 |
B |
1123 |
1116 |
124.14 |
|
|
|
| 48 |
B |
1078 |
1072 |
9.04 |
|
|
|
| 49 |
B |
999 |
994 |
49.40 |
|
|
|
| 50 |
B |
897 |
892 |
0.77 |
|
|
|
| 51 |
B |
871 |
866 |
0.46 |
|
|
|
| 52 |
B |
760 |
756 |
3.38 |
|
|
|
| 53 |
B |
483 |
481 |
1.02 |
|
|
|
| 54 |
B |
316 |
314 |
0.58 |
|
|
|
| 55 |
B |
218 |
217 |
2.59 |
|
|
|
| 56 |
B |
168 |
167 |
2.88 |
|
|
|
| 57 |
B |
74 |
74 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41108.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 40870.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.103 |
| C2 |
0.032 |
1.184 |
0.890 |
| C3 |
-0.032 |
-1.184 |
0.890 |
| C4 |
0.000 |
2.392 |
-0.040 |
| C5 |
0.000 |
-2.392 |
-0.040 |
| C6 |
-1.281 |
2.475 |
-0.873 |
| C7 |
1.281 |
-2.475 |
-0.873 |
| H8 |
-0.846 |
1.207 |
1.588 |
| H9 |
0.952 |
1.200 |
1.528 |
| H10 |
0.846 |
-1.207 |
1.588 |
| H11 |
-0.952 |
-1.200 |
1.528 |
| H12 |
0.887 |
2.340 |
-0.708 |
| H13 |
0.124 |
3.307 |
0.579 |
| H14 |
-0.887 |
-2.340 |
-0.708 |
| H15 |
-0.124 |
-3.307 |
0.579 |
| H16 |
-1.258 |
3.336 |
-1.572 |
| H17 |
-1.417 |
1.550 |
-1.466 |
| H18 |
-2.175 |
2.592 |
-0.224 |
| H19 |
1.258 |
-3.336 |
-1.572 |
| H20 |
1.417 |
-1.550 |
-1.466 |
| H21 |
2.175 |
-2.592 |
-0.224 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4222 | 1.4222 | 2.3961 | 2.3961 | 2.9534 | 2.9534 | 2.0919 | 2.0921 | 2.0919 | 2.0921 | 2.6302 | 3.3430 | 2.6302 | 3.3430 | 3.9386 | 2.6215 | 3.3995 | 3.9386 | 2.6215 | 3.3995 |
C2 | 1.4222 | | 2.3685 | 1.5250 | 3.6948 | 2.5501 | 4.2499 | 1.1212 | 1.1193 | 2.6197 | 2.6566 | 2.1497 | 2.1474 | 3.9765 | 4.5038 | 3.5150 | 2.7906 | 2.8455 | 5.2906 | 3.8660 | 4.4826 | C3 | 1.4222 | 2.3685 | | 3.6948 | 1.5250 | 4.2499 | 2.5501 | 2.6197 | 2.6566 | 1.1212 | 1.1193 | 3.9765 | 4.5038 | 2.1497 | 2.1474 | 5.2906 | 3.8660 | 4.4826 | 3.5150 | 2.7906 | 2.8455 | C4 | 2.3961 | 1.5250 | 3.6948 | | 4.7838 | 1.5309 | 5.1017 | 2.1838 | 2.1869 | 4.0389 | 4.0332 | 1.1117 | 1.1114 | 4.8602 | 5.7333 | 2.1954 | 2.1801 | 2.1922 | 6.0609 | 4.4250 | 5.4410 | C5 | 2.3961 | 3.6948 | 1.5250 | 4.7838 | | 5.1017 | 1.5309 | 4.0389 | 4.0332 | 2.1838 | 2.1869 | 4.8602 | 5.7333 | 1.1117 | 1.1114 | 6.0609 | 4.4250 | 5.4410 | 2.1954 | 2.1801 | 2.1922 | C6 | 2.9534 | 2.5501 | 4.2499 | 1.5309 | 5.1017 | | 5.5748 | 2.8028 | 3.5181 | 4.9130 | 4.4027 | 2.1791 | 2.1854 | 4.8341 | 6.0730 | 1.1084 | 1.1077 | 1.1107 | 6.3799 | 4.8822 | 6.1682 | C7 | 2.9534 | 4.2499 | 2.5501 | 5.1017 | 1.5309 | 5.5748 | | 4.9130 | 4.4027 | 2.8028 | 3.5181 | 4.8341 | 6.0730 | 2.1791 | 2.1854 | 6.3799 | 4.8822 | 6.1682 | 1.1084 | 1.1077 | 1.1107 | H8 | 2.0919 | 1.1212 | 2.6197 | 2.1838 | 4.0389 | 2.8028 | 4.9130 | | 1.7988 | 2.9471 | 2.4099 | 3.0912 | 2.5234 | 4.2243 | 4.6803 | 3.8320 | 3.1258 | 2.6401 | 5.9194 | 4.6953 | 5.1807 | H9 | 2.0921 | 1.1193 | 2.6566 | 2.1869 | 4.0332 | 3.5181 | 4.4027 | 1.7988 | | 2.4099 | 3.0638 | 2.5096 | 2.4537 | 4.5727 | 4.7295 | 4.3644 | 3.8338 | 3.8451 | 5.5023 | 4.0918 | 4.3529 | H10 | 2.0919 | 2.6197 | 1.1212 | 4.0389 | 2.1838 | 4.9130 | 2.8028 | 2.9471 | 2.4099 | | 1.7988 | 4.2243 | 4.6803 | 3.0912 | 2.5234 | 5.9194 | 4.6953 | 5.1807 | 3.8320 | 3.1258 | 2.6401 | H11 | 2.0921 | 2.6566 | 1.1193 | 4.0332 | 2.1869 | 4.4027 | 3.5181 | 2.4099 | 3.0638 | 1.7988 | | 4.5727 | 4.7295 | 2.5096 | 2.4537 | 5.5023 | 4.0918 | 4.3529 | 4.3644 | 3.8338 | 3.8451 | H12 | 2.6302 | 2.1497 | 3.9765 | 1.1117 | 4.8602 | 2.1791 | 4.8341 | 3.0912 | 2.5096 | 4.2243 | 4.5727 | | 1.7814 | 5.0046 | 5.8787 | 2.5179 | 2.5514 | 3.1106 | 5.7527 | 3.9981 | 5.1199 | H13 | 3.3430 | 2.1474 | 4.5038 | 1.1114 | 5.7333 | 2.1854 | 6.0730 | 2.5234 | 2.4537 | 4.6803 | 4.7295 | 1.7814 | | 5.8787 | 6.6178 | 2.5569 | 3.1059 | 2.5384 | 7.0733 | 5.4260 | 6.2965 | H14 | 2.6302 | 3.9765 | 2.1497 | 4.8602 | 1.1117 | 4.8341 | 2.1791 | 4.2243 | 4.5727 | 3.0912 | 2.5096 | 5.0046 | 5.8787 | | 1.7814 | 5.7527 | 3.9981 | 5.1199 | 2.5179 | 2.5514 | 3.1106 | H15 | 3.3430 | 4.5038 | 2.1474 | 5.7333 | 1.1114 | 6.0730 | 2.1854 | 4.6803 | 4.7295 | 2.5234 | 2.4537 | 5.8787 | 6.6178 | 1.7814 | | 7.0733 | 5.4260 | 6.2965 | 2.5569 | 3.1059 | 2.5384 | H16 | 3.9386 | 3.5150 | 5.2906 | 2.1954 | 6.0609 | 1.1084 | 6.3799 | 3.8320 | 4.3644 | 5.9194 | 5.5023 | 2.5179 | 2.5569 | 5.7527 | 7.0733 | | 1.7961 | 1.7917 | 7.1298 | 5.5708 | 6.9812 | H17 | 2.6215 | 2.7906 | 3.8660 | 2.1801 | 4.4250 | 1.1077 | 4.8822 | 3.1258 | 3.8338 | 4.6953 | 4.0918 | 2.5514 | 3.1059 | 3.9981 | 5.4260 | 1.7961 | | 1.7897 | 5.5708 | 4.2000 | 5.6215 | H18 | 3.3995 | 2.8455 | 4.4826 | 2.1922 | 5.4410 | 1.1107 | 6.1682 | 2.6401 | 3.8451 | 5.1807 | 4.3529 | 3.1106 | 2.5384 | 5.1199 | 6.2965 | 1.7917 | 1.7897 | | 6.9812 | 5.6215 | 6.7675 | H19 | 3.9386 | 5.2906 | 3.5150 | 6.0609 | 2.1954 | 6.3799 | 1.1084 | 5.9194 | 5.5023 | 3.8320 | 4.3644 | 5.7527 | 7.0733 | 2.5179 | 2.5569 | 7.1298 | 5.5708 | 6.9812 | | 1.7961 | 1.7917 | H20 | 2.6215 | 3.8660 | 2.7906 | 4.4250 | 2.1801 | 4.8822 | 1.1077 | 4.6953 | 4.0918 | 3.1258 | 3.8338 | 3.9981 | 5.4260 | 2.5514 | 3.1059 | 5.5708 | 4.2000 | 5.6215 | 1.7961 | | 1.7897 | H21 | 3.3995 | 4.4826 | 2.8455 | 5.4410 | 2.1922 | 6.1682 | 1.1107 | 5.1807 | 4.3529 | 2.6401 | 3.8451 | 5.1199 | 6.2965 | 3.1106 | 2.5384 | 6.9812 | 5.6215 | 6.7675 | 1.7917 | 1.7897 | |
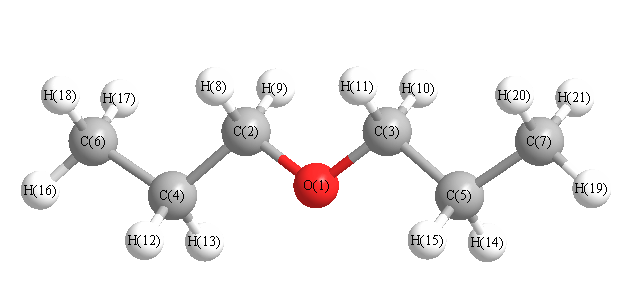 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.736 |
|
O1 |
C2 |
H8 |
110.106 |
| O1 |
C2 |
H9 |
110.237 |
|
O1 |
C3 |
C5 |
108.736 |
| O1 |
C3 |
H10 |
110.106 |
|
O1 |
C3 |
H11 |
110.237 |
| C2 |
O1 |
C3 |
112.749 |
|
C2 |
C4 |
C6 |
113.126 |
| C2 |
C4 |
H12 |
108.212 |
|
C2 |
C4 |
H13 |
108.051 |
| C3 |
C5 |
C7 |
113.126 |
|
C3 |
C5 |
H14 |
108.212 |
| C3 |
C5 |
H15 |
108.051 |
|
C4 |
C2 |
H8 |
110.298 |
| C4 |
C2 |
H9 |
110.651 |
|
C4 |
C6 |
H16 |
111.573 |
| C4 |
C6 |
H17 |
110.397 |
|
C4 |
C6 |
H18 |
111.181 |
| C5 |
C3 |
H10 |
110.298 |
|
C5 |
C3 |
H11 |
110.651 |
| C5 |
C7 |
H19 |
111.573 |
|
C5 |
C7 |
H20 |
110.397 |
| C5 |
C7 |
H21 |
111.181 |
|
C6 |
C4 |
H12 |
110.090 |
| C6 |
C4 |
H13 |
110.599 |
|
C7 |
C5 |
H14 |
110.090 |
| C7 |
C5 |
H15 |
110.599 |
|
H8 |
C2 |
H9 |
106.808 |
| H10 |
C3 |
H11 |
106.808 |
|
H12 |
C4 |
H13 |
106.510 |
| H14 |
C5 |
H15 |
106.510 |
|
H16 |
C6 |
H17 |
108.282 |
| H16 |
C6 |
H18 |
107.691 |
|
H17 |
C6 |
H18 |
107.564 |
| H19 |
C7 |
H20 |
108.282 |
|
H19 |
C7 |
H21 |
107.691 |
| H20 |
C7 |
H21 |
107.564 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.281 |
|
|
|
| 2 |
C |
0.110 |
|
|
|
| 3 |
C |
0.110 |
|
|
|
| 4 |
C |
-0.070 |
|
|
|
| 5 |
C |
-0.070 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
C |
-0.058 |
|
|
|
| 8 |
H |
0.005 |
|
|
|
| 9 |
H |
0.008 |
|
|
|
| 10 |
H |
0.005 |
|
|
|
| 11 |
H |
0.008 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.019 |
|
|
|
| 14 |
H |
0.031 |
|
|
|
| 15 |
H |
0.019 |
|
|
|
| 16 |
H |
0.026 |
|
|
|
| 17 |
H |
0.042 |
|
|
|
| 18 |
H |
0.027 |
|
|
|
| 19 |
H |
0.026 |
|
|
|
| 20 |
H |
0.042 |
|
|
|
| 21 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.821 |
0.821 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.153 |
0.331 |
0.000 |
| y |
0.331 |
-44.254 |
0.000 |
| z |
0.000 |
0.000 |
-46.200 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.926 |
0.331 |
0.000 |
| y |
0.331 |
2.422 |
0.000 |
| z |
0.000 |
0.000 |
-0.497 |
|
| Polar |
|---|
| 3z2-r2 | -0.993 |
|---|
| x2-y2 | -2.899 |
|---|
| xy | 0.331 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.559 |
-0.583 |
0.000 |
| y |
-0.583 |
13.009 |
0.000 |
| z |
0.000 |
0.000 |
10.702 |
<r2> (average value of r
2) Å
2
| <r2> |
347.451 |
| (<r2>)1/2 |
18.640 |
