return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H18 (Octane)

using model chemistry: PBEPBE/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at PBEPBE/cc-pVDZ
 hartrees
Energy at 0K-315.245675
Energy at 298.15K-315.264862
Nuclear repulsion energy372.046479
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3048 3030 0.00      
2 Ag 2963 2946 0.00      
3 Ag 2949 2932 0.00      
4 Ag 2935 2918 0.00      
5 Ag 2934 2917 0.00      
6 Ag 1434 1425 0.00      
7 Ag 1421 1412 0.00      
8 Ag 1411 1403 0.00      
9 Ag 1407 1399 0.00      
10 Ag 1353 1345 0.00      
11 Ag 1346 1339 0.00      
12 Ag 1326 1318 0.00      
13 Ag 1241 1234 0.00      
14 Ag 1124 1117 0.00      
15 Ag 1069 1062 0.00      
16 Ag 1053 1047 0.00      
17 Ag 991 986 0.00      
18 Ag 889 884 0.00      
19 Ag 456 454 0.00      
20 Ag 271 269 0.00      
21 Ag 188 187 0.00      
22 Au 3040 3022 92.75      
23 Au 3001 2983 144.84      
24 Au 2984 2967 0.56      
25 Au 2962 2945 0.27      
26 Au 1419 1411 12.68      
27 Au 1284 1276 0.58      
28 Au 1265 1257 0.21      
29 Au 1181 1174 0.07      
30 Au 999 993 0.36      
31 Au 848 843 1.11      
32 Au 735 731 1.72      
33 Au 721 717 6.78      
34 Au 245 243 0.00      
35 Au 164 163 0.00      
36 Au 75 75 0.00      
37 Au 47 47 0.00      
38 Bg 3040 3022 0.00      
39 Bg 2994 2976 0.00      
40 Bg 2972 2955 0.00      
41 Bg 2958 2941 0.00      
42 Bg 1419 1411 0.00      
43 Bg 1284 1277 0.00      
44 Bg 1273 1265 0.00      
45 Bg 1227 1219 0.00      
46 Bg 1152 1146 0.00      
47 Bg 927 921 0.00      
48 Bg 779 775 0.00      
49 Bg 721 717 0.00      
50 Bg 248 246 0.00      
51 Bg 140 139 0.00      
52 Bg 111 111 0.00      
53 Bu 3048 3030 67.19      
54 Bu 2963 2946 76.56      
55 Bu 2950 2933 158.38      
56 Bu 2942 2925 26.61      
57 Bu 2932 2915 2.60      
58 Bu 1438 1429 6.92      
59 Bu 1428 1420 1.52      
60 Bu 1414 1405 0.84      
61 Bu 1407 1399 0.13      
62 Bu 1350 1342 2.96      
63 Bu 1344 1337 0.89      
64 Bu 1289 1281 3.42      
65 Bu 1192 1185 6.60      
66 Bu 1073 1067 2.57      
67 Bu 1056 1050 1.00      
68 Bu 1015 1009 0.48      
69 Bu 872 867 3.76      
70 Bu 460 457 0.12      
71 Bu 325 324 0.05      
72 Bu 78 78 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 52296.5 cm-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 51993.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at PBEPBE/cc-pVDZ
ABC
0.35343 0.01615 0.01581

See section I.F.4 to change rotational constant units
Geometric Data calculated at PBEPBE/cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.002 0.767 0.000
C2 -0.002 -0.767 0.000
C3 -1.400 1.388 0.000
C4 1.400 -1.388 0.000
C5 -1.400 2.921 0.000
C6 1.400 -2.921 0.000
C7 -2.805 3.529 0.000
C8 2.805 -3.529 0.000
H9 0.567 1.133 0.888
H10 0.567 1.133 -0.888
H11 -0.567 -1.133 0.888
H12 -0.567 -1.133 -0.888
H13 -1.965 1.021 -0.887
H14 -1.965 1.021 0.887
H15 1.965 -1.021 -0.887
H16 1.965 -1.021 0.887
H17 -0.834 3.286 0.887
H18 -0.834 3.286 -0.887
H19 0.834 -3.286 0.887
H20 0.834 -3.286 -0.887
H21 -2.773 4.637 0.000
H22 -3.382 3.211 -0.894
H23 -3.382 3.211 0.894
H24 2.773 -4.637 0.000
H25 3.382 -3.211 -0.894
H26 3.382 -3.211 0.894

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24 H25 H26
C11.53311.53302.56802.57033.94383.93795.12911.11411.11412.17262.17262.17272.17272.79952.79952.79882.79884.23114.23114.76254.26904.26906.07295.29595.2959
C21.53312.56801.53303.94382.57035.12913.93792.17262.17261.11411.11412.79952.79952.17272.17274.23114.23112.79882.79886.07295.29595.29594.76254.26904.2690
C31.53302.56803.94201.53355.13842.56116.46922.17252.17252.79932.79931.11431.11434.23214.23212.17022.17025.25555.25553.52782.83782.83787.32876.69426.6942
C42.56801.53303.94205.13841.53356.46922.56112.79932.79932.17252.17254.23214.23211.11431.11435.25555.25552.17022.17027.32876.69426.69423.52782.83782.8378
C52.57033.94381.53355.13846.47841.53077.69942.80212.80214.23314.23312.17222.17225.25825.25821.11331.11336.65636.65632.19792.19372.19378.63377.82747.8274
C63.94382.57035.13841.53356.47847.69941.53074.23314.23312.80212.80215.25825.25822.17222.17226.65636.65631.11331.11338.63377.82747.82742.19792.19372.1937
C73.93795.12912.56116.46921.53077.69949.01554.23014.23015.24705.24702.78982.78986.65126.65122.17432.17437.77647.77641.10881.11051.11059.88929.19259.1925
C85.12913.93796.46922.56117.69941.53079.01555.24705.24704.23014.23016.65126.65122.78982.78987.77647.77642.17432.17439.88929.19259.19251.10881.11051.1105
H91.11412.17262.17252.79932.80214.23314.23015.24701.77502.53403.09393.09412.53433.12182.56812.56863.12204.42724.76964.92144.80454.46216.24125.47465.1767
H101.11412.17262.17252.79932.80214.23314.23015.24701.77503.09392.53402.53433.09412.56813.12183.12202.56864.76964.42724.92144.46214.80456.24125.17675.4746
H112.17261.11412.79932.17254.23312.80215.24704.23012.53403.09391.77503.12182.56813.09412.53434.42724.76962.56863.12206.24125.47465.17674.92144.80454.4621
H122.17261.11412.79932.17254.23312.80215.24704.23013.09392.53401.77502.56813.12182.53433.09414.76964.42723.12202.56866.24125.17675.47464.92144.46214.8045
H132.17272.79951.11434.23212.17225.25822.78986.65123.09412.53433.12182.56811.77494.42884.77133.09152.53155.43455.13673.81042.60863.15877.43316.81917.0479
H142.17272.79951.11434.23212.17225.25822.78986.65122.53433.09412.56813.12181.77494.77134.42882.53153.09155.13675.43453.81043.15872.60867.43317.04796.8191
H152.79952.17274.23211.11435.25822.17226.65122.78983.12182.56813.09412.53434.42884.77131.77495.43455.13673.09152.53157.43316.81917.04793.81042.60863.1587
H162.79952.17274.23211.11435.25822.17226.65122.78982.56813.12182.53433.09414.77134.42881.77495.13675.43452.53153.09157.43317.04796.81913.81043.15872.6086
H172.79884.23112.17025.25551.11336.65632.17437.77642.56863.12204.42724.76963.09152.53155.43455.13671.77416.78047.00862.52403.10912.54868.75087.94747.7453
H182.79884.23112.17025.25551.11336.65632.17437.77643.12202.56864.76964.42722.53153.09155.13675.43451.77417.00866.78042.52402.54863.10918.75087.74537.9474
H194.23112.79885.25552.17026.65631.11337.77642.17434.42724.76962.56863.12205.43455.13673.09152.53156.78047.00861.77418.75087.94747.74532.52403.10912.5486
H204.23112.79885.25552.17026.65631.11337.77642.17434.76964.42723.12202.56865.13675.43452.53153.09157.00866.78041.77418.75087.74537.94742.52402.54863.1091
H214.76256.07293.52787.32872.19798.63371.10889.88924.92144.92146.24126.24123.81043.81047.43317.43312.52402.52408.75088.75081.78991.789910.806210.013910.0139
H224.26905.29592.83786.69422.19377.82741.11059.19254.80454.46215.47465.17672.60863.15876.81917.04793.10912.54867.94747.74531.78991.787510.01399.32729.4969
H234.26905.29592.83786.69422.19377.82741.11059.19254.46214.80455.17675.47463.15872.60867.04796.81912.54863.10917.74537.94741.78991.787510.01399.49699.3272
H246.07294.76257.32873.52788.63372.19799.88921.10886.24126.24124.92144.92147.43317.43313.81043.81048.75088.75082.52402.524010.806210.013910.01391.78991.7899
H255.29594.26906.69422.83787.82742.19379.19251.11055.47465.17674.80454.46216.81917.04792.60863.15877.94747.74533.10912.548610.01399.32729.49691.78991.7875
H265.29594.26906.69422.83787.82742.19379.19251.11055.17675.47464.46214.80457.04796.81913.15872.60867.74537.94742.54863.109110.01399.49699.32721.78991.7875

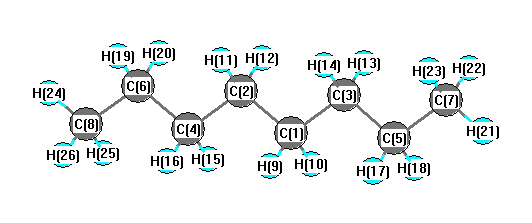
picture of Octane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 113.767 C1 C2 H11 109.283
C1 C2 H12 109.283 C1 C3 C5 113.903
C1 C3 H13 109.297 C1 C3 H14 109.297
C2 C1 C3 113.767 C2 C1 H9 109.283
C2 C1 H10 109.283 C2 C4 C6 113.903
C2 C4 H15 109.297 C2 C4 H16 109.297
C3 C1 H9 109.287 C3 C1 H10 109.287
C3 C5 C7 113.399 C3 C5 H17 109.124
C3 C5 H18 109.124 C4 C2 H11 109.287
C4 C2 H12 109.287 C4 C6 C8 113.399
C4 C6 H19 109.124 C4 C6 H20 109.124
C5 C3 H13 109.217 C5 C3 H14 109.217
C5 C7 H21 111.754 C5 C7 H22 111.315
C5 C7 H23 111.315 C6 C4 H15 109.217
C6 C4 H16 109.217 C6 C8 H24 111.754
C6 C8 H25 111.315 C6 C8 H26 111.315
C7 C5 H17 109.629 C7 C5 H18 109.629
C8 C6 H19 109.629 C8 C6 H20 109.629
H9 C1 H10 105.612 H11 C2 H12 105.612
H13 C3 H14 105.576 H15 C4 H16 105.576
H17 C5 H18 105.643 H19 C6 H20 105.643
H21 C7 H22 107.521 H21 C7 H23 107.521
H22 C7 H23 107.187 H24 C8 H25 107.521
H24 C8 H26 107.521 H25 C8 H26 107.187
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.034      
2 C -0.034      
3 C -0.038      
4 C -0.038      
5 C -0.068      
6 C -0.068      
7 C -0.044      
8 C -0.044      
9 H 0.015      
10 H 0.015      
11 H 0.015      
12 H 0.015      
13 H 0.015      
14 H 0.015      
15 H 0.015      
16 H 0.015      
17 H 0.019      
18 H 0.019      
19 H 0.019      
20 H 0.019      
21 H 0.025      
22 H 0.030      
23 H 0.030      
24 H 0.025      
25 H 0.030      
26 H 0.030      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 0.000 0.000
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -56.377 0.398 0.000
y 0.398 -55.665 0.000
z 0.000 0.000 -54.450
Traceless
 xyz
x -1.319 0.398 0.000
y 0.398 -0.251 0.000
z 0.000 0.000 1.570
Polar
3z2-r23.140
x2-y2-0.712
xy0.398
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 14.718 -2.534 0.000
y -2.534 16.338 0.000
z 0.000 0.000 12.204


<r2> (average value of r2) Å2
<r2> 688.113
(<r2>)1/2 26.232