return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H20 (Nonane)

using model chemistry: PBEPBEultrafine/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at PBEPBEultrafine/cc-pVDZ
 hartrees
Energy at 0K-354.503279
Energy at 298.15K-354.524588
HF Energy-354.503279
Nuclear repulsion energy441.832887
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3048 3030 37.84      
2 A1 2963 2946 27.29      
3 A1 2951 2934 179.69      
4 A1 2943 2926 36.39      
5 A1 2935 2918 1.14      
6 A1 2933 2916 2.18      
7 A1 1437 1428 7.30      
8 A1 1429 1421 0.70      
9 A1 1415 1407 0.14      
10 A1 1410 1402 0.00      
11 A1 1406 1397 0.01      
12 A1 1351 1343 0.09      
13 A1 1348 1340 1.47      
14 A1 1309 1301 0.02      
15 A1 1227 1220 0.00      
16 A1 1123 1117 0.65      
17 A1 1066 1059 0.64      
18 A1 1034 1028 0.35      
19 A1 1000 995 0.23      
20 A1 881 876 1.32      
21 A1 484 481 0.04      
22 A1 282 281 0.03      
23 A1 237 236 0.00      
24 A1 60 60 0.00      
25 A2 3039 3022 0.00      
26 A2 2996 2978 0.00      
27 A2 2977 2960 0.00      
28 A2 2961 2944 0.00      
29 A2 1420 1412 0.00      
30 A2 1285 1277 0.00      
31 A2 1268 1261 0.00      
32 A2 1251 1244 0.00      
33 A2 1174 1168 0.00      
34 A2 1001 995 0.00      
35 A2 872 867 0.00      
36 A2 754 749 0.00      
37 A2 719 715 0.00      
38 A2 249 248 0.00      
39 A2 162 161 0.00      
40 A2 98 97 0.00      
41 A2 59 59 0.00      
42 B1 3039 3022 93.34      
43 B1 3002 2984 173.03      
44 B1 2988 2970 1.44      
45 B1 2968 2950 0.19      
46 B1 2958 2941 0.05      
47 B1 1420 1412 12.67      
48 B1 1280 1273 0.16      
49 B1 1275 1268 0.65      
50 B1 1214 1207 0.00      
51 B1 1152 1145 0.11      
52 B1 941 935 0.61      
53 B1 806 801 1.10      
54 B1 726 722 2.21      
55 B1 720 716 7.00      
56 B1 249 247 0.00      
57 B1 156 155 0.00      
58 B1 116 116 0.00      
59 B1 39 39 0.00      
60 B2 3048 3030 29.71      
61 B2 2963 2946 49.53      
62 B2 2950 2932 0.02      
63 B2 2937 2920 0.01      
64 B2 2933 2916 0.02      
65 B2 1434 1425 0.59      
66 B2 1422 1414 0.63      
67 B2 1410 1402 0.70      
68 B2 1406 1398 0.03      
69 B2 1353 1345 0.00      
70 B2 1348 1340 1.70      
71 B2 1336 1329 1.35      
72 B2 1270 1263 3.16      
73 B2 1184 1177 6.81      
74 B2 1074 1068 1.79      
75 B2 1062 1056 0.08      
76 B2 1055 1049 0.05      
77 B2 994 988 0.12      
78 B2 880 875 2.56      
79 B2 446 444 0.10      
80 B2 386 384 0.03      
81 B2 155 154 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 58323.0 cm-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 57984.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at PBEPBEultrafine/cc-pVDZ
ABC
0.30840 0.01142 0.01123

See section I.F.4 to change rotational constant units
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.360
C2 0.000 1.285 -0.477
C3 0.000 -1.285 -0.477
C4 0.000 2.570 0.359
C5 0.000 -2.570 0.359
C6 0.000 3.855 -0.478
C7 0.000 -3.855 -0.478
C8 0.000 5.131 0.368
C9 0.000 -5.131 0.368
H10 0.887 0.000 1.034
H11 -0.887 0.000 1.034
H12 0.887 1.284 -1.151
H13 -0.887 1.284 -1.151
H14 -0.887 -1.284 -1.151
H15 0.887 -1.284 -1.151
H16 0.888 2.571 1.033
H17 -0.888 2.571 1.033
H18 -0.888 -2.571 1.033
H19 0.888 -2.571 1.033
H20 -0.887 3.852 -1.150
H21 0.887 3.852 -1.150
H22 0.887 -3.852 -1.150
H23 -0.887 -3.852 -1.150
H24 0.000 6.042 -0.263
H25 -0.894 5.179 1.025
H26 0.894 5.179 1.025
H27 0.000 -6.042 -0.263
H28 0.894 -5.179 1.025
H29 -0.894 -5.179 1.025

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24 H25 H26 H27 H28 H29
C11.53321.53322.56972.56973.94463.94465.13105.13101.11421.11422.17232.17232.17232.17232.80162.80162.80162.80164.23154.23154.23154.23156.07455.29785.29786.07455.29785.2978
C21.53322.56941.53303.94402.56995.13933.93806.47102.17242.17241.11421.11422.80032.80032.17292.17294.23464.23462.79852.79855.25605.25604.76264.26884.26887.33036.69616.6961
C31.53322.56943.94401.53305.13932.56996.47103.93802.17242.17242.80032.80031.11421.11424.23464.23462.17292.17295.25605.25602.79852.79857.33036.69616.69614.76264.26884.2688
C42.56971.53303.94405.13931.53346.47862.56137.70062.80102.80102.17232.17234.23334.23331.11441.11445.25975.25972.17022.17026.65606.65603.52822.83772.83778.63467.82877.8287
C52.56973.94401.53305.13936.47861.53347.70062.56132.80102.80104.23334.23332.17232.17235.25975.25971.11441.11446.65606.65602.17022.17028.63467.82877.82873.52822.83772.8377
C63.94462.56995.13931.53346.47867.70931.53109.02534.23434.23432.80112.80115.25845.25842.17222.17226.65996.65991.11331.11337.78667.78662.19832.19362.19369.89949.20169.2016
C73.94465.13932.56996.47861.53347.70939.02531.53104.23434.23435.25845.25842.80112.80116.65996.65992.17222.17227.78667.78661.11331.11339.89949.20169.20162.19832.19362.1936
C85.13103.93806.47102.56137.70061.53109.025310.26195.24955.24954.22964.22966.65216.65212.79022.79027.78117.78112.17432.17439.15349.15341.10881.11051.110511.191310.369810.3698
C95.13106.47103.93807.70062.56139.02531.531010.26195.24955.24956.65216.65214.22964.22967.78117.78112.79022.79029.15349.15342.17432.174311.191310.369810.36981.10881.11051.1105
H101.11422.17242.17242.80102.80104.23434.23435.24955.24951.77492.53393.09363.09362.53392.57073.12393.12392.57074.77034.42804.42804.77036.24355.47705.17936.24355.17935.4770
H111.11422.17242.17242.80102.80104.23434.23435.24955.24951.77493.09362.53392.53393.09363.12392.57072.57073.12394.42804.77034.77034.42806.24355.17935.47706.24355.47705.1793
H122.17231.11422.80032.17234.23332.80115.25844.22966.65212.53393.09361.77483.12232.56882.53433.09414.77294.43063.12112.56765.13645.43434.92074.80384.46147.43366.82017.0488
H132.17231.11422.80032.17234.23332.80115.25844.22966.65213.09362.53391.77482.56883.12233.09412.53434.43064.77292.56763.12115.43435.13644.92074.46144.80387.43367.04886.8201
H142.17232.80031.11424.23332.17235.25842.80116.65214.22963.09362.53393.12232.56881.77484.77294.43062.53433.09415.13645.43433.12112.56767.43366.82017.04884.92074.80384.4614
H152.17232.80031.11424.23332.17235.25842.80116.65214.22962.53393.09362.56883.12231.77484.43064.77293.09412.53435.43435.13642.56763.12117.43367.04886.82014.92074.46144.8038
H162.80162.17294.23461.11445.25972.17226.65992.79027.78112.57073.12392.53433.09414.77294.43061.77505.43915.14143.09162.53166.78367.01193.81083.15882.60868.75537.75007.9521
H172.80162.17294.23461.11445.25972.17226.65992.79027.78113.12392.57073.09412.53434.43064.77291.77505.14145.43912.53163.09167.01196.78363.81082.60863.15888.75537.95217.7500
H182.80164.23462.17295.25971.11446.65992.17227.78112.79023.12392.57074.77294.43062.53433.09415.43915.14141.77506.78367.01193.09162.53168.75537.75007.95213.81083.15882.6086
H192.80164.23462.17295.25971.11446.65992.17227.78112.79022.57073.12394.43064.77293.09412.53435.14145.43911.77507.01196.78362.53163.09168.75537.95217.75003.81082.60863.1588
H204.23152.79855.25602.17026.65601.11337.78662.17439.15344.77034.42803.12112.56765.13645.43433.09162.53166.78367.01191.77417.90567.70392.52422.54843.10909.97369.45889.2896
H214.23152.79855.25602.17026.65601.11337.78662.17439.15344.42804.77032.56763.12115.43435.13642.53163.09167.01196.78361.77417.70397.90562.52423.10902.54849.97369.28969.4588
H224.23155.25602.79856.65602.17027.78661.11339.15342.17434.42804.77035.13645.43433.12112.56766.78367.01193.09162.53167.90567.70391.77419.97369.45889.28962.52422.54843.1090
H234.23155.25602.79856.65602.17027.78661.11339.15342.17434.77034.42805.43435.13642.56763.12117.01196.78362.53163.09167.70397.90561.77419.97369.28969.45882.52423.10902.5484
H246.07454.76267.33033.52828.63462.19839.89941.108811.19136.24356.24354.92074.92077.43367.43363.81083.81088.75538.75532.52422.52429.97369.97361.79011.790112.085011.330811.3308
H255.29784.26886.69612.83777.82872.19369.20161.110510.36985.47705.17934.80384.46146.82017.04883.15882.60867.75007.95212.54843.10909.45889.28961.79011.787611.330810.511710.3586
H265.29784.26886.69612.83777.82872.19369.20161.110510.36985.17935.47704.46144.80387.04886.82012.60863.15887.95217.75003.10902.54849.28969.45881.79011.787611.330810.358610.5117
H276.07457.33034.76268.63463.52829.89942.198311.19131.10886.24356.24357.43367.43364.92074.92078.75538.75533.81083.81089.97369.97362.52422.524212.085011.330811.33081.79011.7901
H285.29786.69614.26887.82872.83779.20162.193610.36981.11055.17935.47706.82017.04884.80384.46147.75007.95213.15882.60869.45889.28962.54843.109011.330810.511710.35861.79011.7876
H295.29786.69614.26887.82872.83779.20162.193610.36981.11055.47705.17937.04886.82014.46144.80387.95217.75002.60863.15889.28969.45883.10902.548411.330810.358610.51171.79011.7876

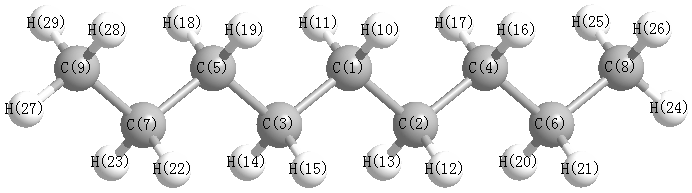
picture of Nonane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 113.873 C1 C2 H12 109.258
C1 C2 H13 109.258 C1 C3 C5 113.873
C1 C3 H14 109.258 C1 C3 H15 109.258
C2 C1 C3 113.844 C2 C1 H10 109.270
C2 C1 H11 109.270 C2 C4 C6 113.880
C2 C4 H16 109.303 C2 C4 H17 109.303
C3 C1 H10 109.270 C3 C1 H11 109.270
C3 C5 C7 113.880 C3 C5 H18 109.303
C3 C5 H19 109.303 C4 C2 H12 109.267
C4 C2 H13 109.267 C4 C6 C8 113.406
C4 C6 H20 109.131 C4 C6 H21 109.131
C5 C3 H14 109.267 C5 C3 H15 109.267
C5 C7 C9 113.406 C5 C7 H22 109.131
C5 C7 H23 109.131 C6 C4 H16 109.222
C6 C4 H17 109.222 C6 C8 H24 111.769
C6 C8 H25 111.292 C6 C8 H26 111.292
C7 C5 H14 96.693 C7 C5 H15 96.693
C7 C9 H27 111.769 C7 C9 H28 111.292
C7 C9 H29 111.292 C8 C6 H20 109.616
C8 C6 H21 109.616 C9 C7 H22 109.616
C9 C7 H23 109.616 H10 C1 H11 105.591
H12 C2 H13 105.588 H14 C3 H15 105.588
H16 C4 H17 105.579 H18 C5 H19 105.579
H20 C6 H21 105.648 H22 C7 H23 105.648
H24 C8 H25 107.530 H24 C8 H26 107.530
H25 C8 H26 107.199 H27 C9 H28 107.530
H27 C9 H29 107.530 H28 C9 H29 107.199
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.034      
2 C -0.034      
3 C -0.034      
4 C -0.037      
5 C -0.037      
6 C -0.068      
7 C -0.068      
8 C -0.044      
9 C -0.044      
10 H 0.016      
11 H 0.016      
12 H 0.015      
13 H 0.015      
14 H 0.015      
15 H 0.015      
16 H 0.015      
17 H 0.015      
18 H 0.015      
19 H 0.015      
20 H 0.019      
21 H 0.019      
22 H 0.019      
23 H 0.019      
24 H 0.025      
25 H 0.030      
26 H 0.030      
27 H 0.025      
28 H 0.030      
29 H 0.030      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 -0.064 0.064
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -61.049 0.000 0.000
y 0.000 -63.063 0.000
z 0.000 0.000 -62.591
Traceless
 xyz
x 1.779 0.000 0.000
y 0.000 -1.243 0.000
z 0.000 0.000 -0.536
Polar
3z2-r2-1.071
x2-y22.014
xy0.000
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 13.607 0.000 0.000
y 0.000 20.752 0.000
z 0.000 0.000 14.366


<r2> (average value of r2) Å2
<r2> 949.812
(<r2>)1/2 30.819