return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-194.695345
Energy at 298.15K-194.703860
HF Energy-193.955511
Nuclear repulsion energy166.477426
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3171 3034 5.88      
2 A' 3168 3030 45.50      
3 A' 3142 3006 12.86      
4 A' 3102 2968 36.37      
5 A' 3082 2948 11.69      
6 A' 1789 1712 9.62      
7 A' 1504 1439 0.04      
8 A' 1474 1411 2.00      
9 A' 1449 1386 1.16      
10 A' 1277 1222 0.27      
11 A' 1227 1174 0.07      
12 A' 1098 1050 0.96      
13 A' 992 949 0.41      
14 A' 911 871 24.97      
15 A' 907 868 7.98      
16 A' 754 721 1.38      
17 A' 663 634 1.20      
18 A' 376 360 5.17      
19 A' 122 117 0.12      
20 A" 3265 3123 10.88      
21 A" 3145 3009 15.48      
22 A" 3078 2945 41.45      
23 A" 1465 1402 2.52      
24 A" 1303 1247 0.33      
25 A" 1259 1204 0.16      
26 A" 1225 1172 0.37      
27 A" 1202 1150 0.74      
28 A" 1046 1000 0.12      
29 A" 947 906 0.90      
30 A" 923 883 0.87      
31 A" 814 778 0.07      
32 A" 649 621 0.07      
33 A" 350 335 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 25438.5 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24337.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.34123 0.15199 0.11476

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.204 -0.425 1.096
C2 0.204 -0.425 -1.096
C3 0.000 0.625 0.000
C4 -0.030 -1.515 0.000
C5 -0.380 1.909 0.000
H6 -0.482 -0.403 1.960
H7 -0.482 -0.403 -1.960
H8 1.246 -0.433 1.466
H9 1.246 -0.433 -1.466
H10 -0.543 2.454 0.937
H11 -0.543 2.454 -0.937
H12 0.646 -2.386 0.000
H13 -1.075 -1.868 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.19251.53121.56392.64401.10333.13231.10572.76642.97843.60302.28972.2184
C22.19251.53121.56392.64403.13231.10332.76641.10573.60302.97842.28972.2184
C31.53121.53122.14001.33972.26472.26472.19552.19552.12622.12623.07892.7147
C41.56391.56392.14003.44242.29852.29852.22492.22494.11074.11071.10221.1032
C52.64402.64401.33973.44243.03273.03273.20593.20591.09651.09654.41603.8409
H61.10333.13232.26472.29853.03273.91961.79753.83763.03504.06943.00772.5178
H73.13231.10332.26472.29853.03273.91963.83761.79754.06943.03503.00772.5178
H81.10572.76642.19552.22493.20591.79753.83762.93303.43694.16072.51513.0982
H92.76641.10572.19552.22493.20593.83761.79752.93304.16073.43692.51513.0982
H102.97843.60302.12624.11071.09653.03504.06943.43694.16071.87505.07124.4547
H113.60302.97842.12624.11071.09654.06943.03504.16073.43691.87505.07124.4547
H122.28972.28973.07891.10224.41603.00773.00772.51512.51515.07125.07121.7974
H132.21842.21842.71471.10323.84092.51782.51783.09823.09824.45474.45471.7974

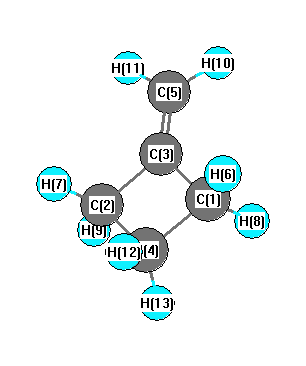
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.443 C1 C3 C5 134.025
C1 C4 C2 89.014 C1 C4 H12 117.327
C1 C4 H13 111.391 C2 C3 C5 134.025
C2 C4 H12 117.327 C2 C4 H13 111.391
C3 C1 C4 87.482 C3 C1 H6 117.634
C3 C1 H8 111.722 C3 C2 C4 87.482
C3 C2 H7 117.634 C3 C2 H9 111.722
C3 C5 H10 121.241 C3 C5 H11 121.241
C4 C1 H6 118.001 C4 C1 H8 111.763
C4 C2 H7 118.001 C4 C2 H9 111.763
H6 C1 H8 108.920 H7 C2 H9 108.920
H10 C5 H11 117.518 H12 C4 H13 109.173
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability