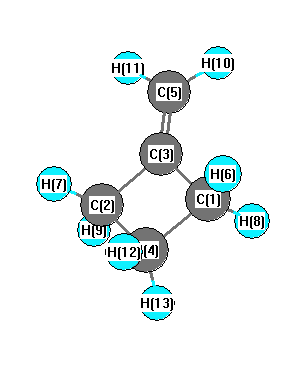
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3069 |
3051 |
5.59 |
|
|
|
| 2 |
A' |
3064 |
3046 |
35.56 |
|
|
|
| 3 |
A' |
3026 |
3009 |
14.64 |
|
|
|
| 4 |
A' |
3002 |
2984 |
41.39 |
|
|
|
| 5 |
A' |
2971 |
2954 |
10.68 |
|
|
|
| 6 |
A' |
1713 |
1703 |
24.63 |
|
|
|
| 7 |
A' |
1421 |
1412 |
0.01 |
|
|
|
| 8 |
A' |
1391 |
1383 |
1.78 |
|
|
|
| 9 |
A' |
1370 |
1362 |
1.28 |
|
|
|
| 10 |
A' |
1204 |
1197 |
0.23 |
|
|
|
| 11 |
A' |
1171 |
1165 |
0.11 |
|
|
|
| 12 |
A' |
1044 |
1038 |
0.39 |
|
|
|
| 13 |
A' |
963 |
958 |
0.45 |
|
|
|
| 14 |
A' |
869 |
864 |
0.60 |
|
|
|
| 15 |
A' |
853 |
848 |
32.27 |
|
|
|
| 16 |
A' |
729 |
725 |
2.68 |
|
|
|
| 17 |
A' |
647 |
643 |
1.73 |
|
|
|
| 18 |
A' |
373 |
370 |
5.71 |
|
|
|
| 19 |
A' |
108 |
107 |
0.04 |
|
|
|
| 20 |
A" |
3163 |
3145 |
10.57 |
|
|
|
| 21 |
A" |
3030 |
3013 |
15.71 |
|
|
|
| 22 |
A" |
2967 |
2950 |
41.55 |
|
|
|
| 23 |
A" |
1376 |
1368 |
4.25 |
|
|
|
| 24 |
A" |
1238 |
1230 |
0.82 |
|
|
|
| 25 |
A" |
1190 |
1183 |
0.10 |
|
|
|
| 26 |
A" |
1166 |
1160 |
0.80 |
|
|
|
| 27 |
A" |
1144 |
1137 |
1.25 |
|
|
|
| 28 |
A" |
998 |
992 |
0.12 |
|
|
|
| 29 |
A" |
919 |
914 |
0.69 |
|
|
|
| 30 |
A" |
883 |
878 |
0.87 |
|
|
|
| 31 |
A" |
778 |
774 |
0.09 |
|
|
|
| 32 |
A" |
613 |
609 |
0.09 |
|
|
|
| 33 |
A" |
347 |
345 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24399.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 24257.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.019 |
|
|
|
| 2 |
C |
0.019 |
|
|
|
| 3 |
C |
-0.162 |
|
|
|
| 4 |
C |
-0.043 |
|
|
|
| 5 |
C |
-0.035 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
0.029 |
|
|
|
| 8 |
H |
0.037 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.013 |
|
|
|
| 11 |
H |
0.013 |
|
|
|
| 12 |
H |
0.013 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.213 |
-0.522 |
0.000 |
0.563 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.945 |
-0.505 |
0.000 |
| y |
-0.505 |
-32.372 |
0.000 |
| z |
0.000 |
0.000 |
-30.808 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.355 |
-0.505 |
0.000 |
| y |
-0.505 |
-0.995 |
0.000 |
| z |
0.000 |
0.000 |
1.350 |
|
| Polar |
|---|
| 3z2-r2 | 2.700 |
|---|
| x2-y2 | 0.427 |
|---|
| xy | -0.505 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.692 |
-1.779 |
0.000 |
| y |
-1.779 |
9.821 |
0.000 |
| z |
0.000 |
0.000 |
8.027 |
<r2> (average value of r
2) Å
2
| <r2> |
117.196 |
| (<r2>)1/2 |
10.826 |