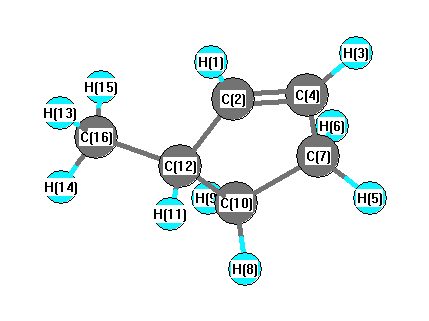
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3134 |
3116 |
23.86 |
|
|
|
| 2 |
A |
3107 |
3089 |
7.36 |
|
|
|
| 3 |
A |
3045 |
3028 |
22.03 |
|
|
|
| 4 |
A |
3044 |
3027 |
36.30 |
|
|
|
| 5 |
A |
3035 |
3017 |
26.35 |
|
|
|
| 6 |
A |
2996 |
2978 |
35.10 |
|
|
|
| 7 |
A |
2969 |
2952 |
29.09 |
|
|
|
| 8 |
A |
2962 |
2944 |
26.56 |
|
|
|
| 9 |
A |
2934 |
2917 |
42.35 |
|
|
|
| 10 |
A |
2900 |
2883 |
32.81 |
|
|
|
| 11 |
A |
1639 |
1629 |
1.61 |
|
|
|
| 12 |
A |
1421 |
1413 |
3.66 |
|
|
|
| 13 |
A |
1418 |
1410 |
4.97 |
|
|
|
| 14 |
A |
1416 |
1408 |
2.46 |
|
|
|
| 15 |
A |
1398 |
1390 |
1.15 |
|
|
|
| 16 |
A |
1342 |
1335 |
1.17 |
|
|
|
| 17 |
A |
1332 |
1325 |
3.81 |
|
|
|
| 18 |
A |
1283 |
1275 |
1.38 |
|
|
|
| 19 |
A |
1270 |
1262 |
1.06 |
|
|
|
| 20 |
A |
1256 |
1248 |
2.98 |
|
|
|
| 21 |
A |
1234 |
1226 |
0.95 |
|
|
|
| 22 |
A |
1178 |
1171 |
0.64 |
|
|
|
| 23 |
A |
1155 |
1148 |
0.26 |
|
|
|
| 24 |
A |
1107 |
1100 |
0.89 |
|
|
|
| 25 |
A |
1075 |
1069 |
4.91 |
|
|
|
| 26 |
A |
1061 |
1055 |
2.52 |
|
|
|
| 27 |
A |
1034 |
1028 |
2.89 |
|
|
|
| 28 |
A |
980 |
974 |
2.29 |
|
|
|
| 29 |
A |
937 |
932 |
2.13 |
|
|
|
| 30 |
A |
926 |
921 |
0.99 |
|
|
|
| 31 |
A |
921 |
915 |
0.32 |
|
|
|
| 32 |
A |
893 |
888 |
2.80 |
|
|
|
| 33 |
A |
838 |
833 |
2.39 |
|
|
|
| 34 |
A |
801 |
796 |
1.18 |
|
|
|
| 35 |
A |
736 |
732 |
1.38 |
|
|
|
| 36 |
A |
711 |
707 |
25.13 |
|
|
|
| 37 |
A |
565 |
562 |
4.31 |
|
|
|
| 38 |
A |
481 |
478 |
2.63 |
|
|
|
| 39 |
A |
333 |
331 |
0.03 |
|
|
|
| 40 |
A |
280 |
278 |
0.15 |
|
|
|
| 41 |
A |
234 |
233 |
0.02 |
|
|
|
| 42 |
A |
120 |
119 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30748.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30569.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.028 |
|
|
|
| 2 |
C |
0.029 |
|
|
|
| 3 |
H |
-0.024 |
|
|
|
| 4 |
C |
-0.041 |
|
|
|
| 5 |
H |
0.034 |
|
|
|
| 6 |
H |
0.025 |
|
|
|
| 7 |
C |
-0.006 |
|
|
|
| 8 |
H |
0.012 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
| 10 |
C |
0.007 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
C |
-0.118 |
|
|
|
| 13 |
H |
0.027 |
|
|
|
| 14 |
H |
0.027 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
| 16 |
C |
-0.011 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.011 |
-0.162 |
0.141 |
0.215 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.692 |
-0.284 |
-0.567 |
| y |
-0.284 |
-37.929 |
-0.136 |
| z |
-0.567 |
-0.136 |
-39.173 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.858 |
-0.284 |
-0.567 |
| y |
-0.284 |
0.503 |
-0.136 |
| z |
-0.567 |
-0.136 |
-1.362 |
|
| Polar |
|---|
| 3z2-r2 | -2.724 |
|---|
| x2-y2 | 0.237 |
|---|
| xy | -0.284 |
|---|
| xz | -0.567 |
|---|
| yz | -0.136 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.925 |
-0.350 |
-0.435 |
| y |
-0.350 |
9.733 |
-0.104 |
| z |
-0.435 |
-0.104 |
7.523 |
<r2> (average value of r
2) Å
2
| <r2> |
166.542 |
| (<r2>)1/2 |
12.905 |